Your new post is loading...

|
Scooped by
mhryu@live.com
August 12, 1:55 PM
|
Nitrogen fixation performed by rhizosphere bacteria has the potential to improve the sustainability of cereal crop cultivation. Deciphering the role of interspecies interactions on nitrogen fixation is crucial for devising strategies to enhance this process. To unravel the contributions of interspecies interactions, we constructed synthetic microbial communities syncom from the bottom-up that contain diazotrophic bacteria that fix nitrogen and maize rhizosphere bacteria that do not have this capability. Interactions that impacted nitrogenase activity via growth-independent mechanisms were prevalent in the system. Nitrogenase activity increased and eventually saturated as a function of the number of inoculated diazotrophs. Using a tailored machine learning model for microbiome dynamics and explainable artificial intelligence, we deciphered species contributions on nitrogenase activity and diazotroph growth. We identified a community containing Klebsiella variicola, Herbaspirillum seropedicae, and Stutzerimonas stutzeri as a starting point for developing microbial inoculants for cereal crops. Taken together, these results provide insights into the role of interspecies interactions on nitrogenase activity.
|
|
Scooped by
mhryu@live.com
August 12, 10:19 AM
|
Prime editing (PE) can make specific local changes to genomic DNA in living systems but its efficient application currently requires extensive optimization of PE guide RNA (pegRNA) sequences. Here we present OptiPrime, a machine learning model of PE efficiency based on current understanding of PE mechanisms. OptiPrime achieves state-of-the-art accuracy on PE efficiency prediction and enables prediction of nicking guide RNA (PE3) and dual pegRNA (twinPE) outcomes. We validate that OptiPrime has learned the determinants of mammalian mismatch repair (MMR) and is well suited for nominating MMR-evasive silent edits that improve PE efficiency. We demonstrate the use of OptiPrime in a variety of prospective therapeutic contexts in primary human and mouse cells. Lastly, we show that OptiPrime can be used to achieve streamlined and efficient in vivo correction of a pathogenic mutation in the brain of a mouse model of KIF1A-associated neurological disorder. We provide a webserver for OptiPrime ( https://optipri.me/ ) as a community resource. OptiPrime incorporates prime-editing biochemistry to predict prime-editing outcomes.
|
|
Scooped by
mhryu@live.com
August 12, 10:07 AM
|
Bacterial diadenylate cyclase (DAC) enzymes synthesize the nucleotide signal 3′3′ cyclic di-AMP (3′3′-c-di-AMP) to control osmoregulation, cell-wall homeostasis, and DNA-damage responses. Here we discover specialized roles for DAC enzymes in bacterial immunity and define a Panoptes-like system we name Panoptoo as a DAC-containing antiphage defense that guards against viral immune evasion. The Panoptoo protein PanS is a minimal DAC that constitutively synthesizes 3′3′ cyclic UMP-AMP (3′3′-cUA) or 3′3′-c-di-AMP to negatively regulate a partnering PanE S2TMβ membrane-targeting effector. We show that Panoptoo decoy signaling acts as a counter-defense to detect phage immune evasion proteins that inhibit nucleotide immune signals. A 1.5 Å crystal structure of PanS in complex with 3′3′-cUA explains how a symmetry break in the canonical DAC active site enables synthesis of asymmetric signaling molecules. Together, our results uncover a role for DAC domains in bacterial anti-phage defense and expand our understanding of nucleotide signaling in antiviral immunity.
|
|
Scooped by
mhryu@live.com
August 12, 9:50 AM
|
Post-translational modifications (PTMs) are chemical changes added to proteins after translation. These changes affect protein function and regulation, and their disruption is linked to disease-associated mechanisms. Because experimentally validating all possible modification sites is impractical, many computational predictors have been developed for PTM site prediction. In this work, we study whether a shared model can represent common residue-background patterns while learning modification-specific background-to-positive offsets. This framing is especially relevant for residues such as lysine (K), which can be acetylated, ubiquitinated, methylated, or sumoylated depending on the surrounding protein context. We propose an anchor-guided rectified flow matching framework for multi-type PTM site prediction from protein language model embeddings. For each PTM--residue pair, the model builds residue-background anchors from PTM-compatible unannotated residues and positive anchors from experimentally annotated modified residues. Given a candidate residue and target modification type, the model compares the residue embedding with these anchor sets and uses a rectified flow module to estimate a modification-conditioned background-to-positive offset. This offset is combined with anchor-based features and used for site scoring. We evaluate the framework on a dbPTM-derived benchmark covering six commonly studied PTMs: phosphorylation, acetylation, ubiquitination, methylation, sumoylation, and N-linked glycosylation. In the shared-model setting, our approach achieves a macro AUPRC of 0.4195, improving over the gated multi-anchor baseline of 0.4154, while independently trained per-modification models achieve 0.4353. These results suggest that multi-type PTM prediction can be modeled within a single shared framework by combining residue-background anchors with modification-conditioned offset features.
|
|
Scooped by
mhryu@live.com
August 12, 1:04 AM
|
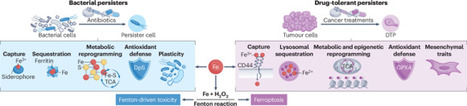
Mirroring bacterial persisters, cancer cells reprogram their iron homeostasis to acquire a drug-tolerant cell state that paradoxically confers vulnerability to ferroptosis. We propose a conserved principle, positioning iron as a driver of cell adaptations across mammalian malignant and microbial persistence. Bacterial persisters and drug-tolerant persister cells in cancer both show reprogrammed iron homeostasis and a vulnerability to ferroptosis. Agostinis and Rodriguez propose that this iron logic is a conserved principle with implications for therapeutics development.
|
|
Scooped by
mhryu@live.com
August 12, 12:41 AM
|
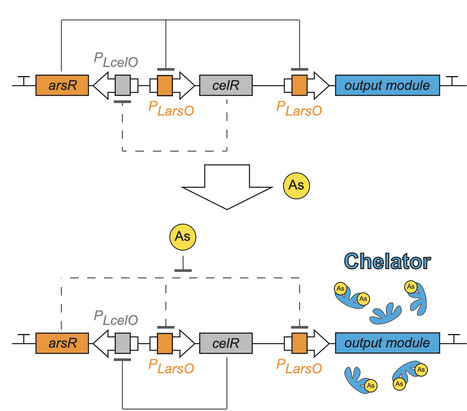
Chronic exposure to arsenite in food and water is a major global health concern, yet no practical strategies exist to prevent ingested arsenite from entering the body through the gastrointestinal tract. Here, we engineered E. coli Nissle 1917 (EcN) to sense and sequester arsenite in situ, creating a probiotic-based approach to reduce host absorption of arsenite. The system involved an arsenite-responsive genetic toggle switch that activated chelator expression upon exposure; after arsenite was removed, it sustained output under biostatic conditions but shut off during active cell division. We also engineered a nontoxic, high-affinity arsenite-binding protein as the chelator. The resulting strain efficiently removed arsenite in vitro while maintaining robust growth. A mass-transfer model guided in vivo dosing, and mouse studies showed that engineered EcN reduced arsenite entry into the bloodstream and promoted its fecal elimination. These findings support the conclusion that this engineered probiotic approach is promising for addressing toxic pollutants in the diet.
|
|
Scooped by
mhryu@live.com
August 12, 12:29 AM
|
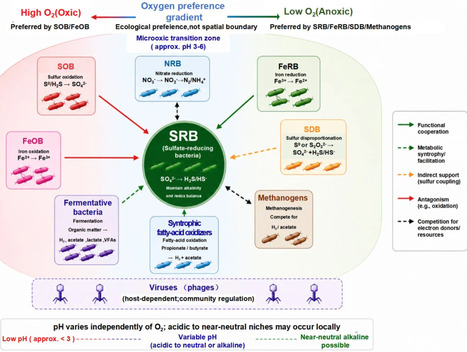
Sulfate-reducing bacteria (SRB) are key functional microorganisms in the bioremediation of acid mine drainage (AMD), simultaneously removing sulfate, generating alkalinity, and precipitating metal sulfides via dissimilatory sulfate reduction. AMD is typically characterized by low pH, high sulfate and metal concentrations, and limited organic carbon, imposing persistent stress on SRB growth, metabolism, and function. Environmental stressors include acidic conditions, heavy metal toxicity, low-temperature stress, and electron-donor limitation. These stressors impair membrane stability, enzymatic activity, and cellular energy conservation. Their combined effects increase maintenance requirements while limiting energy acquisition, ultimately reducing the range of environmental conditions under which SRB can sustain sulfate reduction. At the community level, SRB function is further modulated by complex microbial networks. Various functional groups, including fermenters, methanogens, and sulfur- and iron-cycling microorganisms, interact with SRB through cooperative, competitive, and regulatory processes that influence electron transfer, carbon turnover, and iron–sulfur transformations. Sulfate reduction in AMD depends on these community-level interactions, which can be disrupted under environmental stress and may reduce the stability of sulfate-reducing communities. Engineering strategies such as slow-release carbon supplementation, pH microenvironment optimization, conductive material amendment, mineral–microbe interface regulation, and immobilized reactor design can enhance SRB persistence by stabilizing the extracellular microenvironment, regulating electron flow, and spatially decoupling metabolic and mineralization interfaces. This review highlights how environmental stressors and microbial networks jointly regulate SRB function, emphasizes the roles of metabolic niche constraints and community resilience, and provides mechanistic insights for improving the stability and practical performance of SRB-based AMD treatment systems.
|
|
Scooped by
mhryu@live.com
August 12, 12:18 AM
|
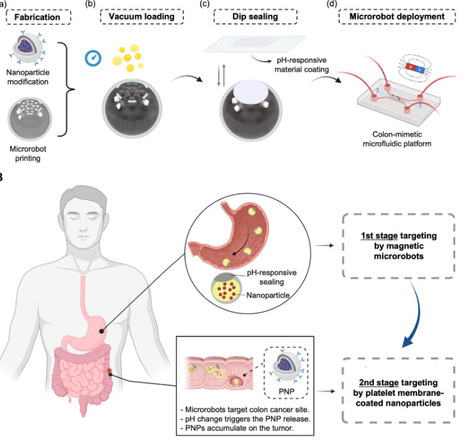
Targeted drug delivery in the gastrointestinal tract remains challenging because therapeutics must overcome multiple hierarchical barriers before reaching diseased tissue. Here, we present a multistage delivery platform that integrates magnetic microrobots, a pH-responsive protective coating, and platelet membrane–coated nanoparticles (PNPs) in one platform. A fillable design enables the formation of an internal magnetic layer for microrobot actuation, while the pH-responsive coating protects the cargo during transit and selectively degrades upon pH change, releasing cancer cell–targeting PNPs. In an in vitro colon cancer model that reproduces key gastrointestinal features, including flow, pH variation, and villi-like structures, this strategy increased nanoparticle retention and enhanced cancer cell cytotoxicity compared to nanoparticles administered alone. Ex vivo studies in porcine stomach and intestine further demonstrated robust locomotion on compliant and folded tissue surfaces. These results establish an environment-responsive hierarchical delivery strategy for more precise oral delivery in complex gastrointestinal settings.
|
|
Scooped by
mhryu@live.com
August 11, 11:57 PM
|
Plant roots are extremely heterogeneous and non-photosynthetic tissue that is drastically different from leaves. Pioneering studies have revealed the existence of pattern-triggered immune responses in roots; however, whether roots mount tissue-specific effector-triggered immunity (ETI) is a fundamental question in plant immunity. By utilizing inducible effector-expressing lines, we systematically characterize non-conventional physiological and molecular responses during ETI activation in roots. Root ETI seems to be weaker than leaf ETI, evidenced by restricted cell death in the transition zone and fewer number of differentially expressed genes (DEGs). Root ETI also shows much higher DEG overlap with Pep1-triggered immunity than with Flg22-triggered responses, which is the opposite trend in leaves. We find that both PEPR1/2 receptors and BIK1, WRKY42 are required for induced root ETI responses. By comparing root responses to wild type and the T3SS mutant of Ralstonia solanacearum in the Nd-1 ecotype, we confirm that natural root ETI responses show similarities to induced ETI. Our work reveals tissue-specific features of root ETI and provides new insights into engineering root disease resistance in agriculture. Roots, unlike leaves, are continuously exposed to a complex soil microbiome. Here the authors reveal root-specific effector-triggered immune responses distinct from those in leaves.
|
|
Scooped by
mhryu@live.com
August 11, 11:37 PM
|
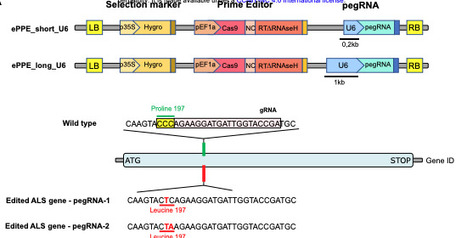
Since the development of CRISPR-based genome editing tools, a number of novel technologies have emerged. This includes Prime-Editing that acts as a search and replace genome editing tool. Prime-Editing has been deployed across multiple clades, including in a few flowering plants. Here, we report on the development of an efficient Prime Editor (PE) for the model bryophyte Marchantia. Initial tests were conducted on Acetolactate Synthase as a target and revealed an average efficiency above 40%. The system has been developed in the GoldenGate cloning system, facilitating construct design. The development of PE in Marchantia expands the Genome-Editing tools available for this emerging model in plant biology.
|
|
Scooped by
mhryu@live.com
August 11, 11:13 PM
|
3′,3'-cyclic diguanylate (cyclic di-GMP) and alternative cyclic di- and oligonucleotides are ancient highly conserved signaling molecules of bacteria and archaea, which can be present in metazoans up to humans. Their impact in fundamental behavioral modes and physiological and metabolic processes, the modular organization of their signaling cascades and the versatility and flexibility of their components in microbes in combination with their far-reaching effects including stimulation of the innate and adaptive immune response in humans makes these molecules and the respective signaling cascades promising targets in antibiofilm therapy, modulation of multicellularity and tools for treatment strategies and biotechnological applications. This review will thus describe the current state-of-the-art of applications and use as therapeutic targets of cyclic di- and oligonucleotides and the limitations and challenges in the application of those molecules and their use as targets. Future possibilities to successfully exploit those molecules and their signaling cascades equally as potential shortcomings are discussed.
|
|
Scooped by
mhryu@live.com
August 11, 10:30 PM
|
Model-based design in synthetic biology is limited because bioparts are typically characterised by relative metrics that vary across genetic and physiological contexts. To address this, we introduce a host-aware framework for quantitatively characterizing parts in combinatorial libraries of plasmid-based constitutive expression constructs. The approach integrates a digital twin of E. coli, conditioned on measured growth rate, with model-in-the-loop parameter identification to separate part-associated properties from host-dependent effects. Using structured combinatorial libraries, we identify mechanistically interpretable, transferable parameters for plasmid origins, promoters and ribosome binding sites. In particular, we define an intrinsic translation initiation capacity that captures the dominant RBS-associated contribution to translation while context-dependent expression emerges from host physiology and local sequence context. The resulting parameterisation accurately predicts protein synthesis across physiological conditions, supports incremental library expansion, and reveals localised failures of modularity, providing a scalable foundation for predictive host-aware design in synthetic biology. Model-based design in synthetic biology is limited because bioparts are typically characterised by relative metrics that vary across genetic and physiological contexts. Here, using combinatorial libraries of plasmid-based expression constructs and a host-aware E. coli digital twin, the authors identify intrinsic biopart parameters that provide a basis for more predictive model-based design of synthetic gene circuits.
|
|
Scooped by
mhryu@live.com
August 11, 4:26 PM
|
The gastrointestinal tract is rich in metabolic, immune, and microbiome-derived signals that can inform the design of live biotherapeutics and diagnosis of intestinal disorders. Engineered cell-based biosensors can tap into this molecular information and report on their environment, yet their development in gut-resident symbionts has been limited by a lack of validated sensor systems. Here, we present a generalizable pipeline that leverages bacterial transcriptional profiling to identify environment-responsive systems for biosensor engineering. Candidate Sensors Systems (CSSs) mined from healthy, disease, and in vitro transcriptomes were assembled into a barcoded library in Bacteroidaceae chassis and screened in high-throughput in vivo to identify responsive promoters. A unique Bacteroidales ECF-type sigma factor operon with ties to sphingolipid metabolism and flux was highly responsive in chemically-induced colitis models. The biosensor responded robustly to disease and returned to baseline upon recovery, establishing an in vivo-driven strategy for discovering functional biosensors in non-model gut-resident bacteria.
|
|
|
Scooped by
mhryu@live.com
August 12, 1:29 PM
|
We present a robust and versatile in vitro transcription (IVT) assay utilizing an optimized Broccoli RNA aptamer sequence. When paired with the fluorophore DFHBI-1T, this system enables real-time monitoring of multi-round transcription over several hours. We developed a dedicated plasmid backbone, pIVT3, designed for the streamlined insertion of diverse promoters. The system was validated using single-subunit T7 RNA polymerase and multi-subunit E. coli RNA polymerase; notably, E. coli enzyme activity remained strictly dependent on the presence of a σ-factor and a cognate promoter sequence. To maximize signal-to-noise ratios, we incorporated two rrnBT1 terminators upstream of the promoter of interest. This modification successfully eliminated background transcription in case of weak promoters (PlivJ) and prevented interference from read-through transcription in strong synthetic promoters (Ptrc*). Furthermore, we demonstrated the assays utility in drug discovery by testing the RNA polymerase inhibitor rifampicin, which showed time- and dosage-dependent inhibitory kinetics. Collectively, our results establish this Broccoli-based IVT system as a highly adaptable platform for quantifying promoter strength and screening small-molecule inhibitors of bacterial transcription.
|
|
Scooped by
mhryu@live.com
August 12, 10:18 AM
|
Cell surface display (CSD) elements are a major class of bioengineering modules, but systematic rules linking CSD sequence to functional potency are lacking. Here we develop DeepSCan, a suite of deep learning and artificial intelligence models to systematically map CSD sequence–function relationships to reliably capture potent elements’ conserved features and iteratively design and develop potent de novo CSD modules for mRNA antigen display. We experimentally quantify surface expression across >570 chimeric antigens, derive cell surface translocation strength labels for ~310 CSD elements and compile ~45 independent training datasets. We train three generations of DeepSCan models and evaluate their performance. Guided by these models, we computationally design 3,700 and experimentally validate approximately 120 generative CSDs, identifying 7 generative CSDs that match or exceed the cell surface translocation strength of the most potent naturally occurring CSDs. Enhanced surface displays are validated across multiple cell types and are functional in an antigen-specific CAR-T cytotoxicity assay. A systematic artificial intelligence-guided approach designs cell surface display elements for therapeutic applications.
|
|
Scooped by
mhryu@live.com
August 12, 10:02 AM
|
Bacteria and phages have co-evolved tit-for-tat strategies to combat one another for survival. To overcome phage infection, bacteria have developed various defense systems (the defensome), and in response, phages have evolved a counter-defensome to inhibit or evade such defensive strategies. While significant progress has been achieved on mechanistic, structural and functional aspects of the defensome, the study of the phage counter-defensome is relatively new, with very limited information available on the diversity and distribution of these systems across complex environmental phageomes. Here we present a large-scale analysis of the counter-defensome of 50,595 DNA phage population genomes reconstructed from soil, marine, and human gut environments. We observed substantial variation in the frequency and composition of the counter-defensome across phage families, which correlated with host range, habitat, and geographic context. A substantial fraction of the counter-defensome was clustered in islands, with gene expression restricted to a limited subset of families. Moreover, both phage counter-defense and bacterial-type defense families were detected across genomes of nucleocytoplasmic large DNA viruses, suggesting cross-clade horizontal gene transfer. Our results reveal the diverse counter-defense strategies across environmentally distinct viral communities and provide a foundation for uncovering novel mechanisms that shape conflicts and alliances in host-virus immunity networks.
|
|
Scooped by
mhryu@live.com
August 12, 9:39 AM
|
Rice paddies are major anthropogenic methane sources, yet the community interactions regulating emissions remain poorly resolved. This field experiment combined genome-resolved metagenomics, metatranscriptomics, and biogeochemistry to identify microbial cohorts directly and indirectly involved in methane cycling over a rice growing season. Cohorts routed reducing equivalents to different terminal electron-accepting processes and changed in abundance in response to depth and time-dependent shifts in environmental conditions. Methanogenic activity peaked late in the season in the rhizosphere and surface soil, whereas activity in 2-10 cm bulk soil peaked earlier, indicating that methanogenesis was temporally partitioned across soil compartments. Geobacter-like extracellular electron transfer was the most highly expressed alternate sink for reducing equivalents. Methane consumption was vertically partitioned between surface type I methanotrophs and deeper type II methanotrophs and anaerobic methane-oxidizing archaea. We infer that methane emissions reflect seasonal shifts in microbial cohorts that redirect methanogenic substrates through competing redox processes.
|
|
Scooped by
mhryu@live.com
August 12, 12:56 AM
|
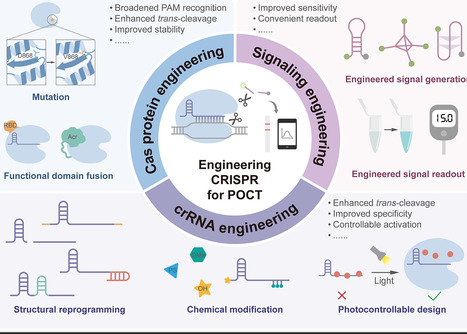
CRISPR-based molecular diagnostics have emerged as powerful and programmable platforms that enable sensitive and specific detection for disease management and epidemiological surveillance. Advances in CRISPR engineering and assay design are driving the emergence of next-generation detection platforms that are highly sensitive, rapid, and amenable to field deployment. These engineering breakthroughs have the potential to reshape point-of-care tests (POCT) and transform how emerging and persistent health threats are monitored in decentralized and resource-limited settings. Herein, we systematically review the recent advancements in CRISPR engineering strategies aimed at improving detection sensitivity and specificity, eliminating the dependence on preamplification, and enabling robust POC deployment. The discussed strategies encompass both the rational engineering of CRISPR ribonucleoproteins (RNPs) and the optimization of downstream signaling modules for molecular diagnostic applications. We further highlight key challenges and future perspectives that may inspire impactful research directions and accelerate the advancement of CRISPR engineering strategies toward robust, field-deployable POCT platforms.
|
|
Scooped by
mhryu@live.com
August 12, 12:39 AM
|
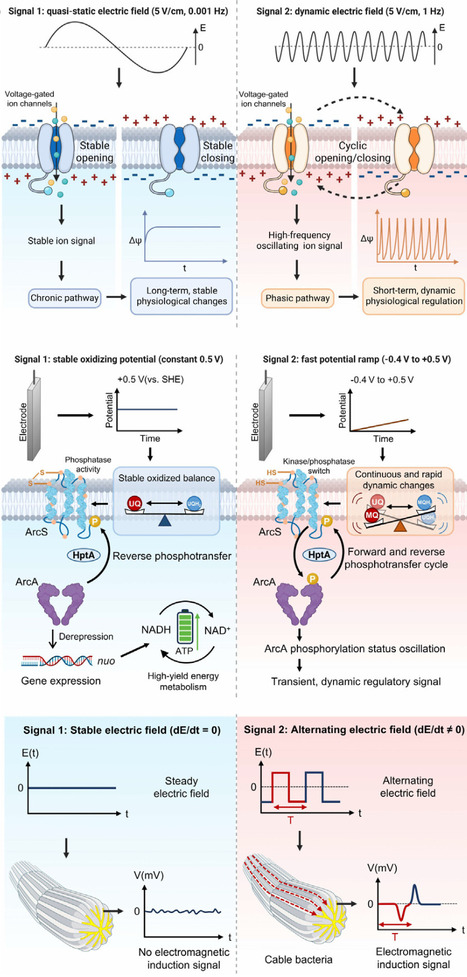
Electric cues (ECs) permeate microbial habitats, yet electrosensing, the ability of microorganisms to detect and respond to these cues, remains largely overlooked. We distinguish four principal EC types [electric fields (EFs), electrode potentials, redox signals, and electromagnetic induction] and map each to its biological sensing mechanism. Recent findings reveal that cable bacteria respond to dynamic EFs through electromagnetic induction, a candidate sensing mechanism that is absent from existing models. We synthesize conserved sensing strategies primarily in bacteria, with emerging evidence in eukaryotes, and assess applications in bioenergy, bioremediation, and electroceutical therapy. Realizing this potential requires moving beyond static-field models toward experimental frameworks that capture the full temporal complexity of natural electric landscapes.
|
|
Scooped by
mhryu@live.com
August 12, 12:18 AM
|
Magicpen’s genetically engineered plants have a heightened bioluminescence. While most traditional plant engineering uses the phenylalanine metabolism pathway, which requires five or six genes working in a coordinated way, the startup developed an efficient dual-gene system. industry Magicpen Bio
|
|
Scooped by
mhryu@live.com
August 12, 12:05 AM
|
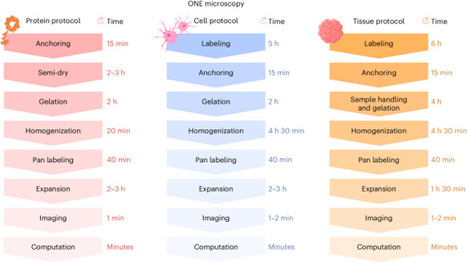
The introduction of expansion microscopy (ExM), a decade ago, marked a shift in super-resolution imaging, by physically separating fluorophores to bypass the diffraction limit. Numerous ExM developments have extended the method’s reach since, yet molecular-scale resolution remained inaccessible. We recently developed one-step nanoscale ExM, which combines ExM with fluctuation-based super-resolution analysis to enable the direct visualization of individual protein shapes, using conventional fluorescence microscopes, a capability that was previously limited to cryo-electron microscopy and averaging-based techniques. Here we provide detailed procedures for gel embedding, labeling, expansion, image acquisition and data analysis. We also introduce a stable, user-friendly software package for efficient fluctuation analysis. Although one-step nanoscale ExM is broadly applicable to a range of samples, including purified proteins, cells and tissues, its most distinctive contribution lies in making single-protein shape analysis accessible and reproducible. Overall, we provide a practical framework for protein imaging on conventional equipment. An approach for combining expansion microscopy with fluctuation-based super-resolution analysis enables the visualization of individual proteins via conventional fluorescence microscopy.
|
|
Scooped by
mhryu@live.com
August 11, 11:42 PM
|
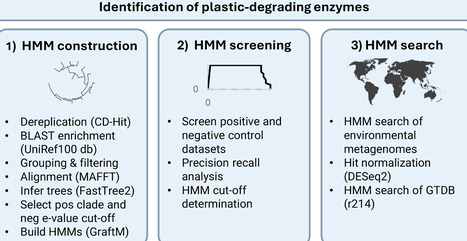
Plastic waste pollution is a global issue that threatens biodiversity and human health. Current plastic waste management practices are not sufficient to keep up with increasing plastic production rates. Microorganisms have the capacity to degrade different types of bio-based and synthetic plastics through enzymatic reactions, offering an alternative solution to traditional plastic recycling techniques. A limited number of plastic-degrading enzymes have been identified, sequenced and characterized; however, studies exploring the distribution of homologues of these enzymes across habitats and microbial taxa have remained scarce. Here, we applied analytical techniques to search for genes encoding potential plastic-degrading enzymes in environmental metagenome datasets and genomes of the Genome Taxonomy Database (GTDB) to explore the geographic and taxonomic distribution patterns of plastic-degrading microorganisms. Hidden Markov Models (HMMs) were constructed from amino acid sequences of known, experimentally verified and putative plastic-degrading enzymes. The HMMs were applied to landfill, soil, river, lake and ocean metagenomes and all archaeal and bacterial genomes in the GTDB. An abundance of hits was discovered across aquatic and terrestrial metagenomes with the majority occurring in polluted rivers, polar oceans and deep ocean samples. GTDB hits were mainly consistent with known plastic-degrading microbial lineages, while also revealing potential plastic-degrading archaeal taxa. The results of this study may be able to assist in the discovery of novel plastic-degrading enzymes for application in plastic waste biodegradation solutions.
|
|
Scooped by
mhryu@live.com
August 11, 11:30 PM
|
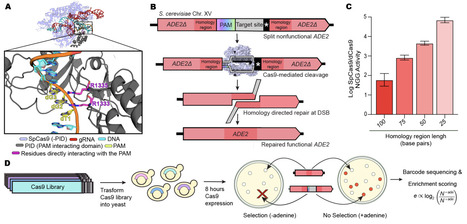
The therapeutic potential of CRISPR-Cas9 genome editing is fundamentally constrained by the requirement for specific short DNA sequences (PAMs) flanking the target site, limiting access to many clinically relevant genomic loci. This stringent PAM requirement is particularly problematic in applications which require precise positioning, such as base editing and allele-specific editing. Although PAM-relaxed variants have expanded the targetable genome, they incur trade-offs in on-target activity, off-target editing, and cleavage kinetics. This highlights an unmet need for variants that are re-targeted to alternative PAMs in order to maintain the specificity and enzymatic performance inherent to stringent dinucleotide PAM recognition. To overcome these limitations, we developed a yeast selection platform capable of engineering SpCas9 variants with re-specified PAM recognition. Using a clinically relevant Huntington's disease gene (HTT) SNP as a proof-of-concept target, we engineered variants with reciprocal NGC and NGT PAM selectivity, as a step toward allele-specific editing in a large percentage of Huntington's disease patients. These yeast-selected SpCas9 variants retained their modified activity across multiple endogenous HEK293T loci, demonstrating that this specificity is robust across diverse genomic contexts. The variants surpassed PAM-broadened variants on their respective on-target PAM while displaying broad loss of activity across alternative PAMs, effectively re-specifying PAM recognition toward a single dinucleotide sequence. Retargeted variants recovered on-target cleavage kinetics approaching wild-type SpCas9, even under competing substrate conditions, demonstrating that PAM re-specification can simultaneously restore catalytic efficiency and improve specificity. Beyond NGC and NGT, we leveraged our high-throughput platform to engineer Cas9 with re-specified activity across multiple additional non-canonical PAMs in yeast, further demonstrating its utility as a general and programmable framework for expanding the therapeutic reach of precision genome editing.
|
|
Scooped by
mhryu@live.com
August 11, 11:09 PM
|
The 21st annual International Genetically Engineered Machine (iGEM) Grand Jamboree took place on the 23rd to 26th of October 2024 at the Paris Expo in Paris, France. It was the final event of the year-long iGEM Competition in microbial synthetic biology, at which more than four hundred high school and university student teams from around the world presented their projects and competed for prizes. At the core, each project applied molecular cloning and microbial engineering to tackle pressing healthcare, human advancement, and environmental challenges. Herein we highlight the efforts of four projects from the latter category: Stockholm, Thessaly, KU Leuven, and Copenhagen, whose work exemplifies the type of research projects presented by iGEM teams each year in advancing sustainability. Through the application of basic principles of synthetic biology to a diverse range of problems, the projects demonstrated the promise of synthetic biology in addressing and indeed resolving a broad spectrum of sustainability challenges.
|
|
Scooped by
mhryu@live.com
August 11, 5:18 PM
|
Accurate detection of off-target activity in primary human cells is crucial for ensuring the safety of gene therapies, yet existing methods often lack sufficient sensitivity. To address this limitation, we develop Tracking-seq2, an advanced technology that integrates exogenous 5′ → 3′ exonuclease treatment and non-homologous end joining (NHEJ) pathway inhibitors with the original Tracking-seq. Tracking-seq2 exhibits enhanced sensitivity in profiling off-target sites of diverse genome editors—including Cas9, Cas12a, cytosine base editors (CBEs), adenine base editors (ABEs), and prime editors (PEs). Critically, Tracking-seq2 is directly applicable to clinically relevant primary human cell types, such as T cells and CD34+ hematopoietic stem and progenitor cells (HSPCs). Furthermore, our findings reveal that genomic variations drive distinct off-target heterogeneity across different individuals, highlighting the necessity for personalized safety assessment in clinical genome editing applications. Tracking-seq2 provides a robust platform for sensitive off-target detection in primary cells, with sensitivity comparable to or exceeding current state-of-the-art methods. Ensuring safe gene editing requires precise detection of unintended DNA edits. Here, authors develop Tracking-seq2, a highly sensitive method to profile off-targets for diverse genome editors, and reveal distinct off-target effects across individuals in primary human cells.
|