Your new post is loading...

|
Scooped by
mhryu@live.com
November 12, 12:11 PM
|
Anti-CRISPR systems are emerging as essential regulators of CRISPR-Cas technologies, providing a controllable “off-switch” for genome editing, transcriptional regulation, and gene-drive applications. This review surveys the expanding landscape of anti-CRISPR strategies, including naturally occurring Acr proteins, engineered Acr switches, small-molecule inhibitors, and nucleic acid-based inhibitors. Acr proteins, discovered in phages and other mobile genetic elements, exploit diverse mechanisms such as blocking CRISPR-Cas complex assembly, preventing target DNA or RNA binding, or directly inhibiting catalysis. Recent work has revealed additional noncanonical mechanisms, including RNA mimicry, host-factor hijacking, and enzymatic modifications of Cas proteins. In parallel, computational screening and machine-learning methods have accelerated discovery of Acr families and enabled de novo design of Cas13 inhibitors. This review also examines engineered control systems that make Acr activity conditional on chemical, light, miRNA, protease, or other environmental cues, enabling spatial and temporal regulation of CRISPR function. Small-molecule inhibitors and nucleic-acid inhibitors offer complementary CRISPR control, including the potential for improved delivery, tunable dosing, and modular design, though further assessment in therapeutic applications is necessary. Across applications, anti-CRISPRs have shown utility in improving editing precision, reducing toxicity, constraining gene drives, controlling CRISPRi/CRISPRa systems, and enhancing safety in therapeutic and agricultural contexts. Despite substantial progress, important challenges remain, including delivery, specificity across natural and engineered CRISPR variants, and governance. Future efforts will likely focus on rational design, multiplexed inhibition, and integration of anti-CRISPR systems into existing CRISPR-Cas therapeutic and environmental frameworks. As CRISPR continues to expand beyond the laboratory into medicine, agriculture, and ecological engineering, anti-CRISPR technologies will play a central role in enabling safer and more precise genome engineering.
|
|
Scooped by
mhryu@live.com
November 12, 11:31 AM
|
RNA structural motifs (RSMs) are recurrent three-dimensional RNA structural elements characterized by conserved three-dimensional geometries and often associated with specific molecular functions. Their identification, validation, and functional characterization frequently require labor-intensive manual inspection. However, existing molecular visualization tools provide limited support for motif-centric exploration, making large-scale visualization and comparative analysis of RNA motifs inefficient and time-consuming. To address this limitation, we present RSMViewer, an open-source PyMOL plug-in that integrates heterogeneous motif annotations and supports batch visualization and efficient structural superimposition of RNA motif instances. RSMViewer streamlines the exploration and comparative analysis of RNA structural motifs, substantially reducing manual effort and facilitating downstream structural and functional interpretation.
|
|
Scooped by
mhryu@live.com
November 11, 11:24 PM
|
RNA structures are active regulators of bacterial translation. Here, we identified a naturally occurring translation-enhancing small open reading frame (tesORF) that controls the expression of the flagellar operon fliFGHIJK in Salmonella enterica and other motile Enterobacterales. Nucleotide-resolution ribosome profiling revealed this previously unannotated 54-nucleotide ORF immediately upstream of fliF. Genetic, transcriptomic, and biochemical analyses support a model in which tesORF translation counteracts an inhibitory RNA structure, exposing the fliF ribosome-binding site and promoting translation initiation independently of the encoded peptide sequence. Disruption of tesORF translation reduced expression of the downstream operon and impaired motility and virulence-associated phenotypes in Escherichia coli, Salmonella, and Yersinia. Together, our findings reveal how translation of a conserved small ORF regulates an entire flagellar operon through RNA structural remodelling, with consequences expending to the wider flagellar regulon, linking RNA structural dynamics to bacterial motility and virulence.
|
|
Scooped by
mhryu@live.com
November 11, 11:00 PM
|
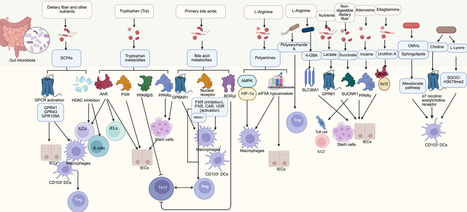
Intestinal immune tolerance enables the host to coexist with commensal microorganisms and dietary antigens while retaining effective defense against pathogens. The review describes mechanisms underlying the development and maintenance of immune tolerance and how the intestinal barrier and local immune system cooperate to maintain immune homeostasis. Particular attention is given to microbial metabolites, and we summarize mechanisms by which they act as key signals between the gut microbiota and the host, regulating barrier function and immune cell responses. We highlight the dynamic, context-dependent nature of metabolite-mediated immune regulation, influenced by local exposure and host factors including age, genetic background, and inflammatory or infectious conditions. Additional metabolites and regulatory pathways are discussed to broaden understanding of microbe-host interactions. Microbial metabolites may serve as potential biomarkers and therapeutic targets for inflammatory bowel disease and other immune-related disorders. However, broad clinical application requires further human validation and more precise intervention strategies.
|
|
Scooped by
mhryu@live.com
November 11, 5:59 PM
|
Antimicrobial resistance (AMR) is an emerging global health threat responsible for antibiotics being ineffective, leading to persistent infections, increased mortality, and healthcare costs. ESKAPE (Enterococcus faecium, Staphylococcus aureus, Klebsiella pneumoniae, Acinetobacter baumannii, Pseudomonas aeruginosa, and Enterobacter species) pathogens have a higher priority worldwide for the development of therapeutic approaches other than traditional antibiotics. Quorum sensing (QS) links population-dependent gene expression with a mechanistic understanding of bacterial communication, virulence regulation, and biofilm-associated persistence. Major QS systems across ESKAPE pathogens are Agr and accessory gene regulator networks in Gram-positive bacteria; LuxS/AI-2 signaling; the LuxI/LuxR-family systems in Gram-negative bacteria; the Las, Rhl, and Pseudomonas quinolone signal (PQS) networks in P. aeruginosa; and the AbaI/AbaR system in A. baumannii. This review focuses on QS regulatory mechanisms in ESKAPE pathogens and discusses the novel approaches to target QS-associated virulence and biofilm formation. A review of the literature on QS signaling pathways, biofilm regulation, and anti-QS therapies was performed with special focus on combating strategies like plant-derived phytochemicals, synthetic QS inhibitors, enzymatic quorum quenching, probiotics, and combinations of QS-targeted agents with conventional antibiotics. QS systems are involved in virulence regulation, adhesion, biofilm maturation, and bacterial persistence. Plant-based compounds such as flavonoids, alkaloids, terpenoids, and polyphenols have been shown to have potential to interfere with QS-related processes. Synthetic inhibitors, quorum-quenching enzymes, probiotics, and antibiotic combination strategies are also promising approaches. By integrating QS regulation with virulence, biofilm biology, and antimicrobial response across ESKAPE pathogens, this review provides a comparative framework to guide future target prioritization, diagnostic development, and evaluation of adjunctive anti-virulence strategies. Mechanistic validation, standardized cross-pathogen comparisons, and long-term resistance-evolution studies are needed to establish the therapeutic readiness and clinical value of these approaches.
|
|
Scooped by
mhryu@live.com
November 11, 5:00 PM
|
Genetic circuit design is central to synthetic biology, yet achieving predictable and programmable circuit performance remains challenging due to diverse circuit- and host-dependent factors. Thus, a systematic perspective is needed to integrate these factors across multiple design layers. In this review, we establish a hierarchical framework for tuning genetic circuits as an engineering process spanning biological part design, DNA synthesis and assembly, carrier selection, host-cell implementation, and cross-host transfer. We further discuss how each layer contributes to circuit behavior and shapes trade-offs among key performance properties. By integrating hierarchical design strategies with iterative troubleshooting, this review provides a practical roadmap for tuning genetic circuits toward more predictable and reliable performance.
|
|
Scooped by
mhryu@live.com
November 11, 3:22 PM
|
Classical small-molecule drugs have traditionally relied on an occupancy-driven paradigm that requires tight and sustained binding to a well-defined pocket on the target protein. Targeted protein degradation (TPD) approaches have emerged to overcome the limitations of traditional pharmacology to promote the selective degradation of disease-associated proteins, harnessing the central proteostasis pathways in eukaryotic cells: the ubiquitin proteasome system and the autophagy-lysosomal machinery. Thus, TPD exploits the idea of reprogramming cellular proteostasis for therapeutic benefits, dictating the fate of the target protein by means of degrader molecules that induce proximity between a target protein and components of the degradation machineries. The present review gives an overview of the molecular mechanisms of cellular proteostasis and how they are exploited for drug discovery purposes through TPD strategies. We describe some of the most explored degrader modalities, such as molecular glues, proteolysis targeting chimeras, and lysosomal targeting chimeras, as well as new modalities of degraders, highlighting their major advantages and limitations, and discuss the challenges associated with their development, including ligand identification, target degradability, or the possibility for their rational design. Furthermore, we consider recent advances in the design of these degraders to potentially address those limitations. Together, these insights established TPD as a transformative pharmacological modality that revolutionized drug discovery campaigns since it expands the druggable proteome, offering numerous advantages over traditional drugs.
|
|
Scooped by
mhryu@live.com
November 11, 3:05 PM
|
Living genetically engineered microorganisms (GEMs) are increasingly used in consumer products and environmental applications, yet U.S. biotechnology regulation relies on statutes that predate their development or are designed for other types of products. This article examines oversight of living GEMs through 3 case studies of consumer and environmental GEM applications. Across these cases, federal premarket review was very limited or absent because oversight depended on distinctions such as the type of modification, the developer’s product classification, or marketing claims. These regulatory pathways may not always align uniformly with the potential health and environmental risk profiles. While not all GEMs require the same level of oversight, current regulatory gaps have allowed for the large-scale, open release of these GEMs without substantial federal oversight. The article recommends baseline federal notification, changes to statutes or regulations to strengthen regulatory foundations, greater transparency, independent evaluation, and post-market monitoring, while also preserving efficient pathways for lower-risk applications. These enhancements to the regulation of GEMs will become increasingly important for effective risk governance as consumer and environmental products incorporate advanced synthetic biology approaches.
|
|
Scooped by
mhryu@live.com
November 11, 2:08 PM
|
RNA-derived 2′,3′-cyclophosphate-terminated nucleotides act as immune signalling molecules in prokaryotes and eukaryotes, but how plants perceive them remains unknown. Sucking insects inject digestive enzymes including RNases into the host plant to acquire nutrients. Here we demonstrate that NIB1, a glycine-rich RNA-binding protein in rice, senses 2′,3′-cyclic nucleotide monophosphates (2′,3′-cNMPs; 2′,3′-cAMP and 2′,3′-cGMP) and adenylyl-(3′→5′)-guanosine 2′,3′-cyclic phosphate (ApG>p) produced by a salivary T2 ribonuclease of green stink bug during feeding. Binding of the RNase products promotes formation of nuclear condensates containing NIB1, which recruit the core immune regulator Enhanced Disease Susceptibility 1 (EDS1) and activate defence, potentially contributing to the immune signalling of Toll/interleukin-1 receptor (TIR) proteins that also produce the nuclease products. Exogenous application of 2′,3′-cAMP, 2′,3′-cGMP or ApG>p elicits NIB1 and EDS1-mediated immune responses. Our findings unveil a conserved pathway in plants, in which glycine-rich RNA-binding proteins perceive the nuclease-generated second messengers to launch defence and point to a potential link between plant TIR nuclease activity and EDS1 signalling. The rice RNA-binding protein NIB1 detects 2′,3′-cyclic nucleotide products from insect saliva RNase, forming nuclear condensates that recruit EDS1 to activate defence, revealing a conserved messenger-based immune pathway in plants.
|
|
Scooped by
mhryu@live.com
November 11, 1:49 PM
|
Saccharomyces boulardii is the only yeast probiotic widely used in clinical practice and consumer products. It combines gastrointestinal robustness, intrinsic antibiotic tolerance and eukaryotic protein-processing capacity, making it an attractive chassis for engineered probiotics, in situ biomanufacturing and live biotherapeutic delivery. However, host-specific characterization of regulatory elements, candidate genomic integration sites, and multicopy integration strategies remains limited in the clinically used S. boulardii CNCM I-745 background. Here, using S. boulardii CNCM I-745 as the host, we developed and characterized a genetic engineering toolkit that integrates transcriptional control elements with genome-integration strategies. We first quantified the activities of 20 endogenous constitutive promoters, two inducible promoters and six terminators. We then established a markerless CRISPR-Cas9 workflow and evaluated 10 chromosomal integration loci; six of which showed Editing efficiency above 80% while supporting comparable transgene expression without detectable growth penalties. Finally, we constructed an rDNA-repeat-based multicopy integration system that increased transgene expression by 9.8- to 22.6-fold relative to single-copy integration and revealed a significant positive but non-proportional association between estimated copy number and reporter output. Together, this toolkit provides S. boulardii with a set of tunable promoters and terminators, candidate chromosomal integration sites compatible with a markerless CRISPR-Cas9 workflow. These resources expand the genetic accessibility of this probiotic yeast and provide a foundation for developing engineered S. boulardii strains with enhanced probiotic, therapeutic and biomanufacturing functions.
|
|
Scooped by
mhryu@live.com
November 11, 1:23 PM
|
Bacterial adherence to gut epithelial cells likely involves host genetic components. We isolated a gut commensal bacterium, Lelliottia jeotgali, that can robustly colonize C. elegans through attachment to intestinal epithelial cells. Here, we developed a novel screen in C. elegans to identify host genes required for microbiome bacterial colonization. Using transgenic L. jeotgali strain expressing tdTomato, we conducted forward genetic screens in C. elegans to select for mutants that completely lost colonization. We identified multiple allelic mutations in five different C. elegans genes, termed knockout of Lelliottia adherence (kola) mutants, with a near complete loss of adherence. Each of kola-1-5 is uncharacterized and predicted to encode for a secreted or membrane-bound protein. We also identified gly-5, a GalNAc transferase that initiates O-linked glycosylation. Overall, we describe the first C. elegans screen to identify host factors that control microbiome bacterial colonization in the gut.
|
|
Scooped by
mhryu@live.com
November 11, 1:13 PM
|
RNAs are emerging as promising therapeutic targets, and small-molecule binding can modulate their structure and function, creating opportunities for therapeutic development. Accurate identification of RNA small-molecule binding sites is therefore important for understanding RNA-ligand interactions and facilitating RNA-targeted drug discovery. However, the structural complexity of RNA and the limited availability of experimentally characterized RNA-ligand complexes pose challenges for computational binding-site prediction. In this study, we propose RSMBind, a graph attention-based deep learning framework for nucleotide-level prediction of RNA-small molecule binding sites. The model integrates geometric descriptors of the local three-dimensional environment with pretrained RNA language model embeddings to capture complementary structural and sequence information. We investigate three pretrained RNA language models RNA-BERT, ERNIE-RNA and RNA-FM and assess the contributions of sequence representations and geometric features through feature-level ablation experiments. The model is developed using the TR60 and HARIBOSS datasets and evaluated on four independent benchmark datasets: RB9, TE18, TL12 and JL10. RSMBind achieves the highest MCC among the compared methods on three of the four benchmarks, with relative improvements of 17.1% on RB9, 7.2% on TL12 and 16.0% on JL10 over the best previously reported results, while achieving competitive performance on TE18. Furthermore, RSMBind retains predictive capability when using only pretrained RNA language model embeddings, highlighting their potential utility when explicit tertiary structural features are unavailable. Structural analysis indicates differences in prediction performance across RNA structural regions, with loop-associated binding sites generally being captured more effectively than those embedded within stem regions. Additionally, 100 ns molecular dynamics simulations of selected RNA-ligand complexes provide insights into the relationship between predicted binding probabilities and dynamic ligand proximity, further highlighting the role of conformational dynamics in evaluating structure-based predictions. The source code is available at [ https://github.com/sharma-nandita/RSMBind].
|
|
Scooped by
mhryu@live.com
November 11, 1:06 PM
|
A key parameter in the experimental design of RNA-seq projects is the choice of sequencing depth. Considering a limited budget, one needs to find a trade-off between the number of samples and the sensitivity of the analysis, particularly concerning lowly expressed genes. While previous studies have proposed a lower bound for the comprehensive analysis of differential gene expression, for the analysis of alternative splicing, it has only been proposed for human adipose tissue. However, alternative splicing differs across tissues and conditions. We analyzed publicly available and newly generated deep-sequenced paired-end RNA-seq samples (between 150 and >500 million reads, read length 50–150 bp) from human buffy coat cells and diverse sets of tissues, including gluteal subcutaneous fat, heart, and hypothalamus. Our results show that the sequencing depth typically used in published cohorts is not sufficient to comprehensively capture the landscape of alternative splicing. This motivates the use of deeper sequencing or long-read technologies in future studies. Toward this goal, we offer guidelines and a website (https://sequencing-depth4splicing.com/) for choosing a suitable sequencing depth.
|
|
|
Scooped by
mhryu@live.com
November 12, 12:08 PM
|
DNA storage has emerged as a promising approach for long-term digital information preservation owing to its ultrahigh storage density, long-term stability, and potential for passive long-term preservation without continuous electrical power under appropriate storage conditions. Recent advances in DNA synthesis, sequencing, and molecular manipulation technologies have accelerated the development of the field, broadening the scope of research from sequence-based encoding to biochemical channel modeling, system implementation, structured DNA storage, and information management. However, existing studies are often focused on individual components of DNA storage systems, and the relationships among these components remain insufficiently characterized from a computational and system-level perspective. This structured narrative review examines selected representative studies based on their technical importance and relevance to a computational and system-level analytical framework for DNA storage. We first outline the development of sequence-based and non-sequence-based DNA storage technologies, highlighting the expansion of information representation beyond nucleotide sequences to encompass multidimensional molecular structures and states. We then examine representative computational tools and system-level platforms, including coding toolchains, biochemical channel modeling, automated storage platforms, and user-oriented information management systems. We present an analytical framework comprising 5 interconnected functional layers: information representation, encoding and decoding, biochemical channel, system implementation, and access and management. Using this framework, we analyze the relationships among existing technologies, current technical challenges, and emerging research directions. DNA storage is evolving from a high-density storage technology toward an integrated molecular information system that supports information representation, storage, retrieval, and management. Further progress will require coordinated advances in multidimensional information representation, content-based querying, cross-layer system optimization, and interdisciplinary research to support the development of practical DNA-based information systems.
|
|
Scooped by
mhryu@live.com
November 11, 11:49 PM
|
Biopharmaceuticals, including antimicrobial peptides and engineered protein-based vaccines, represent key products of the biotechnology sector. However, their reliance on large-scale manufacturing and cold-chain logistics limits deployment in field and resource-limited settings. To address this gap, we developed SporeBot, a platform that exploits the intrinsic stability of Bacillus subtilis spores for versatile and on-demand biopharmaceutical production. The platform comprises polymeric microcapsules that encapsulate engineered spores. These spores exhibit robust stability, retaining engineered functionality under harsh conditions and extended storage. Upon activation, germinated cells secrete the target proteins, which diffuse across a polymeric mesh that physically separates the cellular factory from the secreted product. We validate this platform through the production and functional testing of antimicrobial peptides and vaccine antigens. Our approach resolves key manufacturing and distribution constraints of protein therapeutics and offers a deployable solution for on-demand biomanufacturing in both developed and resource-limited settings.
|
|
Scooped by
mhryu@live.com
November 11, 11:07 PM
|
Proteins often assemble as higher order complexes, underlying a diversity of molecular structures and biological functions. The emergence of protein-protein interactions between binding partners is generally considered to be the primary driver of oligomerization. However, ligand binding has also been demonstrated to affect higher-order assembly, though this phenomenon is not widely considered as a primary driver of oligomerization. Here, we describe the mechanism by which different rubisco oligomeric states are induced via ligand binding. By expanding the phylogenetic sampling of a unique clade of rubisco, we discover a range of oligomeric responses to ligand binding across the phylogeny and identify the structural features responsible for shifts in the oligomeric state that ultimately provide a path to evolving new quaternary structures. Here, we identify a ligand-dependent mechanism giving rise to new protein assemblies, in contrast with the typical process of evolving new protein-protein interfaces directly. Our findings describe a means by which binding and entrenchment of a ligand enables an increase in structural complexity. Proteins often assemble into complexes, and while binding partners typically drive this, ligand binding can too. Here the authors show ligand binding drives distinct rubisco assemblies, revealing an evolutionary path to greater structural complexity.
|
|
Scooped by
mhryu@live.com
November 11, 10:50 PM
|
Bacterial pangenomes provide a shared reservoir of antiphage defenses, yet the ecological and functional constraints shaping this reservoir remain unclear. Here, we combine comparative genomics of 206 Xanthomonas oryzae pv. oryzae (Xoo) genomes with cross-infection assays of 50 strains against 36 phages and heterologous expression of 24 defense systems in a phage-sensitive Xoo chassis. The Xoo pan-defensome comprises a conserved core of 9 subtypes and a variable layer of 35 accessory and cloud subtypes structured by host phylogeography. Phages are more infectious on sympatric than allopatric hosts, consistent with local adaptation. Variable systems show stronger apparent activity than core systems, whereas resistance breadth is not predicted by defense system number. Co-expression of representative co-occurring and mutually exclusive system pairs reveals synergistic and antagonistic interactions underlying non-random repertoire assembly. Together, these findings extend the pan-immunity framework by showing that population-level immune reservoirs are shaped by both ecological structure and functional compatibility. In this study, the authors use comparative genomics and cross-infection assays to show a spatially and evolutionarily structured bacterial immune reservoir in which interactions among defense systems can reinforce or undermine phage resistance, refining the pan-immunity framework.
|
|
Scooped by
mhryu@live.com
November 11, 5:13 PM
|
Transcription factors (TFs) bind specific DNA sequences to control gene expression. Modulating TF activity is of considerable therapeutic interest but very few TFs have been drugged successfully. TF DNA-binding interfaces are large and most TFs have no known allosteric ligands or allosteric sites. Here we introduce TF-MAPs, a scalable and general platform to chart complete functional maps of sequence-specific DNA-binding proteins. We apply TF-MAPS to three human disease TFs: HNF1A, FOXG1 and FOXP1. Allostery in all three proteins is distance-dependent, anisotropic and couples the DNA-binding interface to the protein surface. The allosteric surfaces of FOXG1 and FOXP1 are quite diverged, revealing a potential for protein-specific regulation in protein families. All three TFs have potentially druggable small pockets where mutations affect DNA binding, either within the binding interface or distant from it. Application of TF-MAPS will potentially lead to charting functional and allosteric maps for hundreds of human TFs and other DNA-binding proteins to understand DNA recognition and TF regulation, to interpret clinical variants mechanistically and to guide the development of TF-modulating therapeutics. Li and Lehner present TF-MAPS, a scalable platform to chart functional maps of sequence-specific DNA-binding proteins. The authors map allostery in three transcription factors implicated in human disease and reveal potentially druggable small pockets.
|
|
Scooped by
mhryu@live.com
November 11, 4:26 PM
|
Scientists are editing microbes to supply more nitrogen to staple crops.
|
|
Scooped by
mhryu@live.com
November 11, 3:17 PM
|
Precise regulation of CRISPR-Cas9 activity is critical to limit off-target effects and improve the safety of genome engineering. To achieve reversible and robust control of Cas9, we developed multiple small-molecule-responsive systems that modulate the stability of the anti-CRISPR protein AcrIIA4. By fusing AcrIIA4 to dTAG, SMASh, or ecDHFR degrons, we enabled ligand-dependent degradation or stabilization of the inhibitor, thereby tightly regulating CRISPR cutting, activation, and interference in human cells. To further eliminate background Cas9 activity, we engineered a theophylline-responsive riboswitch into the sgRNA scaffold to block Cas9-sgRNA assembly until ligand addition. Combining riboswitch sgRNAs with degron-controlled AcrIIA4 produced the Cas9-ACROBAT platform, which provided near-complete suppression of CRISPR leakiness, while maintaining potent induction of genome editing and transcriptional modulation, including regulation of endogenous targets such as PLK1, CDKN1A, and CCND1. Cas9-ACROBAT offers a versatile, modular strategy for precise, small-molecule control of CRISPR systems.
|
|
Scooped by
mhryu@live.com
November 11, 2:09 PM
|
Although antimicrobial resistance (AMR) is traditionally viewed as a consequence of antibiotic selection, growing evidence indicates that bacterial immune systems regulate which mobile genetic elements (MGEs) successfully enter, establish, and persist within bacterial populations. Here, we argue that bacterial immunity helps determine access to the mobile resistome and, consequently, shapes the evolutionary opportunities upon which antibiotic selection acts. This review synthesizes current evidence on the interplay between bacterial immunity, counter-defense mechanisms, and MGE-MGE conflict, highlighting how these interactions collectively influence the entry, survival, expression, persistence, and transmission of foreign DNA. We further discuss recent discoveries revealing a far greater diversity of bacterial defence and MGE counter-defence strategies than previously recognized, including conflicts extending beyond genome protection to the regulation of gene expression, and consider how this expanding molecular repertoire influences MGE accessibility and the mobile genetic variation available for AMR evolution. By integrating concepts that are typically discussed separately, we propose that immunity, counter-defense, and MGE competition collectively regulate access to the vast and largely uncatalogued mobile gene pool from which resistance determinants are acquired. Viewed through this framework, AMR emerges not only from antibiotic selection, but also from the evolutionary processes that determine which genetic variants become available for selection, providing a broader perspective on resistance emergence and the uneven distribution of high-risk resistant lineages.
|
|
Scooped by
mhryu@live.com
November 11, 1:52 PM
|
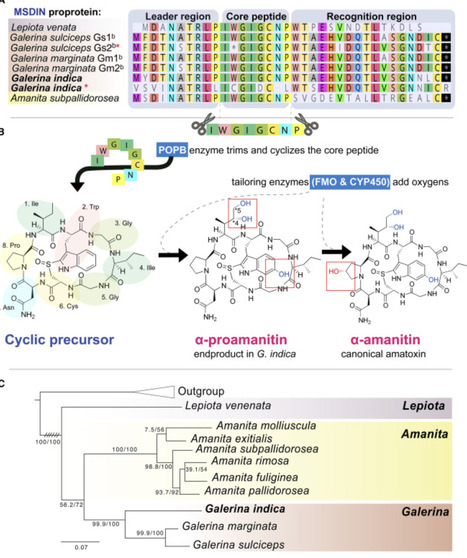
The most famous chemicals produced by mushrooms are the psychedelic compound psilocybin from “magic mushrooms” and amatoxins from deadly poisonous mushrooms. These compounds are known to occur in several distantly related groups of fungi but have never been shown to co-occur within a single species. Here, we show that the “dark magic mushroom” Galerina indica produces both psilocybin and amatoxins. Mass spectrometry revealed psilocybin and amatoxins in mushroom tissues, and genomic analyses identified corresponding biosynthetic genes. Phylogenetic analyses suggest that G. indica acquired psilocybin biosynthesis via horizontal gene transfer after amatoxin biosynthesis was already established. These findings reveal how horizontal gene transfer can combine powerful bioactive systems in a single species, potentially altering the ecological roles of both compound classes and the evolutionary fitness of the species.
|
|
Scooped by
mhryu@live.com
November 11, 1:47 PM
|
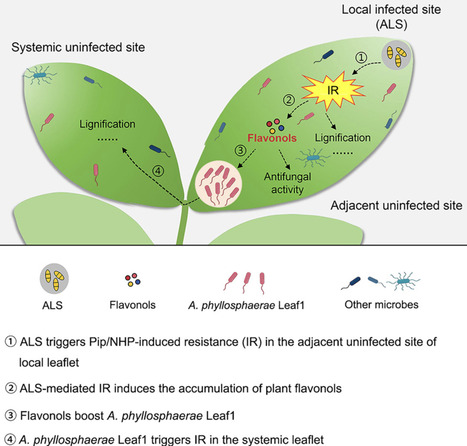
Pathogen-mediated induced resistance (IR) and beneficial microbe-mediated IR are key for plant survival during pathogen attack, yet how these two forms of IR interact remain unclear. Here, we demonstrate that foliar pathogen-triggered induced resistance (IR) orchestrates the assembly of a specific phyllosphere microbiota that can activate IR. Pathogen-triggered IR caused plant flavonol accumulation, which in turn enriched Aureimonas spp., especially the native A. phyllosphaerae Leaf1, in the phyllosphere endophytic microbiota near infection sites. Genetically impairing the capacity of Leaf1 to utilize flavonols reduced its proliferation during pathogen infection. Moreover, Leaf1 protected plants against leaf disease by activating IR and extended the protective range of pathogen-mediated IR. Importantly, pathogen-mediated IR-induced mobilization of native microbiota represented a common yet underrated component of plant resistance to foliar disease. These findings reveal a plant defense “relay” mechanism, whereby pathogen-mediated IR reassembles a specific microbiota that in turn activates IR to jointly combat further pathogen infection.
|
|
Scooped by
mhryu@live.com
November 11, 1:20 PM
|
Intrinsically disordered proteins (IDPs) are central to many biological processes, but their function depends on dynamic conformational ensembles rather than a single structure. Despite the impact of artificial intelligence on protein structure prediction, determining these ensembles at atomistic resolution remains challenging. Recently, we introduced bAIes, a framework that leverages the structural information encoded in AlphaFold2 distograms to generate conformational ensembles of IDPs at atomistic resolution. Here, we extend this framework to six state-of-the-art structure prediction tools (AlphaFold2, AlphaFold3, Boltz-1, Boltz-2, ESMFold, and ESMFold2) and systematically compare the structural information encoded in their predicted inter-residue distance distributions. Despite differences between predictors, we show that bAIes generates ensembles with comparable accuracy against experimental NMR and SAXS data, and competitive performance relative to state-of-the-art all-atom simulations and alternative IDP ensemble generation methods. These results demonstrate that bAIes is robust to the choice of structure predictor and that the distance distributions generated by different deep learning architectures contain valuable information about the conformational preferences of disordered proteins. More broadly, our work highlights the potential of combining AI-derived structural information with physics-based modeling to generate atomistic ensembles of intrinsically disordered proteins and regions.
|
|
Scooped by
mhryu@live.com
November 11, 1:10 PM
|
MobiDB (https://mobidb.org) integrates structural and functional annotations for intrinsically disordered proteins and regions across UniProtKB. The 2027 release strengthens semantic consistency, programmatic access, and reproducible reuse. A refined controlled vocabulary groups features in a hierarchical way and distinguishes evidence classes and types annotation sources as providers, methods, or aggregation rules. A new versioned REST API, along with a developed MCP server, enables scripts, workflows, and tool-using artificial intelligence systems to retrieve and combine evidence-bearing records through documented operations. The redesigned website applies the same vocabulary to protein entries, documentation, and cross-proteome statistics. Additional developments include standardized Nextflow production workflows, updated RING Protein Data Bank interaction processing, integration of Motif Map of the Proteome and Linear Motif mediated Protein–Protein Interaction Database annotations, and a release-aware catalogue of static datasets for large-scale analysis. Following the recent increase in reference proteomes in UniProtKB, MobiDB now provides a more representative estimate of long-disorder with a median residue coverage of 12.3% in eukaryotes, 1.9% in archaea, 1.7% in bacteria, and 3.4% in viruses; and protein prevalence of 48.7%, 11.9%, 11.2%, and 25.0%, respectively. Together, these developments make disorder knowledge easier to discover, relate to function, and reuse with explicit provenance.
|