Your new post is loading...

|
Scooped by
mhryu@live.com
November 11, 1:13 PM
|
RNAs are emerging as promising therapeutic targets, and small-molecule binding can modulate their structure and function, creating opportunities for therapeutic development. Accurate identification of RNA small-molecule binding sites is therefore important for understanding RNA-ligand interactions and facilitating RNA-targeted drug discovery. However, the structural complexity of RNA and the limited availability of experimentally characterized RNA-ligand complexes pose challenges for computational binding-site prediction. In this study, we propose RSMBind, a graph attention-based deep learning framework for nucleotide-level prediction of RNA-small molecule binding sites. The model integrates geometric descriptors of the local three-dimensional environment with pretrained RNA language model embeddings to capture complementary structural and sequence information. We investigate three pretrained RNA language models RNA-BERT, ERNIE-RNA and RNA-FM and assess the contributions of sequence representations and geometric features through feature-level ablation experiments. The model is developed using the TR60 and HARIBOSS datasets and evaluated on four independent benchmark datasets: RB9, TE18, TL12 and JL10. RSMBind achieves the highest MCC among the compared methods on three of the four benchmarks, with relative improvements of 17.1% on RB9, 7.2% on TL12 and 16.0% on JL10 over the best previously reported results, while achieving competitive performance on TE18. Furthermore, RSMBind retains predictive capability when using only pretrained RNA language model embeddings, highlighting their potential utility when explicit tertiary structural features are unavailable. Structural analysis indicates differences in prediction performance across RNA structural regions, with loop-associated binding sites generally being captured more effectively than those embedded within stem regions. Additionally, 100 ns molecular dynamics simulations of selected RNA-ligand complexes provide insights into the relationship between predicted binding probabilities and dynamic ligand proximity, further highlighting the role of conformational dynamics in evaluating structure-based predictions. The source code is available at [ https://github.com/sharma-nandita/RSMBind].
|
|
Scooped by
mhryu@live.com
November 11, 1:06 PM
|
A key parameter in the experimental design of RNA-seq projects is the choice of sequencing depth. Considering a limited budget, one needs to find a trade-off between the number of samples and the sensitivity of the analysis, particularly concerning lowly expressed genes. While previous studies have proposed a lower bound for the comprehensive analysis of differential gene expression, for the analysis of alternative splicing, it has only been proposed for human adipose tissue. However, alternative splicing differs across tissues and conditions. We analyzed publicly available and newly generated deep-sequenced paired-end RNA-seq samples (between 150 and >500 million reads, read length 50–150 bp) from human buffy coat cells and diverse sets of tissues, including gluteal subcutaneous fat, heart, and hypothalamus. Our results show that the sequencing depth typically used in published cohorts is not sufficient to comprehensively capture the landscape of alternative splicing. This motivates the use of deeper sequencing or long-read technologies in future studies. Toward this goal, we offer guidelines and a website (https://sequencing-depth4splicing.com/) for choosing a suitable sequencing depth.
|
|
Scooped by
mhryu@live.com
November 11, 11:40 AM
|
Agricultural biotechnology once found itself in opposition to environmental science. Now it could reduce the impact of food systems on global warming and biodiversity loss.
|
|
Scooped by
mhryu@live.com
November 11, 9:36 AM
|
Microbial fermentation won’t decarbonize the chemical industry and can’t compete with petroleum on cost or scale. Its enduring value lies in the complex molecules petrochemistry cannot make at all.
biochemicals that the US Department of Energy proposed to be well suited for bio-based production in reports from 2004 and 2010.
|
|
Scooped by
mhryu@live.com
November 11, 1:19 AM
|
Agriculture is both humanity’s largest infrastructure and the leading driver of environmental degradation. Many early-stage biotechnologies could mitigate this impact, but their deployability has not been systematically evaluated. We analyze deployability of 77 biotechnologies across eight domains (crops, soils, livestock, watersheds, aquaculture, health and security, food loss/waste and frontiers) by applying a Balanced Readiness Level assessment spanning technological, market, regulatory, acceptance and organizational dimensions. We find that market viability and social acceptance, not technological readiness, represent the primary constraints. High-readiness technologies tend to be ‘drop-in’, contained, transient and perceived as natural. These characteristics indicate strategies to elevate overall deployability, such as addressing genetically modified organism containment. Assessing the scalability and environmental impacts of biotechnologies for prioritization requires additional quantitative analysis, as shown in a Fermi analysis of microbial fertilizers. We combine our findings into a near- to long-term roadmap for living infrastructure: predictively designed multicomponent biological communities that function as systems at the ecosystem level to enhance food security and environmental resilience. Seventy-seven biotechnologies that could advance agricultural sustainability are analyzed for deployment readiness.
|
|
Scooped by
mhryu@live.com
November 11, 1:07 AM
|
DNA-targeting ligands comprise a diverse class of molecules that bind either canonical duplex DNA or non-canonical DNA structures such as G-quadruplexes. The recent shift towards combining distinct omics approaches, including epigenomics, transcriptomics, proteomics, and metabolomics, is significantly expanding the understanding of the biological effects induced by these small molecules, capturing complex cellular perturbations across multiple molecular layers. Here, we report representative examples from recent literature that illustrate how multi-omics approaches have helped to uncover previously unrecognised mechanisms underlying therapeutic efficacy, chemoresistance, toxicity, and chromatin remodelling induced by DNA-binding compounds. Finally, we address key computational bottlenecks in multi-omics data integration and highlight emerging in situ target-engagement mapping strategies that could bridge direct drug occupancy at genomic sites with downstream biological outcomes.
|
|
Scooped by
mhryu@live.com
November 11, 12:54 AM
|
Accurate and efficient EC number prediction is essential for elucidating the catalytic roles of enzymes. Although similarity-based and deep learning-based approaches have advanced this task, they often fail to distinguish active enzymes from closely related decoys inactivated by local functional-site mutations, leading to high false-positive rates (FPRs). We propose DecoyOpt, a two-stage pipeline for decoy-aware enzyme discrimination and EC number prediction. Specifically, we generated putatively inactive functional-site decoys using ESM3 and evaluated 16 protein language models across three splits. An MLP trained with classification and ranking losses distinguishes enzymes from unannotated proteins and decoys before a contrastive learning model assigns EC numbers to predicted enzymes. On the most stringent GraphPart 40% split, DecoyOpt achieves an MCC of 0.899 and a D0 decoy FPR of 6.17%, outperforming retrained baselines. Across all splits, it outperforms these baselines in EC number prediction on the full test set while maintaining performance close to CLEAN on the enzyme subset. It also correctly annotates all four wild-type enzymes and rejects all eight inactive or nearly inactive variants. We believe that DecoyOpt has potential for annotating uncharacterized proteins and their variants, with applications in genomics, synthetic biology, and biocatalysis.
|
|
Scooped by
mhryu@live.com
November 11, 12:48 AM
|
Although bacterial extracellular vesicles (BEVs) are increasingly recognized as mediators of communication, the contribution of their nucleic acid cargo remains poorly understood. Providing an overview of BEV-nucleic acid content and their roles, this review first examines their true significance. In particular, it highlights why they should not be treated as bulk cargo, given that DNA and RNA contents exhibit various sub-vesicular localizations that may define their ecological roles. We propose that this heterogeneity allows viewing BEVs as distinct subpopulations. Furthermore, recent studies increasingly propose the involvement of DNA associated with BEVs in horizontal gene transfer between bacteria and that of vesicular small RNA (sRNA) during host-bacteria interactions. Overall, this review offers insights into considering BEVs in an ecological context based on notable recent studies. omv
|
|
Scooped by
mhryu@live.com
November 11, 12:44 AM
|
Vibrio species adapt to different niches by sensing and responding to environmental signals such as nutrients, host cues, and quorum sensing autoinducers. Vibrio campbellii uses the master quorum sensing transcription factor LuxR to control the expression of hundreds of genes at high cell density. Furthermore, many gamma-proteobacteria use the global transcription factor catabolite activator protein (CAP) to control numerous physiologically relevant pathways, many of which are also modulated by quorum sensing, such as competence, biofilm formation, and carbon metabolism. However, the extent to which these two global transcription factors overlap to co-regulate gene expression and bacterial behaviors is understudied. In this work, we used chromatin immunoprecipitation coupled with high-throughput Illumina sequencing (ChIP-seq) and RNA-seq to determine the individual and combined regulons of CAP and LuxR in V. campbellii. We found that CAP and LuxR co-occupied 11 promoters to synergistically or antagonistically co-regulate genes involved with metabolism, respiration, and virulence. It was previously proposed that CAP and LuxR both bound the bioluminescence (luxCDABE) promoter to co-regulate these genes, and our RNA-seq data showed that these were indeed the most strongly co-regulated genes. However, our ChIP-seq data revealed that only LuxR bound the luxCDABE promoter in vivo. This pattern of co-regulation—where both CAP and LuxR strongly impact transcription, but only LuxR binds the promoter—was the most common mechanism observed. Our model for bioluminescence regulation is that CAP indirectly activates luxCDABE expression through the regulation of an intermediate factor. This study established new connections between the global gene regulatory networks underpinning nutrient sensing and population sensing.
|
|
Scooped by
mhryu@live.com
November 10, 4:46 PM
|
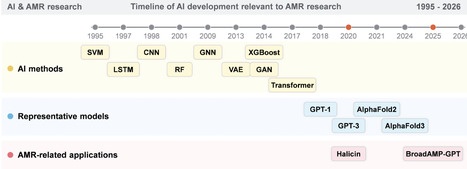
How artificial intelligence (AI) can be rationally applied to antibacterial drug discovery and development to overcome antimicrobial resistance (AMR) remains insufficiently defined, particularly across different resistance mechanisms and translational stages. The global emergence and dissemination of AMR are compromising conventional antibacterial therapies, while slow antibiotic discovery, limited target diversity, and reliance on single-mechanism drug design further increase the risk of inadequate treatment options. With strengths in multi-omics integration, complex pattern recognition, and structure-guided molecular design, AI offers a promising approach to support next-generation antibacterial development. This review focuses on AI-related strategies for antimicrobial development against bacterial resistance and provides a systematic overview of advances in target identification, molecular design, activity prediction, and clinical translation. Rather than proposing a new theoretical framework, the review introduces a pragmatic synthesis tool for evaluating where AI adds value and where important limitations remain, considering application objectives, data properties, validation depth, biological interpretability, and translational feasibility. Major challenges are highlighted, including data heterogeneity, clonal bias, limited external validation, discrepancies between in vitro activity and in vivo therapeutic efficacy, and the absence of regulatory approval for any AI-discovered or AI-designed antibacterial agent. Correspondingly, future directions are delineated, including multimodal data integration, explainable AI methodologies, validation through synthetic biology, and coordinated international regulatory efforts. Overall, this review provides a structured overview of how AI can be integrated with rational antibacterial design while considering both its potential and current limitations for addressing multidrug-resistant pathogens.
|
|
Scooped by
mhryu@live.com
November 10, 4:36 PM
|
Excess cholesterol accumulates in colorectal cancer (CRC) and promotes its progression, yet systemic cholesterol-lowering has limited benefits and safety concerns. Here we show that redirecting intestinal cholesterol metabolism through the gut microbiota generates a metabolite, 4-cholesten-3-one (4-C-3), which suppresses CRC growth. A screen of microbial cholesterol metabolites showed that 4-C-3 is depleted in faecal and tumor samples from patients with CRC. 4-C-3 suppresses tumour growth in CRC cell lines, patient-derived organoids and xenografts. Mechanistically, 4-C-3 suppresses two major oncogenic drivers of CRC by inhibiting epidermal growth-factor receptor activation and stabilizing mutant oncogenic KRAS in an inactive state, thereby overcoming KRAS-driven resistance to anti-epidermal growth-factor receptor therapy. As proof of concept, restoring 4-C-3 using either Oscillibacter ruminantium, a natural 4-C-3 producer reduced in CRC, or a non-pathogenic Escherichia coli engineered to convert intestinal cholesterol into 4-C-3, lowered intratumoral cholesterol and induced tumour regression in preclinical models without overt toxicity. These findings uncover a microbiota–cholesterol metabolic pathway that can be harnessed through microbiome engineering to selectively target CRC and overcome therapy resistance. Modulation of intratumoral cholesterol metabolism through delivery of a microbiota-derived metabolite is shown to induce colorectal cancer regression in mice.
|
|
Scooped by
mhryu@live.com
November 10, 3:55 PM
|
Bioprinting has moved from building organ-like shapes that keep cells alive to constructs judged by function. However, function only emerges when computational design, printing hardware and instructive bioinks are coupled in one process. Making this coupling routine and reproducible is essential for translating printed tissues into credible therapies and models.
|
|
Scooped by
mhryu@live.com
November 10, 3:44 PM
|
Cellular agriculture promises to reduce the environmental burden and inefficiencies of food systems. The question is no longer whether precision fermentation and cultivated meat production can produce alternative proteins and other food components, but whether they can attain the cost, scale and quality required for broad adoption. We examine cellular agriculture through the interrelated lenses of enabling technologies, food function and manufacturing economics. The field is approaching an inflection point where biological technology development is less important for commercial success than achieving scalable, affordable manufacturing. Key challenges include how products and their functional attributes are designed and how production processes are configured. For precision fermentation, the most compelling opportunities lie in functional approaches to improve taste, nutritional quality, health benefits and formulation rather than in direct imitation of existing foods or ingredients. For cultivated meat production, impact will come first from hybrid and ingredient-based products that capture specific values of cellular ingredients while avoiding the costs of full tissue mimicry. Making protein using cells, not animals and plants, promises sustainability and food quality benefits.
|
|
|
Scooped by
mhryu@live.com
November 11, 1:10 PM
|
MobiDB (https://mobidb.org) integrates structural and functional annotations for intrinsically disordered proteins and regions across UniProtKB. The 2027 release strengthens semantic consistency, programmatic access, and reproducible reuse. A refined controlled vocabulary groups features in a hierarchical way and distinguishes evidence classes and types annotation sources as providers, methods, or aggregation rules. A new versioned REST API, along with a developed MCP server, enables scripts, workflows, and tool-using artificial intelligence systems to retrieve and combine evidence-bearing records through documented operations. The redesigned website applies the same vocabulary to protein entries, documentation, and cross-proteome statistics. Additional developments include standardized Nextflow production workflows, updated RING Protein Data Bank interaction processing, integration of Motif Map of the Proteome and Linear Motif mediated Protein–Protein Interaction Database annotations, and a release-aware catalogue of static datasets for large-scale analysis. Following the recent increase in reference proteomes in UniProtKB, MobiDB now provides a more representative estimate of long-disorder with a median residue coverage of 12.3% in eukaryotes, 1.9% in archaea, 1.7% in bacteria, and 3.4% in viruses; and protein prevalence of 48.7%, 11.9%, 11.2%, and 25.0%, respectively. Together, these developments make disorder knowledge easier to discover, relate to function, and reuse with explicit provenance.
|
|
Scooped by
mhryu@live.com
November 11, 1:03 PM
|
Proximity labeling has emerged as a valuable tool for the interrogation of protein–protein and protein–RNA interactions. Almost all proximity labeling methods are deployed using large fusion proteins, which, while powerful in many contexts, can disrupt native biology or obscure important epitopes on the protein under study. Here we develop a photoproximity labeling method that is localized to a short 6-aa peptide sequence. Termed FlAsH-ID, this method uses the combination of the Tsien tetracysteine tag with the FlAsH dye for localized singlet oxygen generation. We demonstrate that this green-light-based method is nontoxic and can be used to label proteins within several cellular compartments. Lastly, we establish that FlAsH-ID can be deployed in primary CD8+ T cells to assess the interactome of the transcription factor RUNX3. Our results suggest this technique will expand the field of proximity labeling to small, functionally sensitive proteins in primary cells without perturbing their function. Commonly used proximity labeling methods implement large fusions of a protein of interest to a proximity labeling enzyme, which can block key protein interactions and disrupt native biology. Now, a method called FlAsH-ID has been developed that enables photoproximity labeling of a protein of interest using a 6-aa peptide tag to bind a small-molecule photocatalyst.
|
|
Scooped by
mhryu@live.com
November 11, 10:31 AM
|
Proteins operate through ligand and solvent interactions governed by thermodynamics, yet the enthalpy–entropy trade-offs that guide functional evolution remain unclear. Although the evolution of ligand specificity has been well studied, the thermodynamic determinants that underlie changes in specificity are not well understood. Here we reveal a shift from entropy-driven binding in the most distant ancestor to the LacI/GalR family (LGF) of transcription factors, to enthalpy-driven binding in the most recent ancestor and extant LacI. The most distant ancestor is characterized by the ability to bind its ligand in an open and dynamic conformation. We propose that entropically driven binding is allowed by the presence of entropic reservoirs. This thermodynamic binding trade-off between the most distant and most recent ancestor is in accordance with the concept that ancient life existed in a hot environment, where higher temperatures enhanced entropically driven binding. This suggests that molecular binding mechanisms evolved not just for ligand specificity but to adapt to environmental pressures such as cooling temperatures where enthalpic binding modes are favored. Transcription factors of the LacI/GalR family have been diverging for over 2 billion years. Ancestral reconstruction, isothermal titration calorimetry and structural analysis have now been used to show that ligand binding switched from entropy-driven, in an ancient repressor that binds ligand in an open and disordered state, to enthalpy-driven and ordered in Escherichia coli LacI.
|
|
Scooped by
mhryu@live.com
November 11, 1:29 AM
|
Diffusion models (DMs) have emerged as a class of generative models for controllable small-molecule design in drug discovery capable of progressively transforming noisy or partially specified molecular states into candidate structures under structural, geometric, and property constraints. This Review provides a comprehensive overview of the methodological foundations and representative applications of diffusion-based molecular generation, emphasizing the alignment among molecular representations, generative dynamics, controllable design, and evaluation and validation. Particular attention is given to recent advances in property- and structure-guided generation, target-aware and fragment-based design, multiobjective optimization, and synthesis-aware generation. We also discuss the roles of data sets, computational evaluation, and prospective experiments in assessing the chemical and biological relevance of generated molecules. By organizing these developments within a unified conceptual framework, we provide computational chemists and drug discovery researchers with a structured roadmap that balances the mathematical rigor with chemical intuition. Throughout, we highlight current methodological limitations and unresolved challenges and conclude with future directions toward chemically credible and experimentally grounded molecular design. To facilitate navigation of this rapidly evolving field, we introduce an interactive companion platform (https://diffsmd.scbdd.com) that provides a curated, continuously updated repository of model architectures and visualizations.
|
|
Scooped by
mhryu@live.com
November 11, 1:11 AM
|
Microbial communities shape immunity through antigens, structural ligands, and metabolites, but how these transient signals are translated into stable, reversible, or disease-promoting immune cell states remains incompletely understood. This review examines epigenetic and epitranscriptomic regulation as interconnected layers of host-microbiota interactions. We discuss how microbial signals engage chromatin- and RNA-regulatory pathways across epithelial, myeloid, lymphoid, and stromal compartments, placing the microbiota-N6-methyladenosine (m6A) axis within the broader landscape of host and microbial RNA modifications. This framework helps explain how the same microbial molecule can promote barrier repair, immune tolerance, inflammation, or antitumor immunity depending on tissue exposure, responding cell type, and host state. We distinguish between association, microbiota dependence, and direct regulatory causality, and highlight knowledge gaps concerning non-m6A RNA modifications and bidirectional regulatory mechanisms. Finally, we outline key experimental and translational considerations, including microbial function, host target engagement, cell specificity, spatial delivery, and disease context. These insights highlight chromatin and RNA regulation as potential mechanistic links between microbial ecology and immune function that welcome further experimental investigation.
|
|
Scooped by
mhryu@live.com
November 11, 1:04 AM
|
Complex microbial communities are found throughout terrestrial, aquatic, and animal-associated ecosystems. Genetic modification of discrete microbial taxa within native communities represents a powerful tool to study and augment microbiome functions. Herein, we develop Synthetic Phage Assisted Reprogrammable and Targetable Addition (SPARTA) as a one-step strategy for packaging, delivering, and integrating large synthetic DNA into a broad range of native microbiomes. SPARTA leverages a broad-spectrum synthetic phage particle for in situ genome engineering within complex soil, freshwater, and mammalian fecal microbiomes, reprogramming 59 native species across 16 families. By modulating SPARTA components, we further demonstrated tunability of SPARTA towards targeted microbiome population. Additionally, we built a novel companion tool to quantitatively identify engineered taxa, named 16S crRNA-guided inference of bacterial engineering sequencing or ScRibe-seq. The SPARTA-ScRibe-seq pipeline delivered and validated functional synthetic gene clusters in diverse native microbiomes while preserving the original microbial community structure. Together, this novel genome engineering framework constitutes an end-to-end programmable platform for rational and customizable reprogramming of native microbiomes.
|
|
Scooped by
mhryu@live.com
November 11, 12:51 AM
|
Since its discovery, CRISPR has revolutionized biomedical science and engineering with its ability to recognize and edit gene sequences with high specificity and precision. Despite the growing interest in using these capabilities to address environmental and sustainability challenges, the broader potential of CRISPR in environmental systems remains unexplored relative to its biomedical applications. Here, we provide a critical perspective on the emerging use of CRISPR for pollution management, ecological conservation, and environmental monitoring. Rather than viewing CRISPR solely as a gene-editing or biosensing technology, we consider both capabilities together to examine how advances in CRISPR engineering can translate into practical environmental solutions. We highlight the fundamental principles behind CRISPR-based editing and sensing and evaluate the challenges that constrain their transition from proof-of-concept experiments to field deployment, including off-target effects, delivery methods, ecological risks, and low sensitivity. Addressing these hurdles requires a multifaceted approach that integrates emerging technologies such as reversible CRISPR technology, amplification and cascade reactions, and machine learning, with strict regulatory oversight and rigorous risk assessments. This perspective identifies critical strengths, weaknesses, and research priorities for advancing CRISPR from laboratory technology toward robust tools for environmental applications.
|
|
Scooped by
mhryu@live.com
November 11, 12:46 AM
|
Urinary tract infections (UTIs) are among the most prevalent bacterial infections worldwide, with ~50% of women experiencing at least one UTI in their lifetime. Uropathogenic E. coli (UPEC) is the causative agent of ~80% of UTIs. In contrast, asymptomatic bacteriuria E. coli 83972 (ABU 83972) colonizes the human urinary bladder without causing symptoms and inflammation. Deliberate colonization with ABU 83972 reduced UTI incidence in patients with voiding disorders. However, the mechanisms by which ABU 83972 outcompetes uropathogens in the urinary tract remain poorly understood and are the focus of this report. We performed comparative metabolomics to identify metabolites that promoted the fitness of ABU 83972 in human urine compared to UPEC strain CFT073. Competition assays were conducted between ABU 83972 and UPEC CFT073 in human urine supplemented with select metabolites that were differentially abundant in metabolome profiling. There was a significant reduction in the fitness of ABU 83972 when supplemented with branched-chain amino acids (BCAAs) such as leucine and isoleucine in a dose-dependent manner. Further, we identified that BrnQ, a low-affinity BCAA importer, is the main determinant of fitness advantage of ABU 83972 over UPEC. Using isogenic mutants, genetic complementation, amino acid analysis, cell culture, chemical inhibition of BCAA biosynthesis, and mouse models of UTI, we demonstrate that BCAA homeostasis represents a unique nutritional vulnerability in UPEC that could be targeted with chemical inhibitors. In summary, our study highlights that intraspecies competition between commensal and pathogenic strains could be used to detect novel fitness mechanisms and metabolic bottlenecks.
|
|
Scooped by
mhryu@live.com
November 11, 12:23 AM
|
Many regulatory mechanisms function by changing a molecule’s conformation such that it is switched between active and inactive states. Engineering macromolecules such as proteins to show robust conformational changes in response to arbitrary, well-defined molecular stimuli is of great interest in fields such as synthetic biology and nanoscale engineering, because it allows the development of “smart” molecular tools. We present a strategy that can convert “simple” affinity binders such as nanobodies into protein switches that undergo clear structural rearrangements in response to their innate targets, by introducing a peptide that competes for the binding site. We demonstrate proof-of-concept implementations of this “NanoBlock” design by developing genetically encoded fluorescent biosensors based on two entirely different binder frameworks, the ALFA nanobody and the PDZ domain of Erbin. We show that our design allows the construction of well-performing protein switches that can reliably detect their targets in vitro, in mammalian cells, as part of fluorescence-activated cell sorting (FACS) experiments, and that also generalizes to the creation of stimulus-responsive enzymes via the integration of a bioluminescent readout. We furthermore develop a theoretical model that rationalizes the observations and provides insight into the underlying molecular mechanisms. Overall, our strategy expands the possibilities to create “smart” protein-based molecules and effectors.
|
|
Scooped by
mhryu@live.com
November 10, 4:44 PM
|
The Gram-positive bacterium Bacillus subtilis is a rapidly growing and easily manipulated microbe with a long history of exploitation for the commercial production of industrial enzymes, high value biochemicals, antibiotics and other secondary metabolites. More recently, it has been developed as a food additive and plant probiotic. It grows on inexpensive substrates and remains productive during extended batch-fed growth conditions. Extensive knowledge of its genetics, biochemistry and gene regulation has facilitated the use of metabolic engineering strategies to optimise substrate utilisation and product yield. As a free-living environmental organism, B. subtilis differentiates into subpopulations with distinct biological functions (e.g. sporulation, biofilm formation, production of antimicrobials, etc.). However, during fermentation, cellular differentiation processes can pose a challenge for its optimal biotechnological utilization, particularly when emerging subpopulations do not contribute to product biosynthesis. Here, we present robust assays that facilitate the analysis of two previously difficult-to-study population properties of B. subtilis: (i) the energisation levels of individual cells within post-exponential but actively growing cultures and (ii) the extent of cell lysis that can occur under such conditions. Our findings reveal an unappreciated level of heterogeneity in cell energisation within post-exponential cultures, and a surprisingly high degree of cell lysis in seemingly healthy, actively growing populations. These data provide insights and add to our understanding of the biological complexities and single-cell heterogeneities present in superficially simple bacterial clonal cultures. They establish robust and well-validated analytical tools with which to study the associated processes and provide a foundation for further optimising B. subtilis as an industrial production host. Considerable research efforts have focused on increasing the productivity of B. subtilis for industrial, medical and agricultural products. However, its ability to undergo physiological and morphological differentiation processes at high cell densities could ultimately limit its productivity. Our research reveals how the resulting heterogeneity impacts the population-level energy status of individual cells in the culture and the surprisingly high extent of population-level cell lysis. The ability to monitor these processes provides tools for evaluating the impact of genetic and metabolic engineering strategies to improve productivity.
|
|
Scooped by
mhryu@live.com
November 10, 4:11 PM
|
Thanks to breakthroughs in genetics, we know how to slash the methane emissions from rice farming. It is time for governments and the private sector to translate the science into commercial rice seeds.
|
|
Scooped by
mhryu@live.com
November 10, 3:55 PM
|
Chemical nitrogen fertilizers produce substantial greenhouse gas emissions. Emerging approaches for developing nitrogen-fixing bacteria could provide a sustainable alternative.
|