 Your new post is loading...

|
Scooped by
mhryu@live.com
April 13, 2018 12:58 PM
|
Diazotrophic Bradyrhizobium spp. are well known for their ability to trigger nodule formation on a variety of legume species. In nodules, Bradyrhizobium utilizes plant-derived carbohydrates in exchange for fixed nitrogen. The genes essential for the nodulation and nitrogen-fixation trait are clustered in a genomic region, which is known as the ‘symbiotic island’. Recently, novel non-diazotrophic Bradyrhizobium spp. have been found to be highly abundant in soils, suggesting that these species can also have a ‘free-living’ life history. However, whether non-diazotrophic Bradyrhizobium spp. can live in association with plants remains elusive. In this study, we show that Bradyrhizobium spp. are common root endophytes of non-legume plant species – including Arabidopsis thaliana (Arabidopsis) – grown in an ecological setting. From a single Arabidopsis root, four Bradyrhizobium sp. strains (designated MOS001 to MOS004) were isolated. Comparative genome analysis revealed that these strains were genetically and functionally highly diverse, but did not harbour the nodulation and the nitrogen fixation gene clusters. Comparative colonization experiments, with MOS strains and nitrogen-fixing symbiotic strains, revealed that all tested Bradyrhizobium spp. can colonize the root endophytic compartment of Arabidopsis. This study provides evidence that both diazotrophic and non-diazotrophic Bradyrhizobium spp. colonize the root endophytic compartment of a wide variety of plant species, including the model species Arabidopsis. This demonstrates that plant roots form a major ecological niche for Bradyrhizobium spp., which might be ancestral to the evolution of the nodulation and nitrogen-fixation trait in this genus.
|
|
Scooped by
mhryu@live.com
April 13, 2018 1:31 AM
|
Elimination or mitigation of the toxic effects of chemical waste released to the environment by industrial and urban activities relies largely on the catalytic activities of microorganisms—specifically bacteria. Given their capacity to evolve rapidly, they have the biochemical power to tackle a large number of molecules mobilized from their geological repositories through human action (e.g., hydrocarbons, heavy metals) or generated through chemical synthesis (e.g., xenobiotic compounds). Whereas naturally occurring microbes already have considerable ability to remove many environmental pollutants with no external intervention, the onset of genetic engineering in the 1980s allowed the possibility of rational design of bacteria to catabolize specific compounds, which could eventually be released into the environment as bioremediation agents. The complexity of this endeavour and the lack of fundamental knowledge nonetheless led to the virtual abandonment of such a recombinant DNA-based bioremediation only a decade later. In a twist of events, the last few years have witnessed the emergence of new systemic fields (including systems and synthetic biology, and metabolic engineering) that allow revisiting the same environmental pollution challenges through fresh and far more powerful approaches. The focus on contaminated sites and chemicals has been broadened by the phenomenal problems of anthropogenic emissions of greenhouse gases and the accumulation of plastic waste on a global scale. In this article, we analyze how contemporary systemic biology is helping to take the design of bioremediation agents back to the core of environmental biotechnology. We inspect a number of recent strategies for catabolic pathway construction and optimization and we bring them together by proposing an engineering workflow.
|
|
Scooped by
mhryu@live.com
April 12, 2018 1:09 PM
|
Our ability to generate bacterial strains with unique and increasingly complex functions has rapidly expanded in recent times. The capacity for DNA synthesis is increasing and costing less; new tools are being developed for fast, large-scale genetic manipulation; and more tested genetic parts are available for use, as is the knowledge of how to use them effectively. These advances promise to unlock an exciting array of 'smart' bacteria for clinical use but will also challenge scientists to better optimize preclinical testing regimes for early identification and validation of promising strains and strategies. Here, we review recent advances in the development and testing of engineered bacterial diagnostics and therapeutics. We highlight new technologies that will assist the development of more complex, robust and reliable engineered bacteria for future clinical applications, and we discuss approaches to more efficiently evaluate engineered strains throughout their preclinical development.
|
|
Scooped by
mhryu@live.com
April 11, 2018 12:37 AM
|
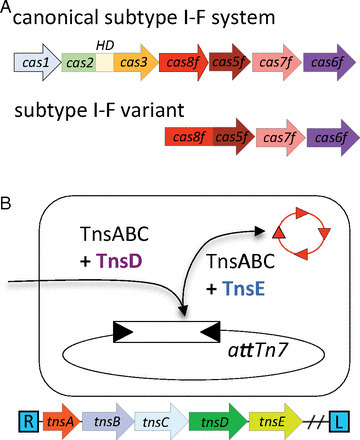
CRISPR-Cas is an adaptive immunity system that protects bacteria and archaea from mobile genetic elements. We present comparative genomic and phylogenetic analysis of minimal CRISPR-Cas variants associated with distinct families of transposable elements and develop the hypothesis that such repurposed defense systems contribute to the transposable element propagation by facilitating transposition into specific sites. Thus, these transposable elements are predicted to propagate via RNA-guided transposition, a mechanism that has not been previously described for DNA transposons.
|
|
Scooped by
mhryu@live.com
April 11, 2018 12:19 AM
|
The number of microbial genomes sequenced each year is expanding rapidly, in part due to genome-resolved metagenomic studies that routinely recover hundreds of draft-quality genomes. Rapid algorithms have been developed to comprehensively compare large genome sets, but they are not accurate with draft-quality genomes. Here we present dRep, a program that reduces the computational time for pairwise genome comparisons by sequentially applying a fast, inaccurate estimation of genome distance, and a slow, accurate measure of average nucleotide identity. dRep achieves a 28 × increase in speed with perfect recall and precision when benchmarked against previously developed algorithms. We demonstrate the use of dRep for genome recovery from time-series datasets. Each metagenome was assembled separately, and dRep was used to identify groups of essentially identical genomes and select the best genome from each replicate set. This resulted in recovery of significantly more and higher-quality genomes compared to the set recovered using co-assembly.
|
|
Scooped by
mhryu@live.com
April 10, 2018 11:52 PM
|
Dissimilatory perchlorate reduction is an anaerobic respiratory pathway that in communities might be influenced by metabolic interactions. Because the genes for perchlorate reduction are horizontally transferred, previous studies have been unable to identify uncultivated perchlorate-reducing populations. Here we recovered metagenome-assembled genomes from perchlorate-reducing sediment enrichments and employed a manual scaffolding approach to reconstruct gene clusters for perchlorate reduction found within mobile genetic elements. De novo assembly and binning of four enriched communities yielded 48 total draft genomes. In addition to canonical perchlorate reduction gene clusters and taxa, a new type of gene cluster with an alternative perchlorate reductase was identified. Phylogenetic analysis indicated past exchange between these gene clusters, and the presence of plasmids with either gene cluster shows that the potential for gene transfer via plasmid persisted throughout enrichment. However, a majority of genomes in each community lacked perchlorate reduction genes. Putative chlorate-reducing or sulfur-reducing populations were dominant in most communities, supporting the hypothesis that metabolic interactions might result from perchlorate reduction intermediates and byproducts. Other populations included a novel phylum-level lineage (Ca. Muirbacteria) and epibiotic prokaryotes with no known role in perchlorate reduction. These results reveal unexpected genetic diversity, suggest that perchlorate-reducing communities involve substantial metabolic interactions, and encourage expanded strategies to further understand the evolution and ecology of this metabolism.
|
|
Scooped by
mhryu@live.com
April 10, 2018 12:24 AM
|
Like all higher organisms, plants have evolved in the context of a microbial world, shaping both their evolution and their contemporary ecology. Interactions between plant roots and soil microorganisms are critical for plant fitness in natural environments. Given this co-evolution and the pivotal importance of plant–microbial interactions, it has been hypothesized, and a growing body of literature suggests, that plants may regulate the composition of their rhizosphere to promote the growth of microorganisms that improve plant fitness in a given ecosystem. Here, using a combination of comparative genomics and exometabolomics, we show that pre-programmed developmental processes in plants (Avena barbata) result in consistent patterns in the chemical composition of root exudates. This chemical succession in the rhizosphere interacts with microbial metabolite substrate preferences that are predictable from genome sequences. Specifically, we observed a preference by rhizosphere bacteria for consumption of aromatic organic acids exuded by plants (nicotinic, shikimic, salicylic, cinnamic and indole-3-acetic). The combination of these plant exudation traits and microbial substrate uptake traits interact to yield the patterns of microbial community assembly observed in the rhizosphere of an annual grass. This discovery provides a mechanistic underpinning for the process of rhizosphere microbial community assembly and provides an attractive direction for the manipulation of the rhizosphere microbiome for beneficial outcomes.
|
|
Scooped by
mhryu@live.com
April 10, 2018 12:08 AM
|
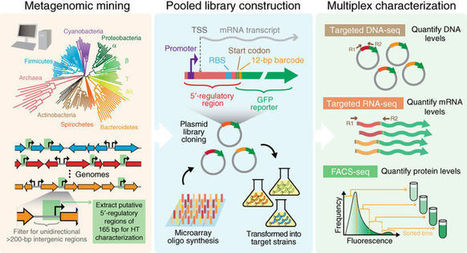
Robust and predictably performing synthetic circuits rely on the use of well-characterized regulatory parts across different genetic backgrounds and environmental contexts. Here we report the large-scale metagenomic mining of thousands of natural 5′ regulatory sequences from diverse bacteria, and their multiplexed gene expression characterization in industrially relevant microbes. We identified sequences with broad and host-specific expression properties that are robust in various growth conditions. We also observed substantial differences between species in terms of their capacity to utilize exogenous regulatory sequences. Finally, we demonstrate programmable species-selective gene expression that produces distinct and diverse output patterns in different microbes. Together, these findings provide a rich resource of characterized natural regulatory sequences and a framework that can be used to engineer synthetic gene circuits with unique and tunable cross-species functionality and properties, and also suggest the prospect of ultimately engineering complex behaviors at the community level.
|
|
Scooped by
mhryu@live.com
April 9, 2018 11:56 AM
|
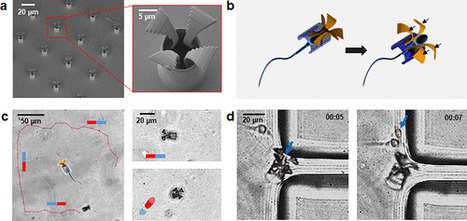
A sperm-driven micromotor is presented as a targeted drug delivery system, which is appealing to potentially treat diseases in the female reproductive tract. This system is demonstrated to be an efficient drug delivery vehicle by first loading a motile sperm cell with an anticancer drug (doxorubicin hydrochloride), guiding it magnetically, to an in vitro cultured tumor spheroid, and finally freeing the sperm cell to deliver the drug locally. The sperm release mechanism is designed to liberate the sperm when the biohybrid micromotor hits the tumor walls, allowing it to swim into the tumor and deliver the drug through the sperm–cancer cell membrane fusion. In our experiments, the sperm cells exhibited a high drug encapsulation capability and drug carrying stability, conveniently minimizing toxic side effects and unwanted drug accumulation in healthy tissues. Overall, sperm cells are excellent candidates to operate in physiological environments, as they neither express pathogenic proteins nor proliferate to form undesirable colonies, unlike other cells or microorganisms. This sperm-hybrid micromotor is a biocompatible platform with potential application in gynecological healthcare, treating or detecting cancer or other diseases in the female reproductive system.
|
|
Scooped by
mhryu@live.com
April 8, 2018 11:10 PM
|
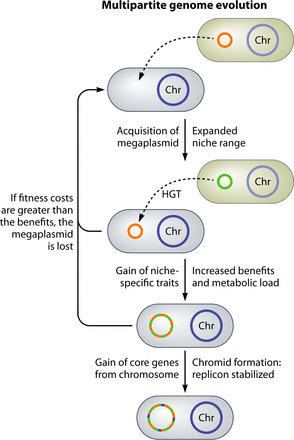
Approximately 10% of bacterial genomes are split between two or more large DNA fragments, a genome architecture referred to as a multipartite genome. This multipartite organization is found in many important organisms, including plant symbionts, such as the nitrogen-fixing rhizobia, and plant, animal, and human pathogens, including the genera Brucella, Vibrio, and Burkholderia. The availability of many complete bacterial genome sequences means that we can now examine on a broad scale the characteristics of the different types of DNA molecules in a genome. Recent work has begun to shed light on the unique properties of each class of replicon, the unique functional role of chromosomal and nonchromosomal DNA molecules, and how the exploitation of novel niches may have driven the evolution of the multipartite genome. The aims of this review are to (i) outline the literature regarding bacterial genomes that are divided into multiple fragments, (ii) provide a meta-analysis of completed bacterial genomes from 1,708 species as a way of reviewing the abundant information present in these genome sequences, and (iii) provide an encompassing model to explain the evolution and function of the multipartite genome structure. This review covers, among other topics, salient genome terminology; mechanisms of multipartite genome formation; the phylogenetic distribution of multipartite genomes; how each part of a genome differs with respect to genomic signatures, genetic variability, and gene functional annotation; how each DNA molecule may interact; as well as the costs and benefits of this genome structure.
|
|
Scooped by
mhryu@live.com
April 5, 2018 5:14 PM
|
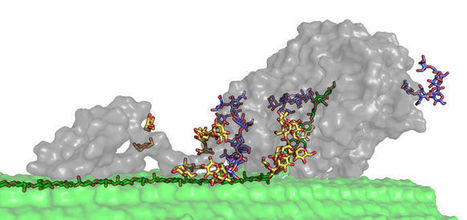
Glycoside Hydrolase Family 7 cellobiohydrolases (GH7 CBHs) catalyze cellulose depolymerization in cellulolytic eukaryotes, making them key discovery and engineering targets. However, there remains a lack of robust structure–activity relationships for these industrially important cellulases. Here, we compare CBHs from Trichoderma reesei (TrCel7A) and Penicillium funiculosum (PfCel7A), which exhibit a multi-modular architecture consisting of catalytic domain (CD), carbohydrate-binding module, and linker. We show that PfCel7A exhibits 60% greater performance on biomass than TrCel7A. To understand the contribution of each domain to this improvement, we measure enzymatic activity for a library of CBH chimeras with swapped subdomains, demonstrating that the enhancement is mainly caused by PfCel7A CD. We solve the crystal structure of PfCel7A CD and use this information to create a second library of TrCel7A CD mutants, identifying a TrCel7A double mutant with near-equivalent activity to wild-type PfCel7A. Overall, these results reveal CBH regions that enable targeted activity improvements.
|
|
Scooped by
mhryu@live.com
April 5, 2018 11:47 AM
|
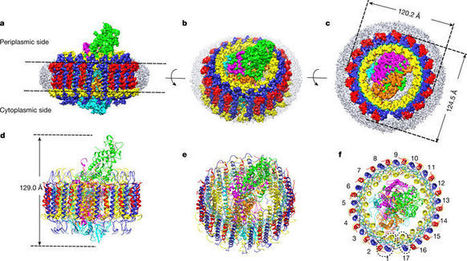
The light-harvesting 1–reaction centre (LH1–RC) complex is a key functional component of bacterial photosynthesis. Here we present a 2.9 Å resolution cryo-electron microscopy structure of the bacteriochlorophyll b-based LH1–RC complex from Blastochloris viridisthat reveals the structural basis for absorption of infrared light and the molecular mechanism of quinone migration across the LH1 complex. The triple-ring LH1 complex comprises a circular array of 17 β-polypeptides sandwiched between 17 α- and 16 γ-polypeptides. Tight packing of the γ-apoproteins between β-polypeptides collectively interlocks and stabilizes the LH1 structure; this, together with the short Mg–Mg distances of bacteriochlorophyll b pairs, contributes to the large redshift of bacteriochlorophyll b absorption. The ‘missing’ 17th γ-polypeptide creates a pore in the LH1 ring, and an adjacent binding pocket provides a folding template for a quinone, Q P, which adopts a compact, export-ready conformation before passage through the pore and eventual diffusion to the cytochrome bc1 complex.
|
|
Scooped by
mhryu@live.com
April 5, 2018 12:56 AM
|
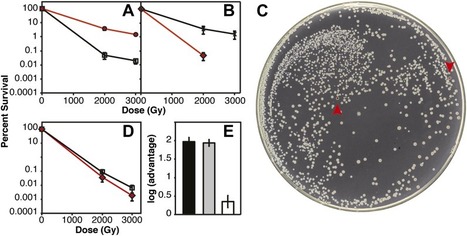
By directed evolution in the laboratory, we previously generated populations of Escherichia coli that exhibit a complex new phenotype, extreme resistance to ionizing radiation (IR). The molecular basis of this extremophile phenotype, involving strain isolates with a 3-4 order of magnitude increase in IR resistance at 3000 Gy, is now addressed. Of 69 mutations identified in one of our most highly adapted isolates, functional experiments demonstrate that the IR resistance phenotype is almost entirely accounted for by only three of these nucleotide changes, in the DNA metabolism genes recA, dnaB, and yfjK. Four additional genetic changes make small but measurable contributions. Whereas multiple contributions to IR resistance are evident in this study, our results highlight a particular adaptation mechanism not adequately considered in studies to date: Genetic innovations involving pre-existing DNA repair functions can play a predominant role in the acquisition of an IR resistance phenotype.
|
|
|
Scooped by
mhryu@live.com
April 13, 2018 11:38 AM
|
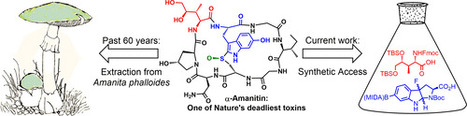
α-Amanitin is an extremely toxic bicyclic octapeptide isolated from the death-cap mushroom, Amanita phalloides. As a potent inhibitor of RNA polymerase II, α-amanitin is toxic to eukaryotic cells. Recent interest in α-amanitin arises from its promise as a payload for antibody–drug conjugates. For over 60 years, A. phalloides has been the only source of α-amanitin. Here we report a synthesis of α-amanitin, which surmounts the key challenges for installing the 6-hydroxy-tryptathionine sulfoxide bridge, enantioselective synthesis of (2S,3R,4R)-4,5-dihydroxy-isoleucine, and diastereoselective sulfoxidation.
|
|
Scooped by
mhryu@live.com
April 12, 2018 10:01 PM
|
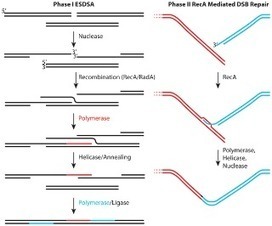
Most genome-editing nucleases introduce a double-strand break at a target site, leaving compatible DNA ends that are repaired by the nonhomologous end joining repair pathway to regenerate the target site. This leads to a cycle of cleavage, target site regeneration by ligation, and subsequent recleavage by the nuclease. Here, we bypass this cycle by developing and testing an RNA-guided dual active site nuclease that introduces two noncompatible DNA breaks at a target site, deleting the majority of the target site so that it cannot be regenerated. The dual nuclease (TevCas9) robustly functions in HEK293 and biases genome-editing outcomes toward defined length deletions at high frequencies. Potential applications include directional oligonucleotide ligation, deletion of DNA-binding sites, and in-frame protein deletions.
|
|
Scooped by
mhryu@live.com
April 11, 2018 9:28 PM
|
Processing of double‐stranded RNA precursors into small RNAs is an essential regulator of gene expression in plant development and stress response. Small RNA processing requires the combined activity of a functionally diverse group of molecular components. However, in most of the plant species, there are insufficient mutant resources to functionally characterize each encoding gene. Here, mutations in loci encoding protein machinery involved in small RNA processing in soya bean and Medicago truncatula were generated using the CRISPR/Cas9 and TAL‐effector nuclease (TALEN) mutagenesis platforms. An efficient CRISPR/Cas9 reagent was used to create a bi‐allelic double mutant for the two soya bean paralogous Double‐stranded RNA‐binding2 (GmDrb2a and GmDrb2b) genes. These mutations, along with a CRISPR/Cas9‐generated mutation of the M. truncatula Hua enhancer1 (MtHen1) gene, were determined to be germ‐line transmissible. Furthermore, TALENs were used to generate a mutation within the soya bean Dicer‐like2 gene. CRISPR/Cas9 mutagenesis of the soya bean Dicer‐like3 gene and the GmHen1a gene was observed in the T0 generation, but these mutations failed to transmit to the T1 generation. The irregular transmission of induced mutations and the corresponding transgenes was investigated by whole‐genome sequencing to reveal a spectrum of non‐germ‐line‐targeted mutations and multiple transgene insertion events. Finally, a suite of combinatorial mutant plants were generated by combining the previously reported Gmdcl1a, Gmdcl1b and Gmdcl4b mutants with the Gmdrb2ab double mutant. Altogether, this study demonstrates the synergistic use of different genome engineering platforms to generate a collection of useful mutant plant lines for future study of small RNA processing in legume crops.
|
|
Scooped by
mhryu@live.com
April 11, 2018 12:20 AM
|
Culture-independent microbiome studies have increased our understanding of the complexity and metabolic potential of microbial communities. However, to understand the contribution of individual microbiome members to community functions, it is important to determine which bacteria are actively replicating. We developed an algorithm, iRep, that uses draft-quality genome sequences and single time-point metagenome sequencing to infer microbial population replication rates. The algorithm calculates an index of replication (iRep) based on the sequencing coverage trend that results from bi-directional genome replication from a single origin of replication. We apply this method to show that microbial replication rates increase after antibiotic administration in human infants. We also show that uncultivated, groundwater-associated, Candidate Phyla Radiation bacteria only rarely replicate quickly in subsurface communities undergoing substantial changes in geochemistry. Our method can be applied to any genome-resolved microbiome study to track organism responses to varying conditions, identify actively growing populations and measure replication rates for use in modeling studies.
|
|
Scooped by
mhryu@live.com
April 10, 2018 11:58 PM
|
pSC101 is a narrow host range, low-copy plasmid commonly used for genetically manipulating Escherichia coli. As a byproduct of a genetic screen for a more sensitive lactam biosensor, we identified multiple novel mutations that increase the copy number of plasmids with the pSC101 origin. All mutations identified in this study occurred on plasmids which also contained at least one mutation localized to the RepA protein encoded within the origin. Homology modelling predicts that many of these mutations occur within the dimerization interface of RepA. Mutant RepA resulted in plasmid copy numbers between ~31 and ~113 copies/cell, relative to ~5 copies/cell in wild-type pSC101 plasmids. Combining the mutations that were predicted to disrupt multiple contacts on the dimerization interface resulted in copy numbers of ~500 copies/cell, while also attenuating growth in host strains. Fluorescent protein production expressed from an arabinose-inducible promoter on mutant origin derived plasmids did correlate with copy number. Plasmids harboring RepA with one of two mutations, E83K and N99D, resulted in fluorescent protein production similar to that from p15a- (~20 copies/cell) and ColE1- (~31 copies/cell) based plasmids, respectively. The mutant copy number variants retained compatibility with p15a, pBBR, and ColE1 origins of replication. These pSC101 variants may be useful in future metabolic engineering efforts that require medium or high-copy vectors compatible with p15a- and ColE1-based plasmids.
|
|
Scooped by
mhryu@live.com
April 10, 2018 12:25 AM
|
In two important reports in this issue of PLoS Genetics, Sommer and colleagues convert much recent speculation into substance. In the first report [6], the Chandler and Sommer laboratories collaborate to explore the mechanism of transposition of element ISDra2. This transposon is a member of a family of elements that transpose via single-stranded DNA intermediates. Transposition is activated by irradiation of Deinococcus. The work not only documents the transposition mechanism, it reinforces the proposition that extensive lengths of single-stranded genomic DNA are generated in the early stages of genome reconstitution in this bacterium. As a bonus, the work provides hope for the development of in vivo transposition as a tool for genetic manipulation of this genome.
|
|
Scooped by
mhryu@live.com
April 10, 2018 12:14 AM
|
mRNAs can fold into complex structures that regulate gene expression. Resolving such structures de novo has remained challenging and has limited our understanding of the prevalence and functions of mRNA structure. We use SHAPE-MaP experiments in living E. coli cells to derive quantitative, nucleotide-resolution structure models for 194 endogenous transcripts encompassing approximately 400 genes. Individual mRNAs have exceptionally diverse architectures, and most contain well-defined structures. Active translation destabilizes mRNA structure in cells. Nevertheless, mRNA structure remains similar between in-cell and cell-free environments, indicating broad potential for structure-mediated gene regulation. We find that the translation efficiency of endogenous genes is regulated by unfolding kinetics of structures overlapping the ribosome binding site. We discover conserved structured elements in 35% of UTRs, several of which we validate as novel protein binding motifs. RNA structure regulates every gene studied here in a meaningful way, implying that most functional structures remain to be discovered.
|
|
Scooped by
mhryu@live.com
April 9, 2018 10:20 PM
|
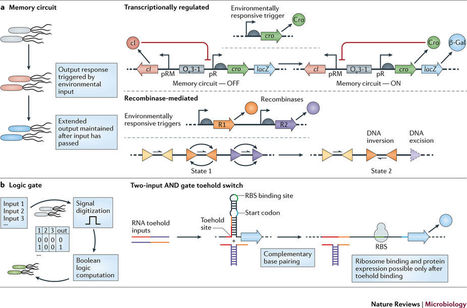
Model ecosystems could provide significant insight into the evolution and behavior of real ecosystems. We discuss the advantages and limitations of common approaches like mesocosms. In this context, we highlight recent breakthroughs that allow for the creation of networks of organisms with independently controlled environments and rates of chemical exchange.
|
|
Scooped by
mhryu@live.com
April 9, 2018 11:35 AM
|
The demand for cellulosic biofuels is on the rise because of the anticipation for sustainable energy and less greenhouse gas emissions in the future. However, production of cellulosic biofuels, especially cellulosic butanol, has been hampered by the lack of potent microbes that are capable of converting cellulosic biomass into biofuels. We report a wild-type Thermoanaerobacterium thermosaccharolyticum strain TG57, which is capable of using microcrystalline cellulose directly to produce butanol (1.93 g/liter) as the only final product (without any acetone or ethanol produced), comparable to that of engineered microbes thus far. Strain TG57 exhibits significant advances including unique genes responsible for a new butyrate synthesis pathway, no carbon catabolite repression, and the absence of genes responsible for acetone synthesis (which is observed as the main by-product in most Clostridium strains known today). Furthermore, the use of glucose analog 2-deoxyglucose posed a selection pressure to facilitate isolation of strain TG57 with deletion/silencing of carbon catabolite repressor genes—the ccr and xylR genes—and thus is able to simultaneously ferment glucose, xylose, and arabinose to produce butanol (7.33 g/liter) as the sole solvent. Combined analysis of genomic and transcriptomic data revealed unusual aspects of genome organization, numerous determinants for unique bioconversions, regulation of central metabolic pathways, and distinct transcriptomic profiles. This study provides a genome-level understanding of how cellulose is metabolized by T. thermosaccharolyticum and sheds light on the potential of competitive and sustainable biofuel production.
|
|
Scooped by
mhryu@live.com
April 7, 2018 10:10 PM
|
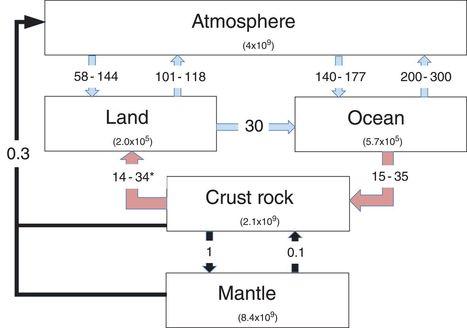
Nitrogen availability is a central controller of terrestrial plant growth and, thereby, of the carbon cycle and global climate change. It has been widely assumed that the atmosphere is the main source of terrestrial nitrogen input. Surprisingly, Houlton et al. now show that bedrock is just as large a nitrogen source across major sectors of the global terrestrial environment. They used three diverse and largely independent assessments of the nitrogen mobility and reactivity of rocks in the surface environment. These approaches yielded convergent estimates pointing to the equal importance of the atmosphere and bedrock as nitrogen sources.
|
|
Scooped by
mhryu@live.com
April 5, 2018 4:52 PM
|
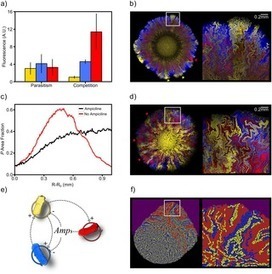
In order to achieve greater levels of complexity, complex systems often display cooperative interactions that enable the formation and stabilisation of mutualisms. Theoretical models have shown that closed chains of cooperative species or hypercycles might have been crucial in the evolution towards complexity in early molecular replicators. However, parasites can easily destroy the cooperative loop, unless the system is embedded in a spatial context where interactions are limited to nearest neighbours. A dynamically similar phenomenon occurs in ecological webs, where closed positive feedback loops contribute to global stability and ecophysiology. Here we explore this problem by engineering synthetic cooperative strains of microbes that grow and interact in a cell culture under the absence and presence of a synthetic parasitic strains. By analysing the impact of cooperation under different conditions, we find that cooperative replication is successful and overcomes competitive interactions in nutrient-poor environments. However, the same closed loop fails to establish in nutrient-rich media. Moreover, parasitic entities that jeopardise cooperation under well-mixed conditions can be overcome by hypercycles when growing in a two-dimensional space.
|
|
Scooped by
mhryu@live.com
April 5, 2018 9:28 AM
|
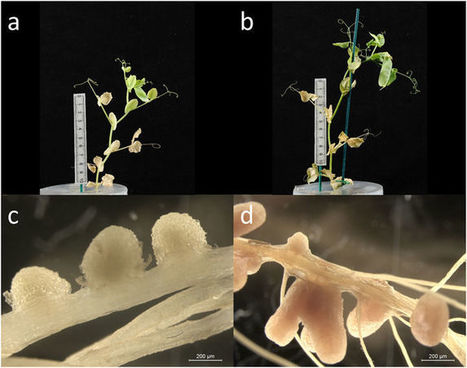
In legume-Rhizobium symbioses, specialised soil bacteria fix atmospheric nitrogen in return for carbon. However, ineffective strains can arise, making discrimination essential. Discrimination can occur via partner choice, where legumes prevent ineffective strains from entering, or via sanctioning, where plants provide fewer resources. Several studies have inferred that legumes exercise partner choice, but the rhizobia compared were not otherwise isogenic. To test when and how plants discriminate ineffective strains we developed sets of fixing and non-fixing strains that differed only in the expression of nifH – essential for nitrogen fixation – and could be visualised using marker genes. We show that the plant is unable to select against the non-fixing strain at the point of entry, but that non-fixing nodules are sanctioned. We also used the technique to characterise mixed nodules (containing both a fixing and a non-fixing strain), whose frequency could be predicted using a simple diffusion model. We discuss that sanctioning is likely to evolve in preference to partner choice in any symbiosis where partner quality cannot be adequately assessed until goods or services are actively exchanged.
|








interesting endophytes