Your new post is loading...

|
Scooped by
mhryu@live.com
Today, 11:50 AM
|
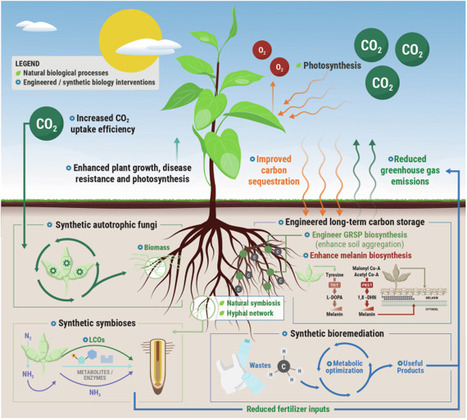
Over the past two decades, there has been a staggering increase in greenhouse gas emissions, particularly carbon dioxide (CO2) from unrestrained anthropogenic activities, leading to notable environmental shifts. Despite continuous efforts to reduce emissions, the world is not making satisfactory progress, which calls for immediate action to implement alternative strategies to enhance environmental sustainability. The Earth naturally absorbs and stores carbon through various processes, but merely keeping pace with ongoing human activities is no longer sufficient. Land plants and marine prokaryotes absorb CO2 during photosynthesis and store carbon as biomass. Eukaryotic organisms also represent a valuable resource that can provide insights into the biological fixation and removal of atmospheric CO2. Expanding the capacities of Earth’s natural systems offers enormous potential for enhancing carbon sinks and promoting environmental resilience. Although extensive synthetic engineering of plant systems has been conducted to enhance their native carbon fixation pathways, research on microorganisms, especially fungi, has been limited. In this review, we specifically explore the role of soil-borne fungi in carbon management, elucidating their responses to environmental shifts and their potential for climate change mitigation. We place particular emphasis on the role of various fungal guilds in this process, highlighting those that should be prioritized for future synthetic biology applications. We then engage with previous endeavors to address primary challenges in advancing genetic tools for these understudied fungi. Lastly, we discuss how integrating synthetic biology practices into the field of soil-associated fungi can drive advances in long-term carbon storage, enhance degradation of environmental pollutants, and improve nutrient exchange and soil health, contributing to more resilient soil ecosystems and sustainable environmental interventions.
|
|
Scooped by
mhryu@live.com
Today, 11:43 AM
|
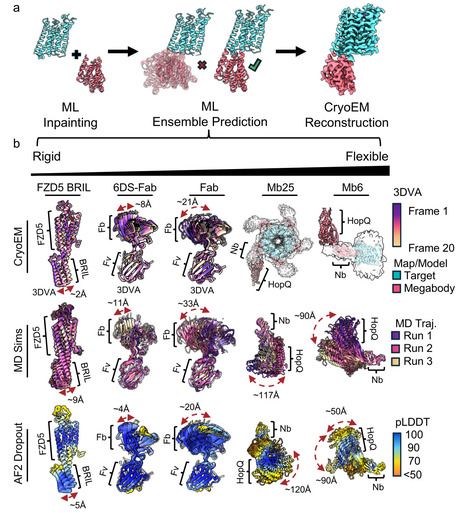
G-protein coupled receptors (GPCRs) mediate precise ligand-specific signaling profiles, yet structural visualization of how ligands alter receptor conformational landscapes in the absence of signaling partners or mimetics has proven challenging. Here we show that by combining generative protein design with deep-learning based ensemble prediction we can reliably design fiducial markers to facilitate cryogenic electron microscopy (cryoEM) of GPCRs at arbitrary fusion points, enabling the visualization of previously intractable states. We validate the approach with high-throughput determination of inactive state structures of four pharmaceutically relevant GPCRs, allowing for key details of receptor pharmacology to be resolved in each case. We then engineered an extracellular fiducial marker for the prototypical β2-adrenergic receptor that enabled direct structural characterization of the rearrangement of key intracellular motifs in the absence of G-protein. Comparison with recent co-folding models highlights major gaps in current methods for predicting ligand-induced GPCR conformations. These results present a generalizable framework for accessing traditionally inaccessible structural states of small, dynamic proteins. Visualizing how drugs reshape GPCR receptors by cryo-EM has been difficult for many conformational states. Here, authors use designed protein fiducials to image elusive GPCR states, revealing details of ligand induced receptor activation.
|
|
Scooped by
mhryu@live.com
October 23, 7:58 PM
|
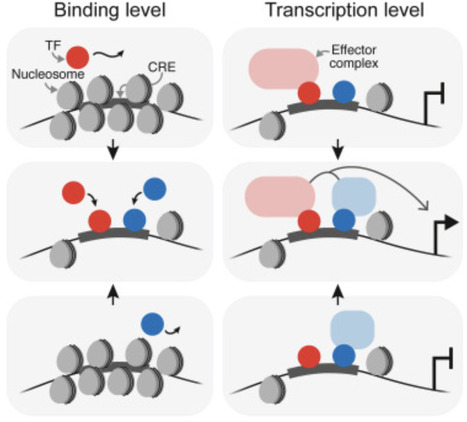
Cis-regulatory elements (CREs) integrate signals from transcription factors (TFs) to control gene expression, yet mechanisms underlying this information processing remain poorly understood. Deep learning demonstrates expression is predictable from CRE sequence, suggesting that unifying principles of regulation are within reach. Here, we propose a framework dividing CRE regulatory logic into two layers: binding-level integration, where cooperative TF interactions modulate DNA occupancy; and transcriptional-level integration, where TF effector domains and cofactors convert binding configurations into expression. At the binding level, we classify TF cooperativity along orthogonal axes—specific versus general, and symmetric versus asymmetric—encompassing protein-protein interactions, nucleosome competition, pioneer activity, and assisted loading. At the transcriptional level, cofactor interactions generate additive, synergistic, or antagonistic logic. Emerging from this complexity is a recurring quantitative pattern: “multiplication plus saturation,” reconciling binding specificity with flexibility, explaining how short DNA motifs generate precise expression while maintaining evolvability, and informing synthetic biology and therapeutic applications.
|
|
Scooped by
mhryu@live.com
October 23, 7:41 PM
|
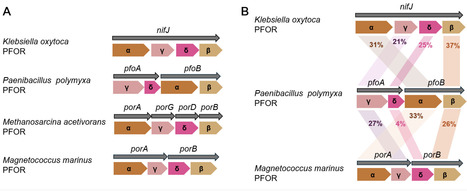
Nitrogenase requires low-potential electrons to reduce N2 to produce ammonia. The direct electron donors to nitrogenase are reduced ferredoxin (Fd) and flavodoxin (Fld). The enzymes that reduce Fd or Fld for N2 fixation mainly include pyruvate-flavodoxin (ferredoxin) oxidoreductase (PFOR), ferredoxin-NADP+ reductase (FNR), Rhodobacter nitrogen fixation protein complex (Rnf), and bifurcating electron transfer flavoprotein complex (Fix). Paenibacillus polymyxa is a model for studies of nitrogen fixation, but the pathways of electron transport to nitrogenase are unknown. In this study, we characterized the electron transport pathway to nitrogenase in P. polymyxa. Deletion analysis showed that among 15 genes encoding Fds, Fld, and Fd-/Fld-reducing enzymes, only cbiW gene and pfoAB operon decreased nitrogenase activity (31% and 37% relative to wild type, respectively). Triple deletion of cbiW and pfoAB led to a reduction of nitrogenase activity by 61%. Elemental analysis and ultraviolet-visible absorption spectroscopy revealed that the purified Fd protein encoded by cbiW is a [2Fe-2S] Fd. The purified Fd protein was able to substitute for Klebsiella NifF (Fld) to transfer electrons from pyruvate to Klebsiella nitrogenase in vitro. The pfoAB operon encodes PFOR, which is composed of two subunits. Klebsiella oxytoca NifJ (PFOR) was able to support nitrogen fixation in the ΔpfoAB mutant. These findings suggest that [2Fe-2S] Fd encoded by cbiW and PFOR encoded by the pfoAB operon constitute an electron transport pathway to Paenibacillus nitrogenase.
|
|
Scooped by
mhryu@live.com
October 23, 7:20 PM
|
The genomic regions regulating gene expression are often themselves transcribed into a variety of noncoding RNAs (ncRNAs). However, the regulatory roles of this noncoding transcription remain largely unknown. By using live imaging, we reveal that the sequential transcription of ncRNAs emanating from distinct regulatory elements underlies gene activation in Drosophila embryos. Single-allele covisualization uncovers that optimal gene activation is achieved by only moderate levels of enhancer activity. Disrupting enhancer-associated ncRNAs causes precocious gene activation, providing evidence that ncRNAs control the timing of gene expression in development. We further show that enhancer transcription can regulate long-range interactions within complex regulatory landscapes. We propose that ncRNAs locally modulate regulatory element activity in cis to shape genome organization and orchestrate the temporal control of gene expression in development.
|
|
Scooped by
mhryu@live.com
October 23, 2:52 PM
|
In this perspective, we discuss the evolving landscape of next-generation probiotics (NGPs), challenging the ambiguity of the term and proposing a framework for unifying the concepts associated with the field. We highlight the potential of novel candidates, including lactic acid bacteria and other commensal microorganisms from untapped taxa and genetically engineered strains, while examining their applications for human health. Rather than defining NGPs as a distinct category, we propose using the term to describe microbial-based health-promoting products in development, particularly where novelty relates to the organism, mechanism, engineering strategy, formulation, or targeted function. We address their safety concerns and regulatory challenges, while clarifying the divergence between therapeutic and dietary supplement pathways. We argue that as these products become more fully characterized and their safety, efficacy, intended use, and regulatory pathway are established, more specific terminology may take precedence, including the long-accepted probiotic category once efficacy and safety are established, or the live biotherapeutic products category if they are specifically developed for therapeutic applications. We suggest that the future of the ‘beneficial microbes for health’ field depends on striking a balance between the benefits of innovation and the necessity of rigorous safety and efficacy considerations.
|
|
Scooped by
mhryu@live.com
October 23, 2:02 PM
|
Replication of low-copy-number iteron plasmids is tightly controlled by initiator proteins that coordinate replication initiation with transcriptional autoregulation. In the F-plasmid system, RepE fulfills both functions through a balance between monomeric and dimeric forms. Here, random mutagenesis of the repE gene combined with high-stringency selection was used to identify variants that modulate plasmid copy number. Two point mutations, A78S and Q45L, were identified and shown by qPCR and ddPCR to increase the copy number of plasmid pFO1 relative to the parental repE(R118P) allele. The mutations exhibited growth-phase-dependent effects, with A78S having a stronger impact during exponential growth and Q45L during stationary phase. A double mutant showed a cumulative effect, particularly in exponential phase. RepE transcription increased in the presence of pFO1 and correlated with plasmid copy number, supporting a titration-based regulatory model. These findings demonstrate that engineering RepE variants enables tunable control of plasmid copy number, offering a useful strategy for optimizing gene dosage in synthetic biology and biotechnological applications.
|
|
Scooped by
mhryu@live.com
October 23, 1:47 PM
|
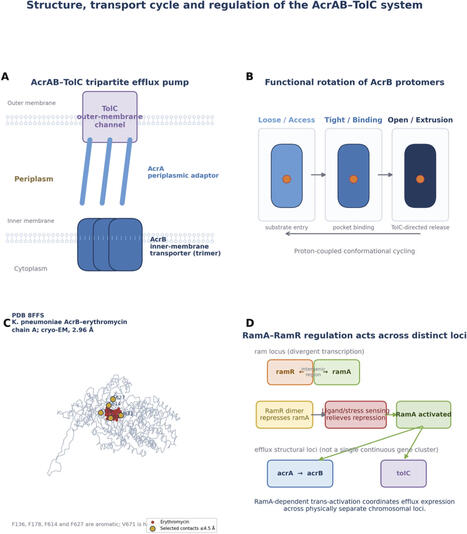
Efficient extracellular secretion remains a major bottleneck in microbial cell factories, motivating the search for chassis organisms with native high-throughput transport capacity. Klebsiella quasipneumoniae (Kqp), a recently delineated member of the Klebsiella pneumoniae species complex, encodes diverse secretion-related systems, including RND-type efflux pumps, type II and type VI secretion systems (T2SS/T6SS), outer membrane vesicle (OMV) pathways, and capsular polysaccharide (CPS) export machinery. In clinical settings, these systems contribute to opportunistic pathogenicity, antimicrobial resistance, and host adaptation; in biotechnology, they may provide transport modules that can be detoxified and redirected for extracellular production. This review examines the feasibility of reprogramming Kqp from a pathogen-related bacterium into a safe industrial secretion chassis. We summarize the molecular architecture, substrate-recognition logic, and regulatory networks of major secretion pathways, then discuss CRISPR-Cas-driven removal of virulence and resistance determinants, biocontainment, and synthetic-biology strategies for reallocating secretory capacity. We further survey potential applications in bulk chemicals, human milk oligosaccharides, recombinant proteins, and industrial enzymes. Finally, we identify key barriers to industrial translation, including growth-secretion energetic trade-offs, periplasmic protein misfolding, endotoxin-associated downstream processing, OMV heterogeneity, scale-up robustness, and quantitative safety validation. Establishing Kqp as an industrial chassis will require evidence-tiered strain selection, quantitative secretion-capacity measurement, regulatory decoupling of resistance-associated pumps, and One Health-aligned safety assessment. Direct evidence in K. quasipneumoniae remains limited; therefore, this review distinguishes direct Kqp observations from studies in other KpSC members or K. oxytoca, evidence from non-Klebsiella hosts, and forward-looking proposals. OMV yield is considered separately from lipid A/endotoxin-related properties.
|
|
Scooped by
mhryu@live.com
October 23, 1:29 PM
|
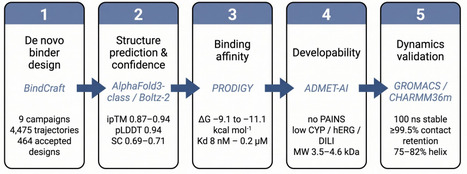
UnibindMD is an integrated computational platform that turns a protein target into validated de novo peptide-binder leads by chaining five orthogonal stages: deep-learning binder design (BindCraft), independent structure prediction and confidence assessment (AlphaFold3-class / Boltz-2), binding-affinity prediction (PRODIGY), developability profiling (ADMET-AI), and all-atom molecular-dynamics validation (GROMACS/CHARMM36m). The platform targets a persistent weakness of single-method design, namely that confidence metrics such as ipTM and pLDDT are self-consistent but not independent, so a design predicted to be confident may still be physically unstable. We demonstrate UnibindMD on three Andes virus (ANDV) intervention points, a setting of urgent need given a case fatality rate approaching 40%, a 2026 MV Hondius cruise-ship outbreak (13 cases, 3 deaths), and no approved therapeutic: the Gn-Gc glycoprotein complex (viral entry), the L-protein endonuclease domain (cap-snatching; PDB 6Q99), and the host VEGFR2 kinase (vascular permeability; PDB 2XIR). Across nine BindCraft campaigns (4,475 trajectories) the platform produced 464 accepted binders (acceptance 2.3-67.6%), with top candidates reaching ipTM = 0.94 and pLDDT = 0.94. The three selected leads were predicted by PRODIGY to bind with G = -9.1 to -11.1 kcal mol-1 (Kd 8 nM-0.2 μM), and all three remained stably bound over 100 ns of unrestrained molecular dynamics (receptor CαRMSD ≤ 2.6 Å, retained peptide fold, ≥ 99.5% of frames preserving more than half of the initial interface contacts, and 75-82% helical content). ADMET-AI flagged the expected poor oral bioavailability of large polar peptides and no PAINS alerts. By converting single-prediction design into a cross-validated, dynamics-tested workflow, UnibindMD provides a rapid, scalable, and generalizable path from any protein target to validated peptide-binder candidates.
|
|
Scooped by
mhryu@live.com
October 22, 11:12 PM
|
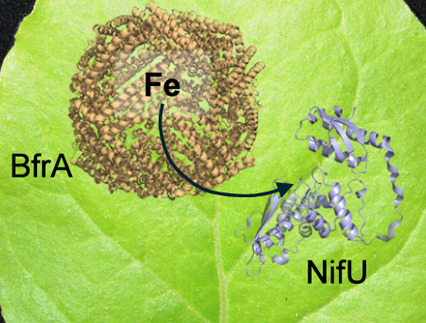
Generating crops that utilize atmospheric nitrogen rather than soil nitrogen requires the transfer of nitrogenase structural genes together with genes required for the biosynthesis of nitrogenase iron−sulfur cofactors. Scaffold proteins, such as NifU, are the starting point of nitrogenase cofactor assembly. While NifU has been successfully produced in plants, its optimal production was achieved with iron supplementation in the nutrient solution. This suggests a faulty connection with the endogenous iron trafficking network, indicating the need for specific elements targeting iron to NifU. Here, we identify bacterioferritin A (BfrA) as an iron donor to NifU. Co-expression with BfrA increases NifU production in Nicotiana benthamiana, leading to doubling the production of NifH, a nitrogenase structural protein that is also required for the synthesis of more complex nitrogenase cofactors. This work underscores the importance of transferring metal delivery systems alongside metalloproteins when engineering heterologous systems.
|
|
Scooped by
mhryu@live.com
October 22, 10:58 PM
|
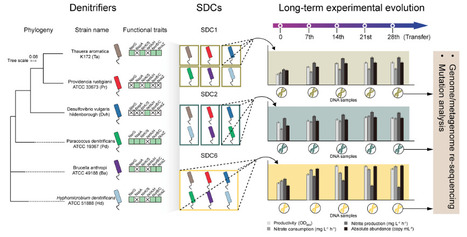
Understanding how interspecific interactions shape ecosystem functioning is a fundamental and longstanding question in ecology, especially along an evolutionary trajectory. Here, we used experimental evolution to evaluate how microbial interactions and adaptive mutations shaped community-level functions in synthetic denitrifying communities (SDCs) along a species richness gradient (one, two, and six species). Our results showed that competitive microbial interactions were observed and significantly (P < 0.05) enhanced denitrification rates in the six-species SDCs during experimental evolution. Genome and metagenome re-sequencing analysis revealed the highest fixed non-synonymous and frameshift mutations within the six-species SDCs, among genes involved in nutrient transport, DNA repair, and secretion pathways. AlphaFold2 protein structure prediction, molecular docking, and Evo 2 host fitness prediction indicated that the fixed mutations could improve host strain fitness, and partial least squares path modeling (PLS-PM) further revealed that interspecific competition accelerated adaptive mutations, thereby reinforcing community-level functions, indicating potential Red Queen dynamics. This study offers novel insights into our understanding of eco-evolutionary mechanisms linking species richness, microbial interactions, adaptive mutations, and community functions, with important implications for active nitrogen removal and greenhouse gas mitigation in diverse ecosystems.
|
|
Scooped by
mhryu@live.com
October 22, 4:34 PM
|
Prokaryotic genomes encode a vast repertoire of noncoding elements such as ribozymes, riboswitches, and CRISPR RNAs. The sequence-level diversity of these intergenic elements precludes alignment-based clustering that would enable annotation and discovery of novel noncoding RNA families. Here, we show that a genomic language model fine-tuned on prokaryotic intergenic sequences learns representations of noncoding elements from DNA sequence. We used these embeddings to organize 376 million intergenic regions from 1.3 terabase pairs of prokaryotic genomes and metagenomes into 5.14 million clusters. This approach nominated 20,149 clusters of putative structured RNAs with a candidate protein partner. Periodicity in token entropies revealed internal architecture, recovering CRISPR, TIGR and VIPR arrays together with their cognate effector proteins from sequence alone. Targeted clustering of bridge recombinase-adjacent sequences identified bridge RNA variants with longer guide recognition sequences than any characterized ortholog. Finally, we directed teams of AI agents to mine the database under researcher-defined criteria, uncovering divergent riboswitches, ribozymes, and other structured RNAs with biological hypotheses for their function. We release a publicly available database that transforms the dark matter of prokaryotic genomes into an explorable resource for human-agent molecular discovery.
|
|
Scooped by
mhryu@live.com
October 22, 12:44 PM
|
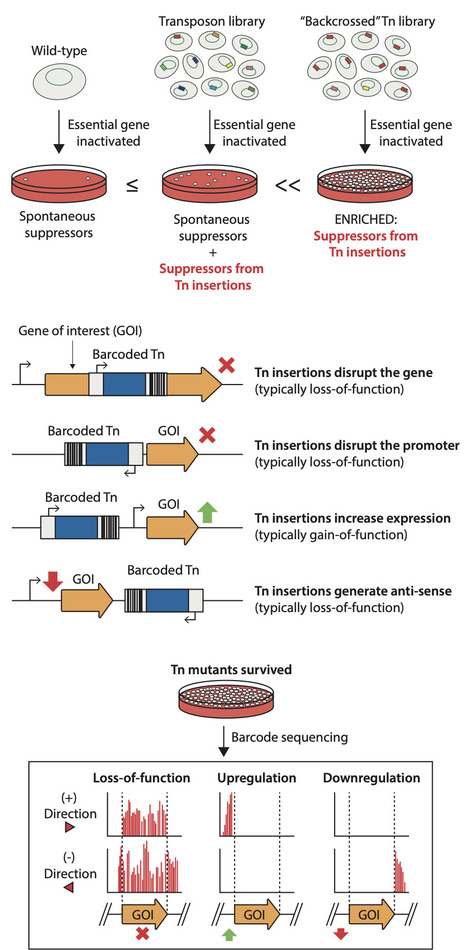
Historically, suppressor analysis has been a powerful tool for uncovering the functions of essential genes. In this study, we report Sup-seq, a transposon-sequencing technique with random DNA barcodes that enables genome-wide detection of genetic suppression. When applied to seven essential genes in Streptococcus pneumoniae, Sup-seq revealed 53 suppressors across various cellular pathways. These suppressors function through diverse mechanisms, such as gene inactivation, overexpression, or downregulation, depending on the locations and orientations of the transposon insertions. Our proof-of-concept experiments identified a previously uncharacterized transcriptional regulator that controls chaperone and ribosomal protein expression, as well as a mechanism that bypasses the requirement for acetate kinase for growth. Furthermore, we found that phosphatidylglycerol is conditionally essential in pneumococcus, revealing an unexpected link between phospholipid synthesis and potassium transport. These findings demonstrate that Sup-seq is a facile approach for providing mechanistic insights into the functions of essential genes. Additionally, Sup-seq can be adapted to other bacteria, aiding in the identification of new antimicrobial targets. Here the authors present ‘Sup-seq’, a transposon-based sequencing technique that enables genome-wide detection of genetic suppression in bacteria. They use it to identify 53 genetic suppressors of seven essential genes in pneumococcus, revealing new links among metabolism, ribosome assembly, and membrane lipid synthesis.
|
|
|
Scooped by
mhryu@live.com
Today, 11:49 AM
|
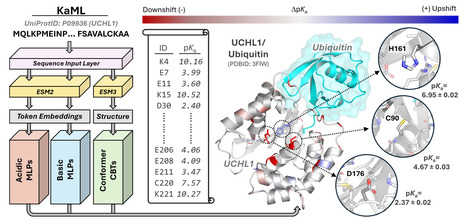
Ionization states are crucial to protein function, yet predicting their equilibrium constants remains challenging. Here we present KaML-ESMs, sequence-based neural networks built on the ESM protein language models and trained on the GAINES-augmented PKAD-3r dataset. KaML-ESM2/ESMC outperform alternative approaches across four benchmarks, achieving root-mean-square errors of about 0.5 units for Asp/Glu/His/Cys/Tyr/Lys residues in native proteins. Performance degrades on 89 buried engineered mutations lacking evolutionary support, revealing a limitation that supervised training may help overcome. These results suggest that protein sequence, through its evolutionary context, encodes not only structure and function but also, indirectly, electrostatic characteristics. Applying KaML-ESM2 to the entire human proteome, we show that predicted pKa values can identify functional sites and infer catalytic mechanisms. We offer KaML, a sequence-based pKa prediction platform supporting biological exploration, protein engineering, and biomolecular simulation. With further development, GAINES could provide a general framework for addressing data scarcity in machine learning. Ionization states are crucial to protein function, yet predicting their equilibrium constants remains challenging. Shen et al. developed KaML-ESMs, sequence-based models that predict pKa values with accuracy rivaling experiment.
|
|
Scooped by
mhryu@live.com
Today, 11:23 AM
|
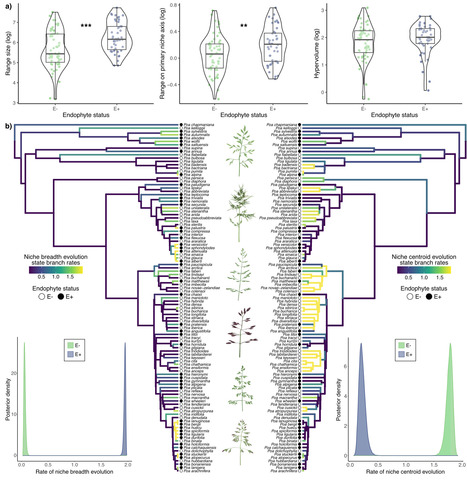
Mutualistic interactions are widely hypothesized to broaden ecological niches and promote diversification, yet evidence for this relationship at macroevolutionary scales remains limited. Here, we investigate how fungal seed endophytes (Epichloë) influence niche breadth and diversification across over 100 species of the globally-distributed grass genus Poa. Combining global-scale climate, environmental, phylogenetic, and symbiosis data, we construct species distribution models for suites of symbiotic and non-symbiotic plant species, and use phylogenetic comparative methods to assess the influence of these symbionts on niche breadth, niche breadth evolution, and diversification. Endophyte-positive species have ~1.4 times broader primary niche axes (most often associated with temperature tolerances), ~2.5 times larger geographic ranges, and >30 times faster rates of niche breadth evolution than non-symbiotic species. Endophytic plants also have significantly drier climate centroids, consistent with endophyte-mediated drought tolerance, while simultaneously showing constrained niche centroid evolution. Despite pronounced niche expansion, hidden state speciation and extinction models reveal two modes of diversification: endophytes decrease diversification rates in young lineages, and have a neutral effect in older lineages. Together, our results indicate that mutualists can be associated with broader niches without necessarily increasing diversification, highlighting a potential decoupling between niche expansion and diversification. Mutualistic interactions are hypothesized to broaden ecological niches and promote diversification, yet evidence for this relationship remains limited. This study finds that, across Poa grasses, fungal endophytes are associated with broader niches and faster niche breadth evolution, but reduced diversification rates.
|
|
Scooped by
mhryu@live.com
October 23, 7:45 PM
|
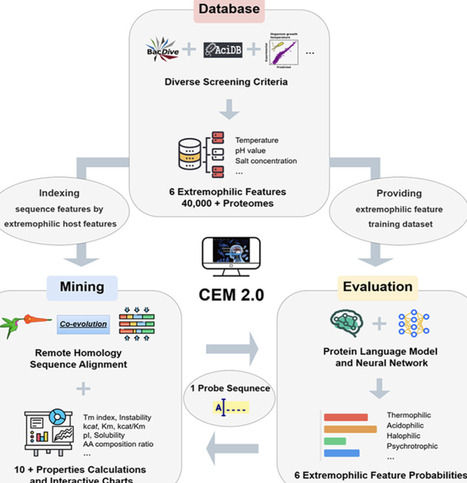
Extremophilic proteins offer immense potential for diverse applications; however, their discovery is hindered by the reliance on predefined templates and domain-specific knowledge. Therefore, new methods are required to mine extremophilic proteins without relying on existing templates, enabling a comprehensive exploration of functional extremophilic proteins across the entire protein universe. Here, we propose a co-evolution–guided protein mining framework, CEM 2.0 (Co-Evolution Mining 2.0), comprising three synergistic components: (1) an extensive and diverse extremophilic proteome database covering six extremophilic features, (2) a multidimensional evaluator for six extremophilic features, leveraging the protein language model ProteinBERT, and (3) an integrated and interactive mining pipeline equipped with advanced tools for computing >20 properties of proteins. This framework was implemented on an interactive web platform ( https://cem.sjtu.edu.cn/) equipped with intuitive visualization tools. We conducted wet-lab validation, performed a series of computational validations using published studies, and made recommendations for novel PETases, thereby demonstrating the effectiveness and potential of CEM 2.0 for the template-independent discovery of extremophilic proteins. CEM 2.0 is expected to mine functional extremophilic proteins across the protein universe in terms of evolutionary distance and serve as a valuable tool for protein engineering and synthetic biology.
|
|
Scooped by
mhryu@live.com
October 23, 7:24 PM
|
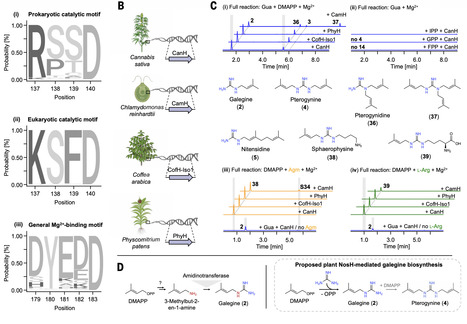
Guanidine exists almost exclusively as a resonance-stabilized cation, making its direct enzymatic functionalization challenging. We identified the cyanobacterial enzyme NosH, which catalyzes direct prenyl transfer to free guanidine. Structural and mechanistic analyses revealed guanidine activation by a two-aspartate clamp coupled to prenyl-donor binding through a distinct catalytic architecture. Guided by these features, we identified a large family of guanidine prenyltransferases across bacteria and plants. Plant enzymes produced guanidine alkaloids such as galegine in vitro and heterologous expression of a plant ortholog in Nicotiana benthamiana produced galegine in planta, providing a direct enzymatic link to plant-specialized metabolism.
|
|
Scooped by
mhryu@live.com
October 23, 2:55 PM
|
Selective oxidation reactions are critical steps in catalysis towards industrial-relevant compounds. Flavin-dependent enzymes of the VAO/PCMH and GMC families provide numerous candidates that can be applied in various formats and scales. Progresses in catalyst production, mechanistic understanding, structural information, and protein engineering enabled their successful applications. Hence, some candidates are already in use for large-scale chemical syntheses and biosensor applications such as vanillin production, biofuel cells, blood glucose sensing, and glucose monitoring in foods and beverages. The most recent advances related to these oxidases are summarized herein, highlighting recent developments beyond well-known applications that pave the way for novel developments.
|
|
Scooped by
mhryu@live.com
October 23, 2:43 PM
|
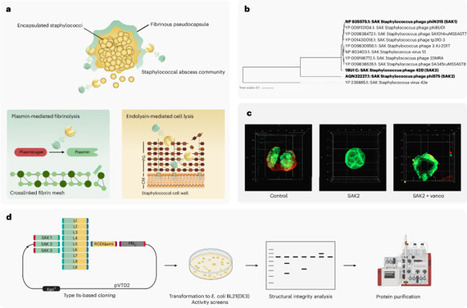
The rise of antibiotic resistance, particularly among pathogens such as methicillin-resistant Staphylococcus aureus, necessitates innovative antibacterial strategies. Here we introduce FibriLysins, a class of engineered enzymes combining staphylokinase and endolysin, which dismantle fibrin-encapsulated multicellular staphylococcal communities, including biofilms and staphylococcal abscess communities (SACs). We characterized diverse staphylokinases, developing a library of 27 FibriLysins, with a 78% positive hit rate. The lead variant, FL-028, has significant antibacterial activity against prevalent clinical S. aureus isolates. In vitro, FL-028 reduced bacterial counts in SACs by 3.4 log and in biofilms by 2.3 log. In vivo, FL-028 exhibits superior efficacy in a mouse model of bone fracture-related infection, outperforming vancomycin by 1 log and eliminating SAC presence in tissues. FL-028 shows no adverse effects on bone healing, as confirmed by histological examination, mechanical testing and micro-computed tomography analyses. These preclinical findings highlight the potential of FibriLysins as a robust and safe therapeutic approach against difficult-to-treat staphylococcal infections. Engineered FibriLysins dismantle fibrin-encapsulated Staphylococcus aureus communities and outperform vancomycin in a preclinical model of fracture-related infection.
|
|
Scooped by
mhryu@live.com
October 23, 2:01 PM
|
Scoring functions built with advanced deep learning algorithms have recently gained prominence in protein–ligand pose generation and binding affinity prediction for computational drug design. However, several shortcomings make it increasingly difficult to evaluate scoring functions for a specific task across studies. Transparent disclosure of training data, code and procedures is essential to permit reproducibility, reuse and independent scrutiny. Guarding against data leakage, sampling bias and flawed design is equally important. Meaningful comparisons of scoring functions also require common data splits, checks for run-to-run variability and performance baselines. Here we propose a set of guidelines to help the community identify the most promising methods for addressing these important problems. Ballester and Tran-Nguyen advocate for transparency in data, code and experimental procedures in the evaluation of protein–ligand predictions. They discuss good practices for preventing data leakage and bias, enabling fair, reproducible and meaningful comparisons across different scoring methods.
|
|
Scooped by
mhryu@live.com
October 23, 1:44 PM
|
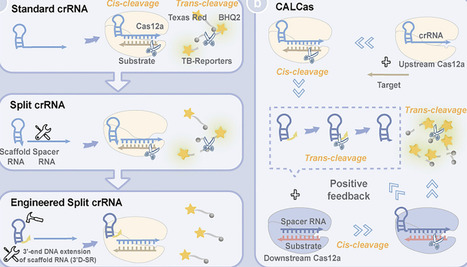
Amplification-free CRISPR detection has attracted significant attention for its capacity to circumvent the intricacies of nucleic acid amplification. Yet, the persistent challenge of high background signals remains a critical bottleneck, motivating intensive efforts toward background suppression strategies. Here, we present a 3’-end DNA extension of scaffold RNA (3’D-SR) conformational switch to drive the cascade-activatable and leakage-suppressed Cas12a system, a mechanism we denote as CALCas. By engineering the crRNA of Cas12a system and introducing steric hindrance as an activation barrier, we reshape crRNA into a “threshold-controlled” smart molecular switch, enabling Cas12a to shift from “inhibition” to “precise activation”. Unlike other Cas cascade reaction generally suffer from internal self-activation that leads to uncontrolled background increase, this newly developed framework, leveraging a DNA-mediated allosteric regulation mechanism and a leakage-suppressed design based on inherent steric hindrance, effectively solves the dilemma between signal enlargement and minor background variation. Through rational design, this cascaded CRISPR system achieves femtomolar sensitivity (over 10,000-fold higher than single Cas12a system), enables rapid HPV16/18 detection, and reduces assay time to under 30 min. This engineered crRNA approach unlocks a new pathway to reprogram Cas12a's enzymatic activities with a stabilized benchmark signal that can be critical in precise quantitative nucleic acid detection.
|
|
Scooped by
mhryu@live.com
October 23, 12:42 PM
|
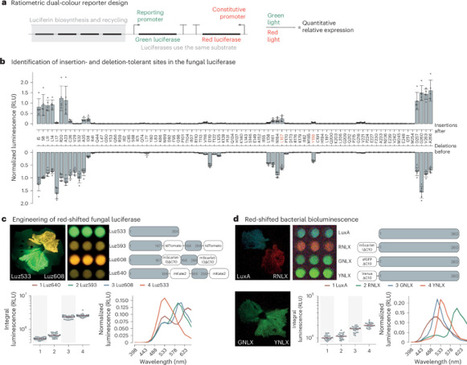
Self-sustained luminescence enables non-invasive monitoring of molecular events at the whole-organism level, but its intensity depends on both gene expression and cellular metabolism, limiting quantitative interpretation. Here we present an approach that disentangles the contributions of gene expression and metabolism. By co-expressing an engineered reference luciferase variant that uses the same substrate but emits light at a different wavelength and monitoring the color ratio, we substantially reduced signal variability, enabling quantitative physiological imaging despite changes in metabolic activity. In glowing plants, monitoring colour shifts instead of changes in luminescence intensity enables quantitative measurements of physiology despite changes in metabolic activity.
|
|
Scooped by
mhryu@live.com
October 22, 11:01 PM
|
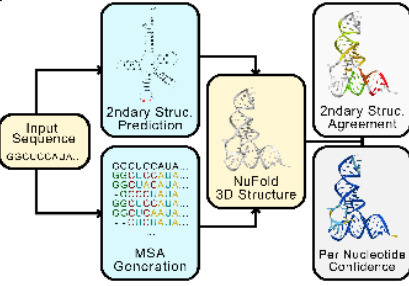
To bridge the gap between rapid RNA sequence discovery and slow experimental characterization, we present the NuFold RNA Structure Database https://nufold.kiharalab.org/ a resource providing large-scale, precomputed full-atom RNA tertiary structures. By removing the computational barriers associated with running the method locally, this database makes high-accuracy RNA structure models readily accessible to the biological research community for a wide range of applications.All predicted structures and corresponding data are available at https://nufold.kiharalab.org/. The source code for the NuFold program is available at https://github.com/kiharalab/NuFold.
|
|
Scooped by
mhryu@live.com
October 22, 10:48 PM
|
Intestinal colonization by Klebsiella pneumoniae is recognized as a pivotal prerequisite for its systemic dissemination and subsequent invasive infection; however, the metabolic basis and regulatory mechanisms underlying this process remain poorly understood. In this study, we employed transposon insertion sequencing (Tn-seq) to systematically identify metabolic genes associated with intestinal colonization in K. pneumoniae. By integrating high-throughput screening with in vivo phenotypic validation, we identified the global transcriptional regulator DksA as a key factor that markedly enhances K. pneumoniae intestinal colonization. Mechanistically, DksA may contribute to intestinal colonization through modulation of central carbon metabolism. Integrated transcriptomic and metabolomic analyses, together with homology modeling and molecular dynamics simulations, suggested that DksA may contribute to allosteric remodeling of key RNA polymerase structural domains, thereby influencing the expression of metabolic genes, carbon source utilization, and intestinal colonization by K. pneumoniae. Collectively, our findings highlight the important role of metabolic plasticity associated with transcriptional regulation in K. pneumoniae intestinal colonization and provide a theoretical framework and potential targets for developing anti-colonization strategies that exploit pathogen metabolic vulnerabilities.
|
|
Scooped by
mhryu@live.com
October 22, 4:28 PM
|
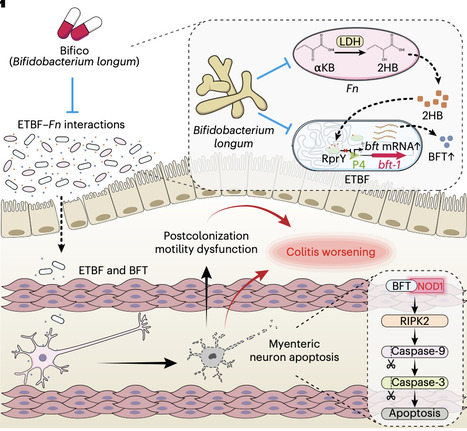
Gut dysmotility is a common long-term effect of enterotoxigenic Bacteroides fragilis (ETBF) colonization but the underlying mechanisms are not clear. Here, using mouse colonization models, we found that ETBF-secreted B. fragilis toxin (BFT) causes loss of colonic myenteric neurons, which are crucial regulators of intestinal motility. BFT induced apoptosis in these neurons through the NOD1–RIPK2–CASP9–CASP3 pathway, leading to gut dysmotility that persisted even after the clearance of ETBF. Analyses of human inflammatory bowel disease and colorectal cancer patient cohorts showed a positive correlation between ETBF and Fusobacterium nucleatum abundance. In mice, F. nucleatum-derived 2-hydroxybutyric acid enhanced BFT production in ETBF by alleviating RprY-mediated suppression and aggravated neurotoxicity and dysmotility. This enteric neuron loss and dysmotility enhanced susceptibility to dextran sulfate sodium-induced colitis in mice. Therapeutically, a Bifidobacterium-based probiotic attenuated neuron toxicity and improved intestinal motility. Our findings reveal bacterial interactions regulating intestinal function and a potential clinical therapeutic approach to preventing outcomes of ETBF colonization. Fusobacterium nucleatum promotes toxin expression in enterotoxigenic Bacteroides fragilis that induces apoptosis in enteric neurons. This eventually leads to gut dysmotility that persists even after bacterial clearance but can be alleviated by using a probiotic cocktail.
|