 Your new post is loading...

|
Scooped by
mhryu@live.com
Today, 5:42 PM
|
Heavy metal contamination poses persistent risks owing to metal mobility and bioavailability. In this study, we explored a biological strategy for altering the fate of metal ions by engineering the yeast Yarrowia lipolytica to secrete alkaline phosphatase (PhoX), thereby driving enzymatic metal-phosphate precipitation. Through genetic copy number optimization and process refinement, a recombinant strain harboring three copies of phoX achieved high extracellular PhoX activity (136.67 U/mL) and demonstrated broad-spectrum precipitation of Cd2+, Mn2+, Co2+, Cr3+, Cu2+, Ni2+ and Pb2+. A designed stepwise process enabled near-quantitative removal efficiencies of 99.23% (Cd2+), 98.22% (Mn2+), 93.16% (Co2+), 90.92% (Cr3+), 89.09% (Cu2+), 78.35% (Ni2+), and 99.28% (Pb2+), with high removal capacities of up to 1251.64 mg Cd2+/g biomass. Microscopic and crystallographic analyses confirmed the formation of cell-associated, crystalline metal phosphate minerals, with Co2+ immobilized as Co3(PO4)2·8H2O. By utilizing a generally recognized as safe microorganism to convert soluble, bioavailable metals into stable mineral phases, we elucidated a biologically mediated immobilization pathway that can reduce metal toxicity and mobility, offering a sustainable mechanism for remediating metal-contaminated environments.
|
|
Scooped by
mhryu@live.com
Today, 5:18 PM
|
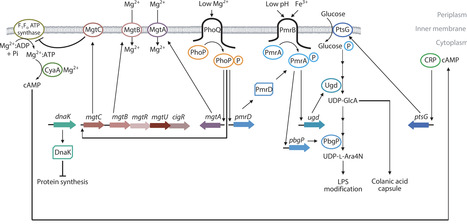
In all living cells, chemical reactions known as metabolism generate energy and building blocks for cellular structures. Metabolism produces and consumes ATP, the major currency of cellular energy. Because approximately 85% of ATP is bound to Mg2+, bacteria reduce their ATP concentration when cytoplasmic Mg2+ availability drops below a certain threshold. This reduction in ATP concentration reshapes metabolism and gene transcription as well as protein synthesis, solubility, and turnover. This article reviews how ATP acts both directly and via ATP-derived signaling molecules, such as 3′,5′-cyclic AMP (cAMP), bis-(3′,5′)-cyclic dimeric-AMP (c-di-AMP), diadenosine tetraphosphate (Ap4A), and guanosine-3′,5′-tetra- and pentaphosphate [(p)ppGpp], which direct adaptations to stresses bacteria experience in their natural environments. It also discusses how Mg2+-responsive regulatory systems supersede canonical control of metabolism and protein homeostasis by governing ATP amounts and how this bacterial response slows growth, modifies cell envelope structures, and favors survival.
|
|
Scooped by
mhryu@live.com
Today, 5:14 PM
|
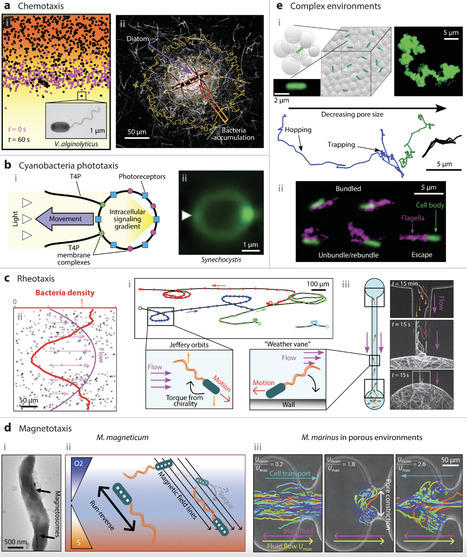
Eppur si muove—the work of Galileo Galilei marked the beginning of the quantitative study of motion for scope-observable entities. Extending this physics-based perspective to the microscope-observable microbial world has uncovered principles of motion that diverge sharply from our everyday intuition, revealing physics as being essential to understanding microbial motility. In this review, we explore the fundamental physical principles that enable and constrain the motility strategies evolved by diverse microorganisms, with a focus on prokaryotes, to enable propulsion in fluids and along surfaces. We examine how microbial motility functions in complex, heterogeneous environments, and we provide a brief evolutionary perspective on microbial motility by highlighting its costs and benefits. In summary, we show how including a physical approach in the study of microbial motility both unveils general solutions and specific adaptations and enables a better understanding of a world for which our intuition is not trained.
|
|
Scooped by
mhryu@live.com
Today, 4:58 PM
|
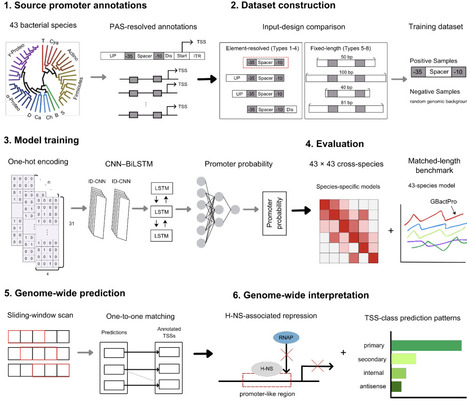
Accurate bacterial promoter prediction remains challenging across species, partly because conventional models often use fixed-length promoter windows that do not explicitly account for promoter architecture. Using promoter annotations derived from the Promoter Architecture Scanner (PAS), we found that training on sequences comprising the -35 element, spacer, and -10 element improved cross-species prediction compared with fixed-length promoter windows. Based on this element-resolved promoter definition, we developed GBactPro, a convolutional neural network-bidirectional long short-term memory (CNN-BiLSTM) model jointly trained on curated promoter data from 43 bacterial species. The jointly trained model achieved a mean AUROC of 0.9943 on held-out test sequences. Under matched-length benchmarking, GBactPro outperformed Promotech, iPro-MP, Prompt, and iPro-WAEL by 0.066-0.314 in mean AUROC. Genome-wide scanning of Escherichia coli achieved an AUROC of 0.979 and an AUPRC of 0.160 under stringent one-to-one promoter-site matching, increasing to 0.997 and 0.323, respectively, with a 100-bp positional tolerance. Unmatched predictions were enriched for H-NS binding, and independent H-NS-knockout data further supported the possibility that some predicted promoter-like regions are transcriptionally repressed. Together, these findings highlight the importance of promoter-sequence definition for cross-species prediction and demonstrate the potential of GBactPro for genome-wide promoter identification and biological interpretation.
|
|
Scooped by
mhryu@live.com
Today, 12:30 PM
|
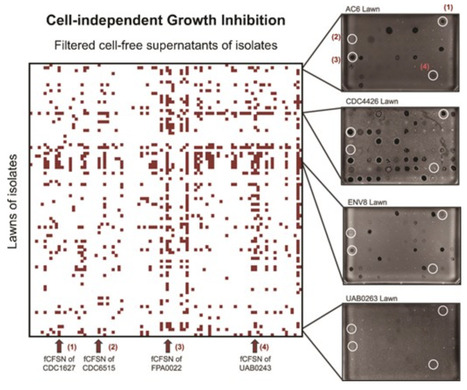
Temperate bacteriophages are increasingly recognized as important drivers of microbial evolution. However, their role in mediating competition among members of the same species during infection remains poorly defined. Pseudomonas aeruginosa (Pa), an opportunistic pathogen associated with acute and chronic infections, frequently exhibits clonal dominance within infected individuals, suggesting the existence of effective mechanisms for excluding conspecific competitors. Here, we investigate the contribution of temperate bacteriophages to intraspecies competition among clinical Pa isolates. Screening of filtered cell-free supernatants from a diverse panel of clinical strains revealed that cell-independent antagonism is widespread, with most strains inhibiting the growth of at least one other Pa isolate. Susceptibility was strongly associated with twitching motility, a type IV pili-dependent phenotype, which also serves as the primary receptor for most Pa bacteriophages. Plaque assays confirmed the presence of infectious phage particles with strain-specific host ranges. Exposure of susceptible strains to infective phages resulted in the emergence of phage-induced small colony variants (pSCVs) that retained and propagated the acquired phages and associated inhibitory capacity. Whole genome sequencing revealed extensive prophage diversity and mosaicism across clinical isolates, supporting a model in which poly-lysogeny expands competitive potential. Using isogenic pSCV systems and murine wound infection models, we demonstrate that acquisition of specific temperate phages confers a competitive advantage both in vitro and in vivo. These findings establish temperate bacteriophages as dynamic ecological weapons that shape intraspecies competition providing a mechanistic basis for clonal dominance during Pa infection and highlighting the broader role of prophages in structuring microbial populations.
|
|
Scooped by
mhryu@live.com
Today, 12:11 PM
|
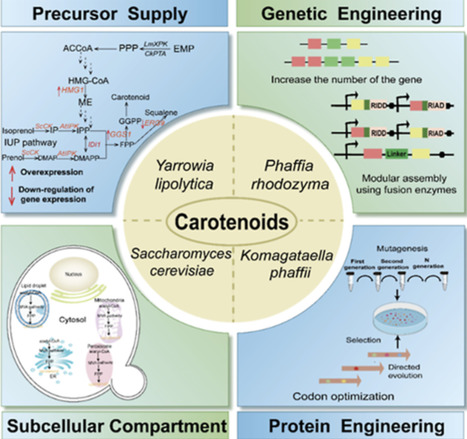
Carotenoids are versatile natural pigments with potent antioxidant, anti-cancer, and anti-inflammatory activities, making them valuable ingredients in cosmetics, animal feed, and pharmaceuticals. While traditionally sourced from plants, algae, and microorganisms, conventional extraction methods struggle to meet the rising market demand for natural products. With the rapid advancement of synthetic biology, microbial fermentation has emerged as a scalable and efficient alternative for carotenoid production. This review paper examines four promising yeast platforms Phaffia rhodozyma, Saccharomyces cerevisiae, Yarrowia lipolytica, and Komagataella phaffii for the production of carotenoids. We systematically outline key modification strategies for the de novo biosynthetic process of carotenoids, primarily including heterologous pathway adaptation, precursor supply enhancement, protein engineering of rate-limiting enzymes, subcellular compartmentalization, and optimization of the fermentation process. Furthermore, we compare the inherent limitations of each yeast chassis and discuss potential solutions to overcome production bottlenecks, thereby providing a strategic roadmap for future strain and process development in support of industrial-scale microbial carotenoid production.
|
|
Scooped by
mhryu@live.com
Today, 12:01 PM
|
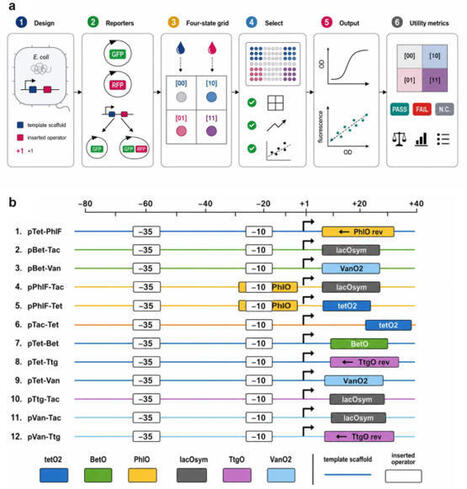
Engineering compact bacterial promoters that integrate two chemical inputs would simplify biosensors, synthetic circuits, and dynamic metabolic control, but such promoters remain difficult to build because a second operator can increase partial-state leak, reduce the fully induced state, and create sequence- or reporter-context effects. Here we engineered 12 compact Marionette-derived two-input promoter architectures in Escherichia coli by combining defined template scaffolds with added operators, then quantified 19 reporter-specific four-state truth tables under a predefined operational criterion for chemical-input AND behavior. Single-input template-promoter controls in matched reporter contexts helped separate weak parent-scaffold output from effects introduced by the second operator. Nine curated architectures passed in at least one reporter, and selected constructs delivered high-utility AND responses, including [11]/max-off separation up to > 1,367. The full pass–fail dataset showed that a statistically higher [11] state does not necessarily create a useful switch. Practical utility depended mainly on suppressing [10] and [01] leakage, matching the template scaffold to the inserted operator, and controlling long-operator orientation and local sequence context. Reciprocal architectures behaved differently, and 4 of 7 dual-reporter architectures changed operational classification with reporter context. Thus, this work delivers a compact set of candidate Marionette-compatible two-input promoter parts and a source-traceable design framework for engineering bacterial chemical-input AND switches.
|
|
Scooped by
mhryu@live.com
Today, 11:49 AM
|
Ionization states are crucial to protein function, yet predicting their equilibrium constants remains challenging. Here we present KaML-ESMs, sequence-based neural networks built on the ESM protein language models and trained on the GAINES-augmented PKAD-3r dataset. KaML-ESM2/ESMC outperform alternative approaches across four benchmarks, achieving root-mean-square errors of about 0.5 units for Asp/Glu/His/Cys/Tyr/Lys residues in native proteins. Performance degrades on 89 buried engineered mutations lacking evolutionary support, revealing a limitation that supervised training may help overcome. These results suggest that protein sequence, through its evolutionary context, encodes not only structure and function but also, indirectly, electrostatic characteristics. Applying KaML-ESM2 to the entire human proteome, we show that predicted pKa values can identify functional sites and infer catalytic mechanisms. We offer KaML, a sequence-based pKa prediction platform supporting biological exploration, protein engineering, and biomolecular simulation. With further development, GAINES could provide a general framework for addressing data scarcity in machine learning. Ionization states are crucial to protein function, yet predicting their equilibrium constants remains challenging. Shen et al. developed KaML-ESMs, sequence-based models that predict pKa values with accuracy rivaling experiment.
|
|
Scooped by
mhryu@live.com
Today, 11:23 AM
|
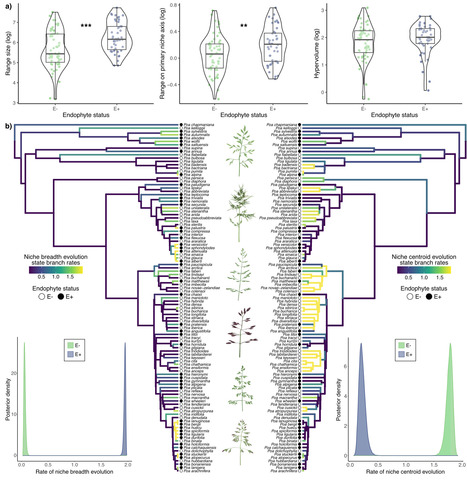
Mutualistic interactions are widely hypothesized to broaden ecological niches and promote diversification, yet evidence for this relationship at macroevolutionary scales remains limited. Here, we investigate how fungal seed endophytes (Epichloë) influence niche breadth and diversification across over 100 species of the globally-distributed grass genus Poa. Combining global-scale climate, environmental, phylogenetic, and symbiosis data, we construct species distribution models for suites of symbiotic and non-symbiotic plant species, and use phylogenetic comparative methods to assess the influence of these symbionts on niche breadth, niche breadth evolution, and diversification. Endophyte-positive species have ~1.4 times broader primary niche axes (most often associated with temperature tolerances), ~2.5 times larger geographic ranges, and >30 times faster rates of niche breadth evolution than non-symbiotic species. Endophytic plants also have significantly drier climate centroids, consistent with endophyte-mediated drought tolerance, while simultaneously showing constrained niche centroid evolution. Despite pronounced niche expansion, hidden state speciation and extinction models reveal two modes of diversification: endophytes decrease diversification rates in young lineages, and have a neutral effect in older lineages. Together, our results indicate that mutualists can be associated with broader niches without necessarily increasing diversification, highlighting a potential decoupling between niche expansion and diversification. Mutualistic interactions are hypothesized to broaden ecological niches and promote diversification, yet evidence for this relationship remains limited. This study finds that, across Poa grasses, fungal endophytes are associated with broader niches and faster niche breadth evolution, but reduced diversification rates.
|
|
Scooped by
mhryu@live.com
October 23, 7:45 PM
|
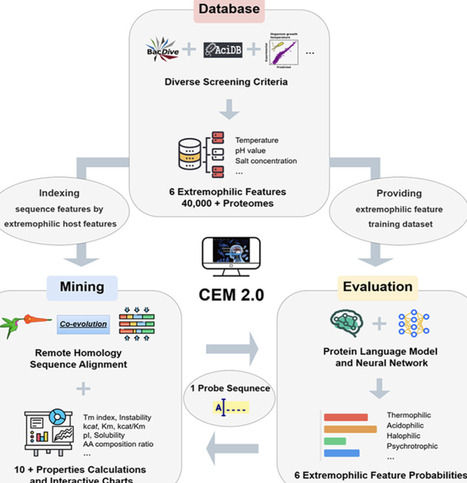
Extremophilic proteins offer immense potential for diverse applications; however, their discovery is hindered by the reliance on predefined templates and domain-specific knowledge. Therefore, new methods are required to mine extremophilic proteins without relying on existing templates, enabling a comprehensive exploration of functional extremophilic proteins across the entire protein universe. Here, we propose a co-evolution–guided protein mining framework, CEM 2.0 (Co-Evolution Mining 2.0), comprising three synergistic components: (1) an extensive and diverse extremophilic proteome database covering six extremophilic features, (2) a multidimensional evaluator for six extremophilic features, leveraging the protein language model ProteinBERT, and (3) an integrated and interactive mining pipeline equipped with advanced tools for computing >20 properties of proteins. This framework was implemented on an interactive web platform ( https://cem.sjtu.edu.cn/) equipped with intuitive visualization tools. We conducted wet-lab validation, performed a series of computational validations using published studies, and made recommendations for novel PETases, thereby demonstrating the effectiveness and potential of CEM 2.0 for the template-independent discovery of extremophilic proteins. CEM 2.0 is expected to mine functional extremophilic proteins across the protein universe in terms of evolutionary distance and serve as a valuable tool for protein engineering and synthetic biology.
|
|
Scooped by
mhryu@live.com
October 23, 7:24 PM
|
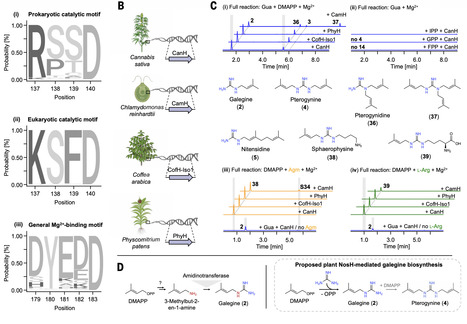
Guanidine exists almost exclusively as a resonance-stabilized cation, making its direct enzymatic functionalization challenging. We identified the cyanobacterial enzyme NosH, which catalyzes direct prenyl transfer to free guanidine. Structural and mechanistic analyses revealed guanidine activation by a two-aspartate clamp coupled to prenyl-donor binding through a distinct catalytic architecture. Guided by these features, we identified a large family of guanidine prenyltransferases across bacteria and plants. Plant enzymes produced guanidine alkaloids such as galegine in vitro and heterologous expression of a plant ortholog in Nicotiana benthamiana produced galegine in planta, providing a direct enzymatic link to plant-specialized metabolism.
|
|
Scooped by
mhryu@live.com
October 23, 2:55 PM
|
Selective oxidation reactions are critical steps in catalysis towards industrial-relevant compounds. Flavin-dependent enzymes of the VAO/PCMH and GMC families provide numerous candidates that can be applied in various formats and scales. Progresses in catalyst production, mechanistic understanding, structural information, and protein engineering enabled their successful applications. Hence, some candidates are already in use for large-scale chemical syntheses and biosensor applications such as vanillin production, biofuel cells, blood glucose sensing, and glucose monitoring in foods and beverages. The most recent advances related to these oxidases are summarized herein, highlighting recent developments beyond well-known applications that pave the way for novel developments.
|
|
Scooped by
mhryu@live.com
October 23, 2:43 PM
|
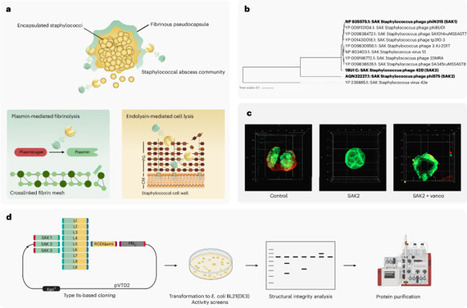
The rise of antibiotic resistance, particularly among pathogens such as methicillin-resistant Staphylococcus aureus, necessitates innovative antibacterial strategies. Here we introduce FibriLysins, a class of engineered enzymes combining staphylokinase and endolysin, which dismantle fibrin-encapsulated multicellular staphylococcal communities, including biofilms and staphylococcal abscess communities (SACs). We characterized diverse staphylokinases, developing a library of 27 FibriLysins, with a 78% positive hit rate. The lead variant, FL-028, has significant antibacterial activity against prevalent clinical S. aureus isolates. In vitro, FL-028 reduced bacterial counts in SACs by 3.4 log and in biofilms by 2.3 log. In vivo, FL-028 exhibits superior efficacy in a mouse model of bone fracture-related infection, outperforming vancomycin by 1 log and eliminating SAC presence in tissues. FL-028 shows no adverse effects on bone healing, as confirmed by histological examination, mechanical testing and micro-computed tomography analyses. These preclinical findings highlight the potential of FibriLysins as a robust and safe therapeutic approach against difficult-to-treat staphylococcal infections. Engineered FibriLysins dismantle fibrin-encapsulated Staphylococcus aureus communities and outperform vancomycin in a preclinical model of fracture-related infection.
|
|
|
Scooped by
mhryu@live.com
Today, 5:20 PM
|
Antibiotic-resistant bacteria are common in natural environments, including the microbiota of animal and human hosts. The local physical, chemical, and biological conditions of environmental patches and matrices vary both qualitatively and quantitatively. Often, these conditions are found as gradients, creating intersections that can either facilitate or inhibit the spread and evolution of antibiotic resistance (AbR). Ecological kinetics (EcoK) describes how combinations of these conditions influence the abundance and transmission of antibiotic-resistant populations, as well as the mobile genetic elements that carry resistance genes. Evolutionary dynamics (EvoD) concerns the effects of environmental conditions on genetic variation, the expression of resistance traits, and the evolvability of host bacteria, ultimately contributing to the emergence and selection of AbR. This review examines the influence of numerous environmental conditions on EcoK and EvoD. This field remains poorly understood, and further research is essential to identify high-risk areas where surveillance for the emergence and spread of AbR should be implemented.
|
|
Scooped by
mhryu@live.com
Today, 5:16 PM
|
Peptides are among the smallest functional biomolecules. Learning to systematically design their activity would revolutionize biology and medicine. Yet achieving this goal remains an open challenge. Recent advances in machine learning enable the discovery and design of peptides at scale, not only yielding new antibiotic agents effective against priority pathogens but also revealing unexpected roles for peptides in host immunity. In this review, we survey how artificial intelligence (AI) methods can be used to mine genomes, proteomes, and even ancient sequence data to uncover “encrypted” peptide antibiotics across the tree of life and over evolutionary time. We highlight emerging approaches for activity prediction, structure–activity modeling, sequence optimization, and de novo peptide generation, and we discuss workflows that tightly couple in silico design with high-throughput experimental validation. We further introduce the concept of “encrypted immunity,” in which ordinary proteins encode latent antimicrobial fragments, and we outline translational opportunities as well as remaining challenges on the path to bringing AI-designed peptides to the clinic.
|
|
Scooped by
mhryu@live.com
Today, 5:11 PM
|
Protein glycosylation involves the co-translational or post-translational addition of glycans to proteins and is a crucial protein modification in health and disease. The aim of glycoproteomics is to understand how glycosylation shapes biological processes by understanding peptide sequences, glycan structures and sites of modification in a system-wide context. Over the past two decades, mass spectrometry (MS) has emerged as the primary technique for studying glycoproteins, with intact glycopeptide analysis — the study of glycopeptides decorated with their native glycan structures — now a preferred approach across the community. In this Primer, we discuss glycoproteomic methods for studying glycosylation classes, including best practices and critical considerations. We summarize how glycoproteomics is used to understand glycosylation at a systems level, with a specific focus on N-linked and O-linked glycosylation (both mucin-type and O-GlcNAcylation). We cover topics that include sample selection; techniques for protein isolation, proteolytic digestion, glycopeptide enrichment and MS fragmentation; bioinformatic platforms and applications of glycoproteomics. Finally, we give a perspective on where the field is heading. Overall, this Primer outlines the current technologies, persistent challenges and recent advances in the exciting field of glycoproteomics. Glycoproteomic techniques give information on the structure and location of glycan protein modifications. In this Primer, Bagdonaite et al. summarize these techniques, discuss best practices for their use and explore their applications, including identifying biomarkers of disease.
|
|
Scooped by
mhryu@live.com
Today, 4:39 PM
|
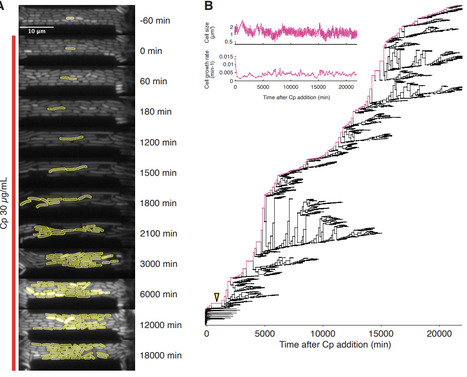
Exposing bacterial cell populations to antibiotics often results in the emergence of descendants with enhanced resistance. Phenotypic heterogeneity and global physiological responses are thought to contribute to resistance evolution. However, identifying the specific stages of drug exposure where phenotypic heterogeneity and physiological responses play critical roles remains elusive. Using long-term single-cell lineage tracking of an Escherichia coli resistance model, here we reveal that resistance-enhanced cells emerge rapidly within a few generations in a clonal bacterial cell population under chloramphenicol (Cp) exposure and stably transmit the enhanced resistance for hundreds of generations in the presence of Cp. High expression of a resistance protein was advantageous during the initial period of drug exposure, but conferred only a marginal fitness advantage during the period of resistance enhancement. Transcriptomic profiles of resistance-enhanced cell populations were lineage-specific, suggesting that multiple gene expression states can support enhanced resistance. Altogether, these results suggest that an interplay between initial phenotypic heterogeneity and divergent remodeling of gene expression drives the rapid acquisition of enhanced antibiotic resistance.
|
|
Scooped by
mhryu@live.com
Today, 12:28 PM
|
Anaerobic methanotrophic ‘Candidatus Methanoperedenaceae’ play a key role in mitigating methane emissions from freshwater sediments. Members of the family are metabolically versatile, with species coupling anaerobic oxidation of methane (AOM) to the reduction of nitrate and metal oxides. Metagenomic analyses have indicated further uncharacterized metabolic diversity, including the potential for AOM coupled to arsenate reduction (As-AOM). While As-AOM has been confirmed as an important methane sink and contributor to arsenic mobilization in freshwater environments, the underlying microbiological mechanisms remain to be empirically demonstrated. In this study stoichiometric coupling of dissimilatory arsenate reduction to methane oxidation was demonstrated using enrichment cultures of the uncultured type species ‘Ca. Methanoperedens nitroreducens’. Meta-omics revealed that arsenate oxidoreductases were exclusively encoded by ‘Ca. M. nitroreducens’ and upregulated under As-AOM conditions, confirming this species was directly responsible. These findings expand our understanding of ‘Ca. Methanoperedenaceae’ physiology and demonstrate microbial mechanisms underpinning arsenic mobilization in freshwater systems, with important implications for climate and public health.
|
|
Scooped by
mhryu@live.com
Today, 12:10 PM
|
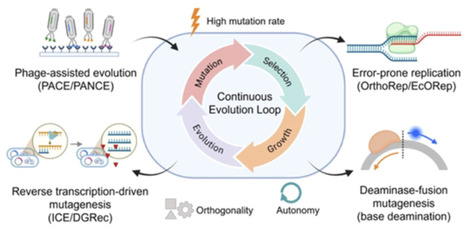
Traditional in vitro directed evolution is constrained by discrete workflows, high labor demands, and separation from native cellular contexts. In vivo continuous evolution systems overcome these limitations by enabling autonomous and targeted mutagenesis coupled with cellular proliferation. This review outlines the key parts of these systems, including elevated mutation rates (10−5−10−4 substitutions per base), orthogonality, and autonomy. We classify current platforms into some categories: phage-assisted continuous evolution, error-prone polymerase-based DNA replication systems, reverse transcription-driven mutagenesis systems, and deaminase-fusion transcription-coupled mutagenesis systems, detailing their mechanisms, optimization strategies, and applications. Finally, we address technical bottlenecks and emerging directions, including AI-driven component design, automated platforms, expanded host compatibility, and broadened industrial applications, providing a roadmap for advancing continuous evolution technology.
|
|
Scooped by
mhryu@live.com
Today, 11:50 AM
|
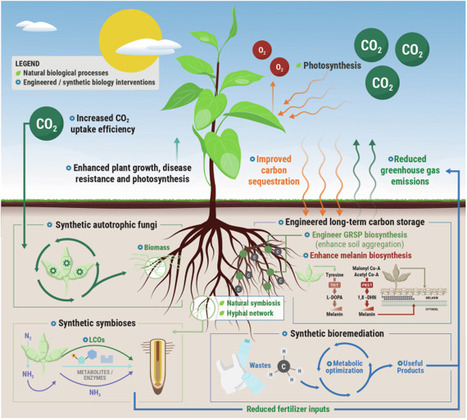
Over the past two decades, there has been a staggering increase in greenhouse gas emissions, particularly carbon dioxide (CO2) from unrestrained anthropogenic activities, leading to notable environmental shifts. Despite continuous efforts to reduce emissions, the world is not making satisfactory progress, which calls for immediate action to implement alternative strategies to enhance environmental sustainability. The Earth naturally absorbs and stores carbon through various processes, but merely keeping pace with ongoing human activities is no longer sufficient. Land plants and marine prokaryotes absorb CO2 during photosynthesis and store carbon as biomass. Eukaryotic organisms also represent a valuable resource that can provide insights into the biological fixation and removal of atmospheric CO2. Expanding the capacities of Earth’s natural systems offers enormous potential for enhancing carbon sinks and promoting environmental resilience. Although extensive synthetic engineering of plant systems has been conducted to enhance their native carbon fixation pathways, research on microorganisms, especially fungi, has been limited. In this review, we specifically explore the role of soil-borne fungi in carbon management, elucidating their responses to environmental shifts and their potential for climate change mitigation. We place particular emphasis on the role of various fungal guilds in this process, highlighting those that should be prioritized for future synthetic biology applications. We then engage with previous endeavors to address primary challenges in advancing genetic tools for these understudied fungi. Lastly, we discuss how integrating synthetic biology practices into the field of soil-associated fungi can drive advances in long-term carbon storage, enhance degradation of environmental pollutants, and improve nutrient exchange and soil health, contributing to more resilient soil ecosystems and sustainable environmental interventions.
|
|
Scooped by
mhryu@live.com
Today, 11:43 AM
|
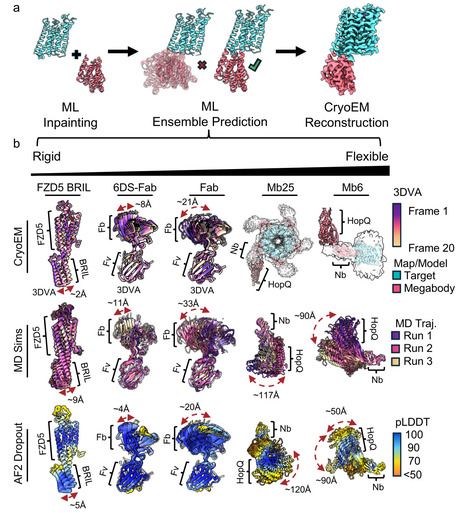
G-protein coupled receptors (GPCRs) mediate precise ligand-specific signaling profiles, yet structural visualization of how ligands alter receptor conformational landscapes in the absence of signaling partners or mimetics has proven challenging. Here we show that by combining generative protein design with deep-learning based ensemble prediction we can reliably design fiducial markers to facilitate cryogenic electron microscopy (cryoEM) of GPCRs at arbitrary fusion points, enabling the visualization of previously intractable states. We validate the approach with high-throughput determination of inactive state structures of four pharmaceutically relevant GPCRs, allowing for key details of receptor pharmacology to be resolved in each case. We then engineered an extracellular fiducial marker for the prototypical β2-adrenergic receptor that enabled direct structural characterization of the rearrangement of key intracellular motifs in the absence of G-protein. Comparison with recent co-folding models highlights major gaps in current methods for predicting ligand-induced GPCR conformations. These results present a generalizable framework for accessing traditionally inaccessible structural states of small, dynamic proteins. Visualizing how drugs reshape GPCR receptors by cryo-EM has been difficult for many conformational states. Here, authors use designed protein fiducials to image elusive GPCR states, revealing details of ligand induced receptor activation.
|
|
Scooped by
mhryu@live.com
October 23, 7:58 PM
|
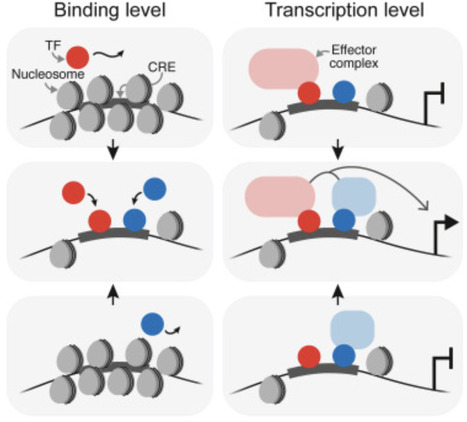
Cis-regulatory elements (CREs) integrate signals from transcription factors (TFs) to control gene expression, yet mechanisms underlying this information processing remain poorly understood. Deep learning demonstrates expression is predictable from CRE sequence, suggesting that unifying principles of regulation are within reach. Here, we propose a framework dividing CRE regulatory logic into two layers: binding-level integration, where cooperative TF interactions modulate DNA occupancy; and transcriptional-level integration, where TF effector domains and cofactors convert binding configurations into expression. At the binding level, we classify TF cooperativity along orthogonal axes—specific versus general, and symmetric versus asymmetric—encompassing protein-protein interactions, nucleosome competition, pioneer activity, and assisted loading. At the transcriptional level, cofactor interactions generate additive, synergistic, or antagonistic logic. Emerging from this complexity is a recurring quantitative pattern: “multiplication plus saturation,” reconciling binding specificity with flexibility, explaining how short DNA motifs generate precise expression while maintaining evolvability, and informing synthetic biology and therapeutic applications.
|
|
Scooped by
mhryu@live.com
October 23, 7:41 PM
|
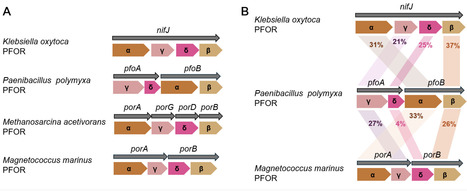
Nitrogenase requires low-potential electrons to reduce N2 to produce ammonia. The direct electron donors to nitrogenase are reduced ferredoxin (Fd) and flavodoxin (Fld). The enzymes that reduce Fd or Fld for N2 fixation mainly include pyruvate-flavodoxin (ferredoxin) oxidoreductase (PFOR), ferredoxin-NADP+ reductase (FNR), Rhodobacter nitrogen fixation protein complex (Rnf), and bifurcating electron transfer flavoprotein complex (Fix). Paenibacillus polymyxa is a model for studies of nitrogen fixation, but the pathways of electron transport to nitrogenase are unknown. In this study, we characterized the electron transport pathway to nitrogenase in P. polymyxa. Deletion analysis showed that among 15 genes encoding Fds, Fld, and Fd-/Fld-reducing enzymes, only cbiW gene and pfoAB operon decreased nitrogenase activity (31% and 37% relative to wild type, respectively). Triple deletion of cbiW and pfoAB led to a reduction of nitrogenase activity by 61%. Elemental analysis and ultraviolet-visible absorption spectroscopy revealed that the purified Fd protein encoded by cbiW is a [2Fe-2S] Fd. The purified Fd protein was able to substitute for Klebsiella NifF (Fld) to transfer electrons from pyruvate to Klebsiella nitrogenase in vitro. The pfoAB operon encodes PFOR, which is composed of two subunits. Klebsiella oxytoca NifJ (PFOR) was able to support nitrogen fixation in the ΔpfoAB mutant. These findings suggest that [2Fe-2S] Fd encoded by cbiW and PFOR encoded by the pfoAB operon constitute an electron transport pathway to Paenibacillus nitrogenase.
|
|
Scooped by
mhryu@live.com
October 23, 7:20 PM
|
The genomic regions regulating gene expression are often themselves transcribed into a variety of noncoding RNAs (ncRNAs). However, the regulatory roles of this noncoding transcription remain largely unknown. By using live imaging, we reveal that the sequential transcription of ncRNAs emanating from distinct regulatory elements underlies gene activation in Drosophila embryos. Single-allele covisualization uncovers that optimal gene activation is achieved by only moderate levels of enhancer activity. Disrupting enhancer-associated ncRNAs causes precocious gene activation, providing evidence that ncRNAs control the timing of gene expression in development. We further show that enhancer transcription can regulate long-range interactions within complex regulatory landscapes. We propose that ncRNAs locally modulate regulatory element activity in cis to shape genome organization and orchestrate the temporal control of gene expression in development.
|
|
Scooped by
mhryu@live.com
October 23, 2:52 PM
|
In this perspective, we discuss the evolving landscape of next-generation probiotics (NGPs), challenging the ambiguity of the term and proposing a framework for unifying the concepts associated with the field. We highlight the potential of novel candidates, including lactic acid bacteria and other commensal microorganisms from untapped taxa and genetically engineered strains, while examining their applications for human health. Rather than defining NGPs as a distinct category, we propose using the term to describe microbial-based health-promoting products in development, particularly where novelty relates to the organism, mechanism, engineering strategy, formulation, or targeted function. We address their safety concerns and regulatory challenges, while clarifying the divergence between therapeutic and dietary supplement pathways. We argue that as these products become more fully characterized and their safety, efficacy, intended use, and regulatory pathway are established, more specific terminology may take precedence, including the long-accepted probiotic category once efficacy and safety are established, or the live biotherapeutic products category if they are specifically developed for therapeutic applications. We suggest that the future of the ‘beneficial microbes for health’ field depends on striking a balance between the benefits of innovation and the necessity of rigorous safety and efficacy considerations.
|


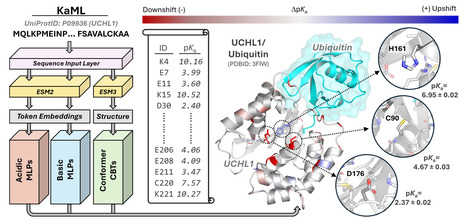







KaML accepts a user-provided protein sequence through the sequence input layer. ESM2 (or ESMC) is used to generate token embeddings, while ESM3 predicts a three-dimensional structure if not provided. Embeddings are fed to the acid and base task heads (ensemble of 500 MLPs), respectively. Optionally, the ESM3-derived or user-provided structure is processed for conformer-dependent predictions by KaMLs-CBT2. The predicted acid and base pKa's, standard errors, and pKa shifts are given in the output.