Your new post is loading...

|
Scooped by
mhryu@live.com
Today, 2:03 PM
|
Archaea play essential roles in global biogeochemical cycles, yet their diversity in environmental samples remains understudied. Limited coverage of universal primers towards archaea, their very low abundance relative to bacteria in the studied samples, inefficient cell lysis and the incompleteness of reference databases complicate archaeal 16S rRNA gene sequencing. Furthermore, standardization lacks due to the absence of commercially available mock communities. Here, we developed a mock community composed of mixtures of DNA from six archaeal and four bacterial strains with known genome sizes and 16S rRNA gene copy numbers. Five mock compositions with different archaeal–bacterial abundance ratios were used to evaluate two universal primer pairs and eight archaea-specific methods. The universal primer pairs detected all taxa but consistently overestimated bacterial abundance, indicating to be less suitable for environments where archaea are more dominant. Archaeal specific primer pairs showed variable specificity, with some exhibiting substantial bacterial co-amplification under certain conditions. The Arch349F–A806R primer pair demonstrated the highest archaeal specificity, the least technical variability, the most consistent agreement with expected mock compositions, and minimal artefactual amplification when applied to a complex bacterial mock community. To show its applicability on environmental samples, this method was applied on Boom Clay pore water samples, which indicated a much higher archaeal diversity than previously shown with a standard commercial method. Overall, this study showed the necessity of primer validation through mock communities and identified Arch349F–A806R as a promising primer pair for archaeal 16S rRNA gene sequencing, especially in samples containing methanogenic archaea.
|
|
Scooped by
mhryu@live.com
Today, 1:48 PM
|
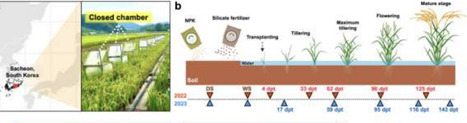
Methane emissions from rice paddies are a major source of greenhouse gases, and silicate fertilizer (SF) derived from blast furnace slag has emerged as an environmentally friendly amendment that reduces methane release. SF modifies the soil redox environment by supplying electron acceptors such as iron, but the microbial mechanisms underlying methane mitigation remain unclear. Here, we combined metagenomic profiling, microbial isolation, and greenhouse experiments to show that SF consistently enriched Burkholderia vietnamiensis together with the methanotrophs in the rice rhizosphere. Inoculation of B. vietnamiensis strain HU01 alone reproduced the methane-suppressing effect of SF and increased methanotroph abundance. Cell-free culture supernatants of HU01 stimulated the growth of Methylococcus geothermalis and Methylocystis parvus through secreted, low-molecular-weight, heat-stable metabolites. Although metabolic modeling predicted primary metabolites such as L-glutamate and cobalamin as candidate cross-feeding factors, LC–MS/MS analysis of the highly active fractions indicated that the robust bioactivity was primarily driven by a chemically complex metabolite mixture rather than these ubiquitous single compounds. These results provide direct evidence that enrichment of a single heterotrophic species can enhance methane oxidation by stimulating methanotroph activity through metabolite exchange. B. vietnamiensis HU01 represents a promising microbial resource for methane mitigation and illustrates how targeted heterotroph–methanotroph interactions can be leveraged for climate-smart rice cultivation.
|
|
Scooped by
mhryu@live.com
Today, 12:51 PM
|
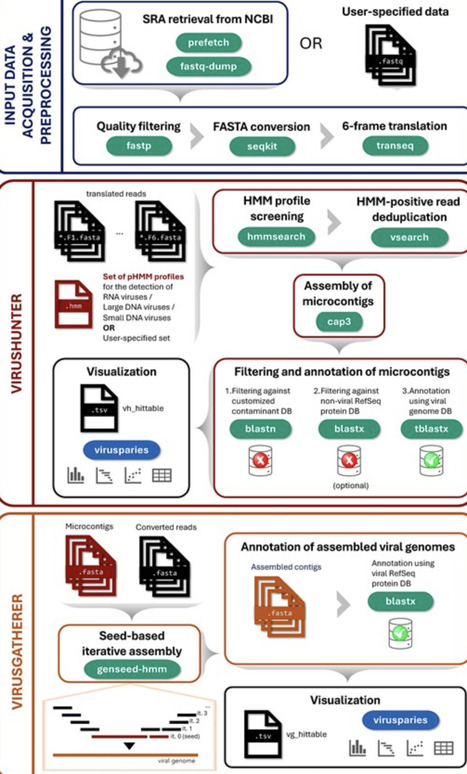
We present Virushunter and Virusgatherer, a bipartite computational workflow for discovering viral genomes from unprocessed next-generation sequencing data. Virushunter conducts a profile Hidden Markov Model-based sequence homology search for viral hallmark protein-coding sequence reads to construct microcontigs, which are used as seeds for targeted viral genome assembly by Virusgatherer. We showcase the application of Virushunter and Virusgatherer through an RNA virus-wide screen in 3348 bat transcriptome sequencing experiments from the Sequence Read Archive. Many of the discovered viruses are likely prototypes of new putative virus species, including a novel alphacoronavirus identified in a South American velvety free-tailed bat and a novel rhabdovirus in a European barbastelle bat. The tools are implemented as a reproducible Snakemake pipeline, which is available at https://github.com/lauberlab/VirusHunterGatherer.
|
|
Scooped by
mhryu@live.com
Today, 12:44 PM
|
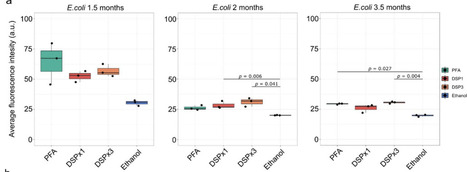
Phylogeny- or activity-based fluorescent labelling of microorganisms coupled with fluorescence activated cell sorting (FACS) is a powerful approach to rapidly isolate cells of interest from complex communities for targeted genomics. However, the method has not been widely adopted because standard paraformaldehyde or ethanol fixation procedures hamper nucleic acid recovery and/or long-term sample storage. Here, we present dithiobis(succinimidyl propionate) (DSP) as a reversible fixative for microbial cells that enables fluorescence imaging and DNA extraction and amplification for sequencing. DSP fixation is compatible with widely used fluorescence in situ hybridization (FISH) approaches and bioorthogonal non-canonical amino acid tagging, and can be applied to diverse sample types, where it consistently performed as well as paraformaldehyde. Furthermore, high-quality genomes were obtained from individual and multiple bacterial cells selectively extracted from complex samples using FISH-FACS. Thus, DSP is a dual-use fixative for microbiome imaging and sequencing, improving targeted microbial genomics.
|
|
Scooped by
mhryu@live.com
Today, 12:07 PM
|
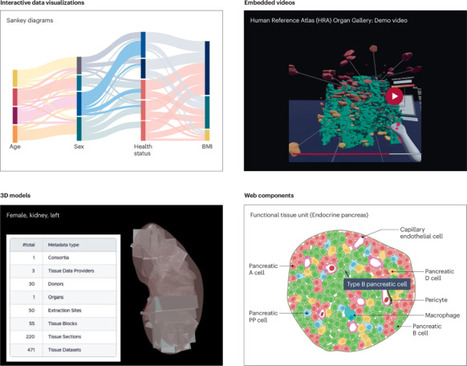
How biologists can make the most of supplementary materials for impact. To overcome the limitations of traditional supplementary items, companion websites can be created with relatively little effort today. These can function as dynamic, rich repositories for additional content, especially figures. They enable the authors to embed interactive visualizations, videos and annotated figures that are impractical to publish in static PDF form. They can also help readers navigate complex datasets and analyses by providing exploratory interfaces. Researchers may already be producing figures suitable for such sites during the research process; interactive plots, animated visualizations or exploratory tools can be valuable when shared alongside the paper.
|
|
Scooped by
mhryu@live.com
Today, 8:46 AM
|
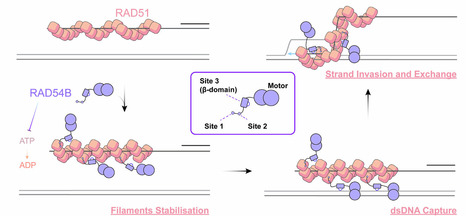
Genome stability is essential for cellular viability yet constantly threatened by DNA-damaging agents. Among these, DNA double-strand breaks (DSBs) are particularly harmful and in S/G2 phases are faithfully repaired through homologous recombination (HR). The RAD51 recombinase forms nucleoprotein filaments on single-stranded DNA (ssDNA) that performs critical functions in HR. The efficiency of HR depends on precise regulation of RAD51 filament dynamics by accessory factors, including RAD54B, a double-strand DNA-dependent ATPase. Here, using cryogenic electron microscopy, mutagenesis, and biochemical assays, we show that RAD54B stabilizes RAD51–DNA filaments, inhibits RAD51 ATPase activity, and promotes D-loop formation and strand exchange. RAD54B contains two main RAD51-interacting sites and a β-domain that bridges RAD51 protomers, is positioned to contact donor dsDNA, and regulates RAD54B’s ATPase activity. Cellular assays reveal that the RAD51-interacting sites are required for repairing DSBs by HR in human cells. Our findings uncover a modular architecture and mechanistic framework for RAD54B’s critical role in genome maintenance. RAD54B contains two main RAD51-interacting sites and a β domain that bridges RAD51 protomers, is positioned to contact donor dsDNA, and regulates RAD54B’s ATPase activity. Cellular assays reveal that the RAD51-interacting sites are required for repairing DSBs by HR in human cells
|
|
Scooped by
mhryu@live.com
Today, 1:04 AM
|
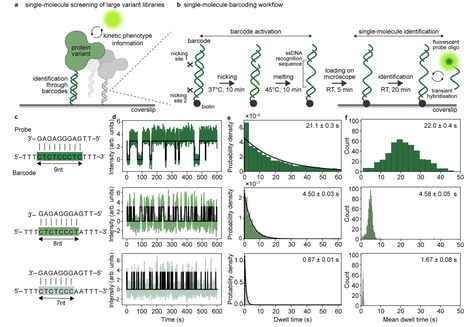
Single-molecule techniques provide exceptional resolution of biomolecular dynamics but are limited by low throughput. Here, we present a DNA barcoding strategy that enables parallel kinetic profiling of multiple protein variants in a single experiment. Each protein is covalently linked to a unique DNA barcode, decoded through transient hybridisation of fluorescent probes distinguished by color and binding duration. Applied to a library of 16 ClpS variants, this approach revealed ~5-fold variation in dissociation kinetics and uncovered previously unrecognised bimodal binding behavior arising from distinct structural subpopulations. By enabling scalable, multiplexed functional characterization at the single-molecule level, this platform provides a general framework for protein engineering, directed evolution and the development of next-generation single-molecule technologies. Single-molecule methods typically analyse only a few protein variants at a time, limiting high-throughput screening. Here, authors develop a DNA barcoding strategy that enables parallel identification and kinetic profiling of protein variants in a single experiment.
|
|
Scooped by
mhryu@live.com
Today, 12:50 AM
|
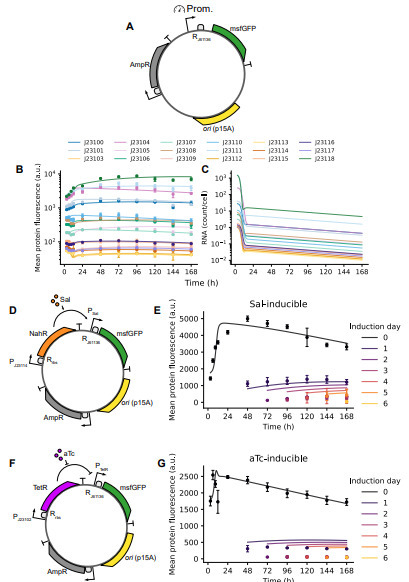
The development of biological engineering tools have improved the ability to embed novel functions into living organisms. Concurrently, our understanding of the interface between synthetic genetic circuits and their host has accelerated in the past decade, enabling the development of context-aware predictive models. However, long-term circuit function remains difficult to predict when designs operate in field-relevant conditions. Specifically, while synthetic genetic circuits are typically characterized during exponential growth, these must operate for extended periods of time in nutrient-limited conditions and where bacteria enter stationary growth phase. In this work, we construct a mathematical model that captures the dynamics of a genetic circuit as cells transition into stationary growth phase. To achieve this, we construct synthetic genetic circuits with fluorescent reporter output and measure plasmid DNA copy number, mRNA level, protein level, and live cell count over the course of one week in stationary growth phase. We find that transcription rates plummet shortly after entering stationary phase but settle at a basal value that depends on both the promoter and ribosome binding site. Translation rates also decline in stationary phase, but protein accumulation due to lack of cell division can increase protein levels well beyond exponential phase. Our model recapitulates these findings and is capable of predicting the output of new genetic circuits. This model can serve as a tool to determine how a genetic circuit design may change on longer time scales where cells enter stationary growth phase.
|
|
Scooped by
mhryu@live.com
Today, 12:02 AM
|
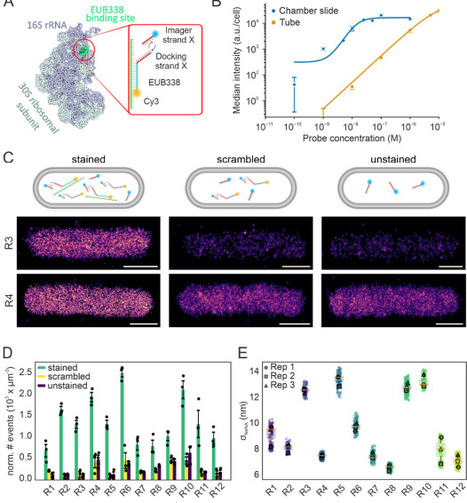
DNA-PAINT enables super-resolution imaging with high multiplexing capacity, yet it is rarely used in nanoscale bacteriology. A likely reason is the lack of straightforward protocols and of systematic characterization of imager-docking strand pairs inside intact bacterial cells, whose envelope and dense cytoplasm differ markedly from both the DNA origami in vitro systems used to develop them, and eukaryotic cells. In this work, we establish a screening platform based on RNA-FISH, in which we extend the 16S rRNA-directed EUB338 probe with docking sequences of choice. This allows imager-docking pairs to be benchmarked directly inside Escherichia coli cells and characterized for parameters such as specificity, binding kinetics, and localization precision. We screened 12 different DNA-PAINT sequences and used an optimized set to perform four-target 3D Exchange-PAINT in single E. coli cells. We further applied our strategy to image three effector mRNAs in the Gram-negative pathogen Yersinia enterocolitica and quantified RNA clusters and their 3D distribution. Our work provides a framework and practical guidelines for high-quality 3D DNA-PAINT imaging in Gram-negative bacteria and should facilitate its broader adoption in nanoscale bacteriology.
|
|
Scooped by
mhryu@live.com
October 12, 11:28 PM
|
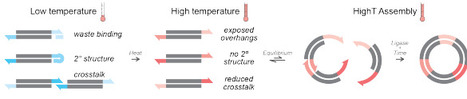
Our ability to efficiently construct synthetic DNA products is essential to studying and engineering biology. There is an abundance of widely adopted DNA assembly technologies suitable for generating synthetic constructs from diverse DNA inputs, but these techniques suffer from low specificity of connection during assembly, which decreases the accuracy, efficiency, and throughput of construction. Here we propose a broadly applicable paradigm for DNA assembly, High Temperature (HighT) assembly, that lowers misconnection rates by ligating complementary single strand DNA overhangs at reaction temperatures significantly higher than their predicted melting temperatures. By expanding the concept of melting temperature to demarcate smaller association fractions of DNA duplexes, we characterize HighT and demonstrate 40 percentage points increase in accuracy, and 4 orders of magnitude increase in yield of transformants when applying the technique to conventional assembly technologies. HighT is then applied to develop cloning pipelines for the development of efficient assembly-to-integration protocols, the construction of specific low-abundance isoforms of medically relevant proteins from human gDNA and oligos, and the de novo construction of a 10 kb, GC rich genomic region from the functionally extinct Northern White Rhinoceros created entirely in vitro.
|
|
Scooped by
mhryu@live.com
October 12, 11:08 PM
|
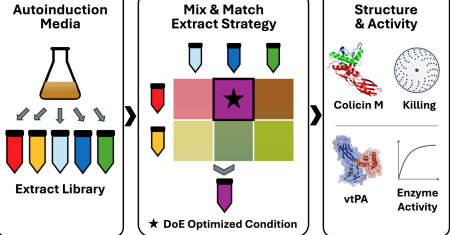
Cell-free protein synthesis (CFPS) is a powerful platform for rapid protein prototyping and biomanufacturing, but expression of complex proteins remains limited by labor-intensive extract preparation as well as the inability of conventional E. coli extracts to efficiently support disulfide bond formation and prevent protein aggregation. We previously developed a Cell-Free Autoinduction (CFAI) workflow that simplifies extract preparation, however, CFAI-derived extracts retain the folding limitations of standard E. coli systems for producing traditionally difficult proteins. Here, we address these challenges by utilizing extracts enriched with molecular chaperones or Erv1p-based oxidative folding catalysts and combining them through a modular extract mixing strategy to enable independent tuning of the cell-free folding environment. This workflow supplies folding factors through enriched extracts, avoiding recurring purchases of purified supplements and enabling a modeled reagent-cost advantage. To evaluate real-world applications and utility of lysate mixing for protein production, we demonstrate a 10-fold increase in solubility of the aggregation-prone bacterial toxin Colicin M, achieving >95% soluble protein while retaining antimicrobial activity. We further used a Design of Experiments (DoE) framework to optimize combinations of chaperone-enriched and Erv1p-based oxidative folding extracts containing either protein disulfide isomerase (PDI) or DsbC. The optimized compositions produced >95% soluble truncated human tissue plasminogen activator (vtPA), comprising the kringle 2 and protease domains, while preserving enzymatic activity. Together, these results demonstrate that CFAI-derived modular extract mixing provides a tunable, high-throughput strategy with low modeled materials costs for tailoring cell-free biochemical environments for difficult-to-express proteins, expanding CFPS utility for protein engineering and biomanufacturing.
|
|
Scooped by
mhryu@live.com
October 12, 10:27 PM
|
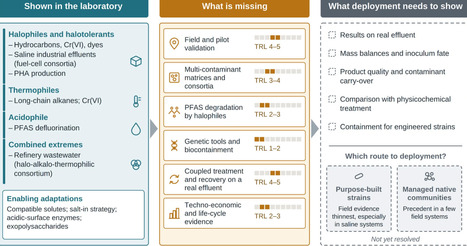
Extremophiles are a recurring answer to a recurring problem in environmental biotechnology: many waste streams most in need of treatment are too saline, hot, acidic, or alkaline for conventional microorganisms to treat effectively. Organisms that not only tolerate but require those conditions can, in principle, act on such streams directly, and their extreme niche doubles as a barrier against contamination, the premise for open, non-sterile processing. The literature cataloging which extremophile degrades which pollutant is now large and, in places, mature. Far less settled is whether any of it works outside the laboratory. This review surveys the demonstrated degradative capabilities of extremophiles and then concentrates on the translational gap separating a capable strain from a deployable process. Six problems recur: the scarcity of field and pilot validation, especially for halophilic and engineered systems; performance on the multi-contaminant, multi-stressor matrices real waste presents, and the open question of whether managed or designed consortia outperform single isolates; activity against recalcitrant contaminants such as per- and polyfluoroalkyl substances (PFAS), where we identified no halophilic reports; immature genetic toolkits that constrain engineering and biocontainment; the little-explored combination of treatment with resource recovery; and the scarcity of techno-economic and life-cycle evidence. For each, we identify the experiments and analyses that would close it, and we distinguish, throughout, between what is genuinely absent from the literature and what may simply be underreported.
|
|
Scooped by
mhryu@live.com
October 12, 5:47 PM
|
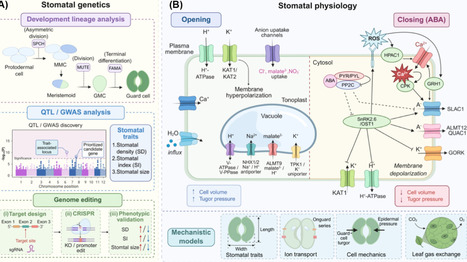
Stomata regulate plant growth and environmental responses through their control of gas exchange and transpiration. However, high-throughput stomatal phenotyping and the integration of phenotypic, genetic, and multiomics data remain major challenges for biological discovery and for understanding stomatal function in plants. Here, we review emerging imaging technologies and foundation models for stomatal biology and discuss how advances in AI could transform plant stomatal research. The rapid accumulation of large-scale phenotypic, genetic, and omics datasets requires systematic integration of heterogeneous data and their interpretation to uncover mechanisms of stomatal regulation within a unified reasoning framework. Therefore, we propose a stomatal phenotyping and analysis agent framework that connects these pipelines to accelerate hypothesis generation, phenotype–genotype analysis, discovery of biological function, and crop improvement.
|
|
|
Scooped by
mhryu@live.com
Today, 2:01 PM
|
Fluorescence-based CRISPR diagnostic assays have become a popular platform for nucleic acid detection due to their programmability, configurability, specificity, and compatibility with standard laboratory equipment. However, reported enzymatic kinetic rates and limits of detection for CRISPR trans-cleavage assays vary by several orders of magnitude across the literature. This variation in performance parameters is coupled with, and exacerbated by, inconsistent calibration, incomplete correction of measurement biases, and nonstandardized or incomplete data analysis procedures. We present an experimental protocol and quantitative analysis framework for fluorescence-based enzyme assays using routine laboratory instrumentation, including thermocyclers and fluorescence microplate readers. Building on previous studies of CRISPR enzyme kinetics and fluorescence calibration, we describe procedures for flat-field and background correction; comprehensive fluorescence calibration, including correction for the inner filter effect; quantification and implications of reporter degradation; extraction of Michaelis–Menten kinetic parameters; and determination of assay's limits of detection. We provide step-by-step experimental guidelines and open-source Python implementations for each stage of the workflow. Using representative Cas12 trans-cleavage datasets, we demonstrate that explicit fluorescence calibration and correction procedures substantially reduce systematic bias in measured kinetic rates and improve consistency between experiments. Our framework aims to establish standardized practices for quantitative fluorescence-based CRISPR assays and provides researchers with practical tools for reproducible kinetic characterization and rational assay design.
|
|
Scooped by
mhryu@live.com
Today, 1:34 PM
|
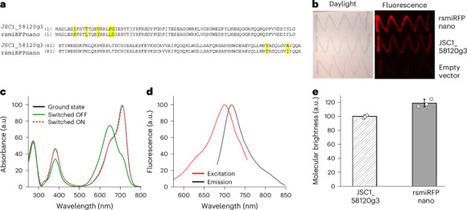
Using near-infrared photoswitchable photochromic genetically encoded probes, differential photoacoustic tomography (PAT) enables deep-tissue imaging with high resolution and sensitivity; however, current bacteriophytochrome-based probes are bulky multidomain proteins exceeding 55 kDa, which constrains their use for designing PAT molecular reporters. Here we engineer rsmiRFPnano, a compact cyanobacteriochrome-derived photoswitchable PAT probe. With a molecular weight below 19 kDa, rsmiRFPnano efficiently incorporates biliverdin, exhibits high extinction coefficients in both photochromic states and demonstrates robust performance in live cells. We established rsmiRFPnano-based reporters for NF-κB and AP-1 pathways, which display low basal activity and strong activation-dependent responses. These reporters effectively visualized acute inflammation using noninvasive PAT in vivo, further validated through cellular-level spatial omics mapping. We then created photoswitchable nanobody probes, rsNIR-Fbs, stabilized upon antigen binding and rapidly degraded otherwise, enabling selective photoacoustic immuno-imaging of intracellular antigens in vivo. Together, these results establish rsmiRFPnano as an ultracompact and versatile platform for molecular PAT. A small, photoswitchable and genetically encodable protein operating in the near-infrared window is reported for photoacoustic tomography applications.
|
|
Scooped by
mhryu@live.com
Today, 12:46 PM
|
With the rapid growth of plastic production, plastic pollution has become increasingly widespread and persistent in the environment. This persistent pollution poses potential risks to ecosystems and human health. Microbial biodegradation offers a promising approach to plastic waste management because it operates under mild reaction conditions and has potential for sustainable application. This review summarizes recent advances in the major microbial groups involved in plastic biodegradation, methods for assessing degradation efficiency, underlying mechanisms, and strategies for improving degradation performance. Assessment methods are discussed, from physicochemical characterization to biochemical validation. Meanwhile, it emphasizes that real plastic biodegradation and degradation efficiency require multiple complementary lines of evidence to judge in a more complete and objective way. The complete process and molecular mechanisms of microbial plastic degradation are then described, including microbial attachment and biofilm formation, enzymatic degradation, intracellular metabolism, and mineralization of degradation products. In addition, it summarizes various efficiency optimization strategies for degradation, including plastic pretreatment, bioaugmentation, and regulation of environmental parameters. The lack of standardized assessment, limited practical translation, and ecological risks are the identified key challenges. Finally, standardized efficiency evaluation strategies, plastic upcycling, and emerging technologies for subsequent development were discussed. This review aims to provide a reference for understanding microbial plastic degradation.
|
|
Scooped by
mhryu@live.com
Today, 12:39 PM
|
Microbial communities are the main drivers of global carbon cycling. Yet, predicting their respiratory responses to warming is challenging because the underlying mechanisms remain elusive. Here, we developed a spatially explicit model, integrating single-cell metabolism with metabolic exchange and endogenous heat production, to disentangle the mechanisms driving community-level temperature responses. We reveal that mutualism amplifies respiratory temperature sensitivity compared to isolated taxa, as exchanged metabolites sustain a secondary growth phase with higher respiratory demand, and this phase creates disproportionately large CO2; emissions under warming (apparent activation energy Ea; increases from 0.65 to 0.77 eV). Spatial structure amplifies this further through threshold effects, whereby diffusion limitation shifts from constraining to enabling mutualistic benefits as metabolic rates increase with temperature (Ea; increases to 1.18 ± 0.09 eV). Endogenous heat production drives additional amplification through positive feedback between local temperature rises and resource turnover rates (Ea; further increasing to 1.78 ± 0.14 eV). Collectively, our results demonstrate that community-level temperature responses are not determined by enzyme kinetics alone but emerge from the interplay of ecological, physical, and metabolic interactions. Our model provides a general framework for understanding the combined effects of the underlying mechanisms on microbial temperature responses. Future climate carbon models should incorporate these considerations to improve predictions.
|
|
Scooped by
mhryu@live.com
Today, 12:05 PM
|
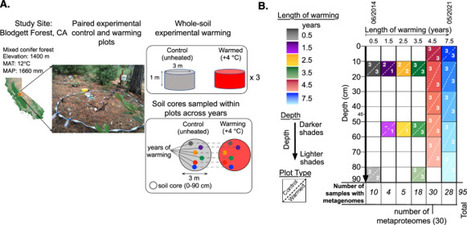
Soils are Earth’s largest carbon reservoir, yet microbial mechanisms controlling decomposition, and their warming response, remain poorly resolved, especially in subsoils. Using genome-resolved proteogenomics within a replicated whole-soil warming experiment (0–90 cm, +4 °C, 7.5 years) we map microbial traits, their depth stratification, and warming responses. Depth is the dominant driver of microbial community and trait structure. Depth-wise, metabolism converge on low-molecular-weight and C1 compounds, challenging glucose-centric models, with surface soils enriched in methanol oxidation and deep soils in alcohol fermentation, CO oxidation and N₂ fixation. Abundant sialidases in deep soils implicate glycoproteins as a key carbon source. Warming effects are smaller and decrease over time but are depth-specific: surface soils show enhanced amino sugar and sugar alcohol transport, phosphorus mobilization, and stress tolerance, whereas mid- and deep soils exhibit stimulated C1 metabolism, CO oxidation, and altered nitrogen turnover. These depth-specific microbial trait shifts may critically influence soil carbon–climate feedbacks. In this study, the authors use proteogenomics of a whole-soil warming experiment to show that soil depth dictates microbial community and trait structure more than soil warming. Deep-soil microbes rely on alternative survival strategies and actively produce enzymes that break down complex glycoproteins.
|
|
Scooped by
mhryu@live.com
Today, 8:39 AM
|
Aminoacyl-tRNA synthetases are the guardians of translational fidelity. Their complex function is mirrored by an elaborate structure, which includes multiple nested domains. While the evolutionary pressures that promoted the emergence of some domains, such as the editing domain, are clear, the pressures acting on other domains, particularly those at the C-terminus, are not. Here, we combine kinetic analysis, X-ray crystallography, and bioinformatics to unveil the history and evolutionary forces that have shaped isoleucyl-tRNA synthetase (IleRS) domain structure. We find that the traditional classification of IleRSs based on the C-terminal tRNA-recognition domains is incomplete, as it fails to capture features of the synthetic domain. Guided by a crystal structure of the Priestia megaterium IleRS2:tRNAIle complex, we removed key interactions between IleRS2 and its cognate tRNA and characterised their impact on enzyme activity. We found that D-loop interactions with the IleRS2 C-terminal domains are non-essential in prokaryotes, and their loss can even increase catalytic turnover. Further, we show that the zinc-binding domain of IleRS1 recognises the anticodon less stringently than the canonical C-terminal domain of IleRS2. Our data suggest that C-terminal evolutionary remodelling of IleRSs is an ongoing process with a historical precedent and is consistent with selection for faster aminoacylation rate. Aminoacyl-tRNA synthetases decipher the genetic code with remarkable speed and accuracy. Here, the authors show how evolution can increase aminoacylation speed by repeatedly reshaping the interactions between an enzyme and its tRNA substrate.
|
|
Scooped by
mhryu@live.com
Today, 12:54 AM
|
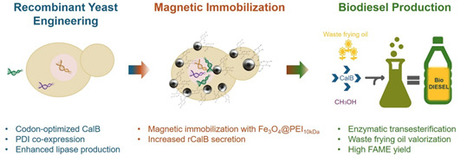
Conventional bio-diesel synthesis is both chemical- and energy-intensive, whereas lipase-catalyzed transesterification provides a more sustainable alternative. Nevertheless, the industrial application of native Candida antarctica lipase B (CalB) is limited by its low catalytic activity and high production costs. To overcome these challenges, we developed a recombinant Komagataella phaffii expression platform to efficiently secrete recombinant CalB (rCalB). To the best of our knowledge, this study demonstrates a proof-of-concept platform that integrates CalB secretion through a dual-promoter expression system regulated by AOX1/GAP in PDI-assisted and magnetically immobilized K. phaffii cells for biodiesel production. This integrated platform enabled extracellular production of rCalB and magnetic recovery of the engineered yeast cells. The maximum biodiesel yield was reached after 96 h using a methanol-to-oil ratio of 4:1 and a catalyst amount of 8 mL (2.4 EU) at 30 °C. The Gas Chromatography Flame Ionization Detector analysis results indicated that the produced biodiesel had higher amounts of C16 and C18 fatty acid methyl esters. These results highlight the potential to produce biodiesel via transesterification catalyzed by rCalB secreted from magnetically immobilized yeast cells.
|
|
Scooped by
mhryu@live.com
Today, 12:31 AM
|
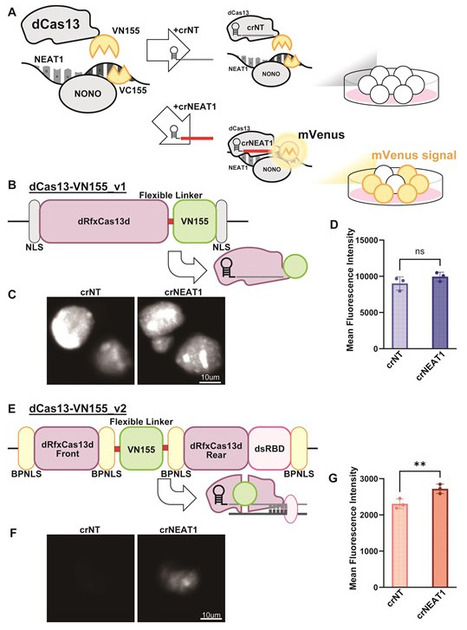
RNA–protein interactions (RPIs) are as important as protein–protein interactions (PPI) for the formation of membraneless organelles (MLOs) and play vital roles in diverse biological processes. Despite remarkable advances in PPI analysis technologies, the development of tools for RPI analysis has lagged behind. To address this challenge, we integrated three established PPI technologies—bimolecular fluorescence complementation (BiFC), NanoBiT, and split-TurboID—with RNA-targeting CRISPR/dCas13. We applied these platforms to paraspeckles, a representative MLO formed through interactions between the long non-coding RNA NEAT1 and the RNA-binding protein NONO. BiFC-dCas13 functions as a fluorescence-based recorder that enables quantitative detection of NEAT1–NONO interaction events. NanoBiT-dCas13 precisely monitors dynamic changes in the NEAT1–NONO interaction through a reversible luminescence readout. Split-TurboID-dCas13 induces proximity-dependent biotinylation of proteins surrounding paraspeckles, identifying the N6-methyladenosine reader YTHDC1 as a paraspeckle-associated protein. Furthermore, BiFC-dCas13 was readily reconfigured to detect additional RPI pairs, demonstrating the modularity of the platform. Together, these complementary systems provide versatile approaches for recording, monitoring, and discovering RPI-associated biology and molecular environments.
|
|
Scooped by
mhryu@live.com
October 12, 11:38 PM
|
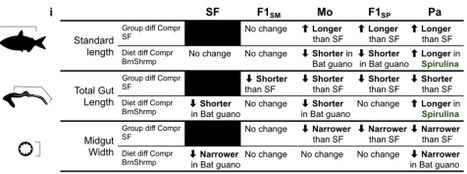
Animals colonizing food-limited environments need to overcome growth constraints by adapting to low, inconsistent nutritional inputs. Comparative studies of surface and cave populations of the Mexican tetra fish species Astyanax mexicanus provide a powerful system to explore evolution to food scarcity. In nutrient-scarce caves, evolved traits such as eye loss and increased fat storage have been shown to conserve energy in cavefish. However, the role of microbiota-mediated metabolism in sustaining growth and survival under food limitation remains poorly understood. Here, we demonstrate that surface and cave populations distinctly modulate their gut microbiota, with cavefish recruiting microbes whose predicted metabolites specifically promote growth under nutrient-limited conditions. Using microbiota data from three distinct dietary regimes and a fasting treatment, integrated topological data analysis and sparse modeling identify sugar and fatty acid degradation pathways coupled with growth in cavefish. Notably, the metabolites of these pathways are beneficial only for cavefish growth, and not for surface fish. Fasting in cavefish led to the recruitment of gut microbes capable of metabolizing aromatic compounds inaccessible to the host, whereas it recruited few microbial associates in surface fish. Our findings indicate that cavefish evolved novel host-microbiota coupling to sustain growth and resilience in their nutrient-scarce niche. We anticipate that our findings will broadly inform studies of evolution under extreme nutritional conditions and provide insights into the role of host-microbiota interactions in resilience to food scarcity at varied time scales.
|
|
Scooped by
mhryu@live.com
October 12, 11:16 PM
|
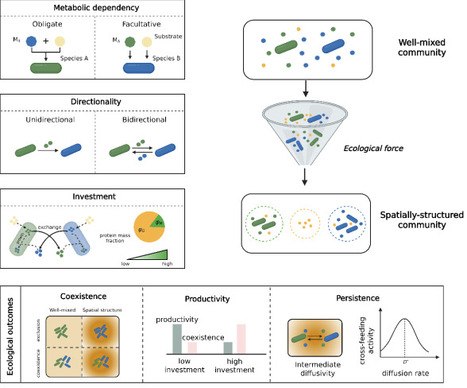
Metabolic cross-feeding is central to microbial life. Theory predicts that spatial structure favors cross-feeding by affording cells preferential access to the exchanged metabolites. Yet recent evidence shows that spatial structure can, in some contexts, disfavor cross-feeding. Here, we resolve this contradiction by structuring cross-feeding interactions into three dimensions: (i) directionality, (ii) investment, and (iii) metabolic dependency. We then investigated their combined effects using a spatially explicit individual-based model. We find that the effect of spatial structure on cross-feeding is determined not by physical proximity alone, but by how microbial growth depends on exchanged metabolites. Metabolic dependence shapes the spatial organization of cells and metabolite fluxes, and this emergent organization, in turn, reshapes the ecological outcome of cross-feeding. Our results reconcile previously conflicting observations and provide a general framework for predicting and engineering metabolically interdependent microbial communities.
|
|
Scooped by
mhryu@live.com
October 12, 11:01 PM
|
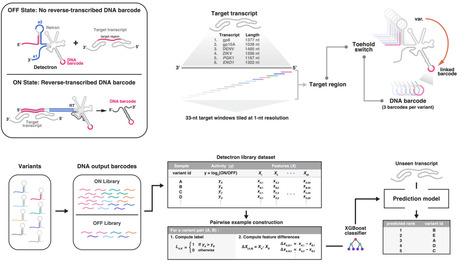
Programmable RNA sensors enable sequence-specific detection of RNA inputs for diagnostics, genetic circuit control, and multiplexed molecular readouts. Although readily programmable, their performance varies substantially with the target site selected within a transcript, making target-site selection a major challenge. Here, we report DeSTAR (Detectron Sensor TARget-site ranker), a machine-learning-assisted target-site ranking model for programmable RNA sensors. To train this model, we generated transcript-wide, single-nucleotide-resolution tiling sensor libraries across six diverse RNA targets, enabling generalization to be tested across transcripts rather than within a single RNA. In leave-one-transcript-out evaluation, DeSTAR's top-ranked sites were enriched for high-performing sensors in held-out transcripts, driven predominantly by interpretable sequence grammar in the sensor stem rather than transcript-specific patterns. Prospectively, on an unseen SARS-CoV-2 transcript, the model's top-ranked sites yielded working sensors roughly five-fold more often than random selection. These results support DeSTAR as a generalizable strategy for target-site selection in RNA sensors across transcripts.
|
|
Scooped by
mhryu@live.com
October 12, 10:22 PM
|
tRNA are short, ~76 nucleotide-long, modification-rich molecules that play an important role in protein translation. Nanopore sequencing is the only platform capable of directly sequencing full-length, native tRNA molecules. This capability is enabled by splinted ligation of tRNA-specific adapters. As a result, the measured ionic current signal reflects a composition of both adapter and tRNA sequence features. Despite these advantages, analysis of nanopore tRNA sequencing data poses distinct computational challenges. Existing ionic current-based signal-to-sequence decoding (basecalling) models are optimized for mRNA and perform suboptimally on short, modification-dense RNA such as tRNA. Furthermore, alignment algorithms for reference-guided analysis of short-length reads are not optimized for nanopore reads with lower accuracy than Next-generation sequencing reads. To overcome these limitations, domain-specific computational approaches are required. We present tRNAZAP, a nanopore ionic current signal analysis toolkit designed to enable precise reference-guided sequence alignment. tRNAZAP operates directly on raw ionic current signals and decomposes the analysis into two complementary tasks: 1) segmentation of adapter and tRNA regions, and 2) tRNA classification, both implemented using transformer-based models. The segmentation component isolates the tRNA signal portion and extracts the corresponding nucleotide sequence as the alignment query, while the classification component identifies the tRNA type as the alignment reference without requiring sequence-level information. Together, these outputs allow direct optimal pairwise alignment between the tRNA subsequence of the full query and reference sequences. Trained on in vitro transcribed RNA data and evaluated on yeast tRNA, these models achieved ~98% classification accuracy and intersection-over-union of ~0.98 for segmentation.
|