 Your new post is loading...

|
Scooped by
mhryu@live.com
Today, 2:55 PM
|
Selective oxidation reactions are critical steps in catalysis towards industrial-relevant compounds. Flavin-dependent enzymes of the VAO/PCMH and GMC families provide numerous candidates that can be applied in various formats and scales. Progresses in catalyst production, mechanistic understanding, structural information, and protein engineering enabled their successful applications. Hence, some candidates are already in use for large-scale chemical syntheses and biosensor applications such as vanillin production, biofuel cells, blood glucose sensing, and glucose monitoring in foods and beverages. The most recent advances related to these oxidases are summarized herein, highlighting recent developments beyond well-known applications that pave the way for novel developments.
|
|
Scooped by
mhryu@live.com
Today, 2:43 PM
|
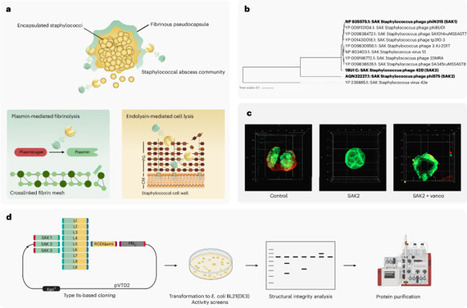
The rise of antibiotic resistance, particularly among pathogens such as methicillin-resistant Staphylococcus aureus, necessitates innovative antibacterial strategies. Here we introduce FibriLysins, a class of engineered enzymes combining staphylokinase and endolysin, which dismantle fibrin-encapsulated multicellular staphylococcal communities, including biofilms and staphylococcal abscess communities (SACs). We characterized diverse staphylokinases, developing a library of 27 FibriLysins, with a 78% positive hit rate. The lead variant, FL-028, has significant antibacterial activity against prevalent clinical S. aureus isolates. In vitro, FL-028 reduced bacterial counts in SACs by 3.4 log and in biofilms by 2.3 log. In vivo, FL-028 exhibits superior efficacy in a mouse model of bone fracture-related infection, outperforming vancomycin by 1 log and eliminating SAC presence in tissues. FL-028 shows no adverse effects on bone healing, as confirmed by histological examination, mechanical testing and micro-computed tomography analyses. These preclinical findings highlight the potential of FibriLysins as a robust and safe therapeutic approach against difficult-to-treat staphylococcal infections. Engineered FibriLysins dismantle fibrin-encapsulated Staphylococcus aureus communities and outperform vancomycin in a preclinical model of fracture-related infection.
|
|
Scooped by
mhryu@live.com
Today, 2:01 PM
|
Scoring functions built with advanced deep learning algorithms have recently gained prominence in protein–ligand pose generation and binding affinity prediction for computational drug design. However, several shortcomings make it increasingly difficult to evaluate scoring functions for a specific task across studies. Transparent disclosure of training data, code and procedures is essential to permit reproducibility, reuse and independent scrutiny. Guarding against data leakage, sampling bias and flawed design is equally important. Meaningful comparisons of scoring functions also require common data splits, checks for run-to-run variability and performance baselines. Here we propose a set of guidelines to help the community identify the most promising methods for addressing these important problems. Ballester and Tran-Nguyen advocate for transparency in data, code and experimental procedures in the evaluation of protein–ligand predictions. They discuss good practices for preventing data leakage and bias, enabling fair, reproducible and meaningful comparisons across different scoring methods.
|
|
Scooped by
mhryu@live.com
Today, 1:44 PM
|
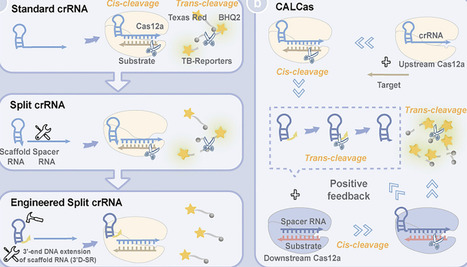
Amplification-free CRISPR detection has attracted significant attention for its capacity to circumvent the intricacies of nucleic acid amplification. Yet, the persistent challenge of high background signals remains a critical bottleneck, motivating intensive efforts toward background suppression strategies. Here, we present a 3’-end DNA extension of scaffold RNA (3’D-SR) conformational switch to drive the cascade-activatable and leakage-suppressed Cas12a system, a mechanism we denote as CALCas. By engineering the crRNA of Cas12a system and introducing steric hindrance as an activation barrier, we reshape crRNA into a “threshold-controlled” smart molecular switch, enabling Cas12a to shift from “inhibition” to “precise activation”. Unlike other Cas cascade reaction generally suffer from internal self-activation that leads to uncontrolled background increase, this newly developed framework, leveraging a DNA-mediated allosteric regulation mechanism and a leakage-suppressed design based on inherent steric hindrance, effectively solves the dilemma between signal enlargement and minor background variation. Through rational design, this cascaded CRISPR system achieves femtomolar sensitivity (over 10,000-fold higher than single Cas12a system), enables rapid HPV16/18 detection, and reduces assay time to under 30 min. This engineered crRNA approach unlocks a new pathway to reprogram Cas12a's enzymatic activities with a stabilized benchmark signal that can be critical in precise quantitative nucleic acid detection.
|
|
Scooped by
mhryu@live.com
Today, 12:42 PM
|
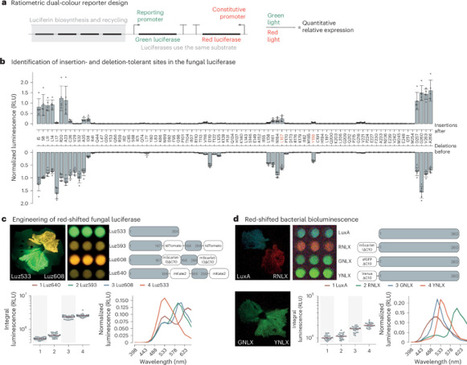
Self-sustained luminescence enables non-invasive monitoring of molecular events at the whole-organism level, but its intensity depends on both gene expression and cellular metabolism, limiting quantitative interpretation. Here we present an approach that disentangles the contributions of gene expression and metabolism. By co-expressing an engineered reference luciferase variant that uses the same substrate but emits light at a different wavelength and monitoring the color ratio, we substantially reduced signal variability, enabling quantitative physiological imaging despite changes in metabolic activity. In glowing plants, monitoring colour shifts instead of changes in luminescence intensity enables quantitative measurements of physiology despite changes in metabolic activity.
|
|
Scooped by
mhryu@live.com
October 22, 11:01 PM
|
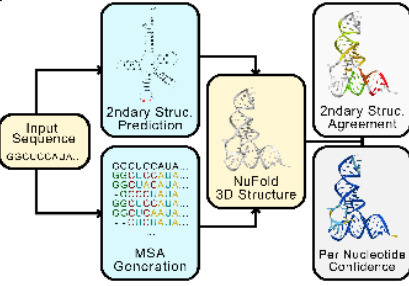
To bridge the gap between rapid RNA sequence discovery and slow experimental characterization, we present the NuFold RNA Structure Database https://nufold.kiharalab.org/ a resource providing large-scale, precomputed full-atom RNA tertiary structures. By removing the computational barriers associated with running the method locally, this database makes high-accuracy RNA structure models readily accessible to the biological research community for a wide range of applications.All predicted structures and corresponding data are available at https://nufold.kiharalab.org/. The source code for the NuFold program is available at https://github.com/kiharalab/NuFold.
|
|
Scooped by
mhryu@live.com
October 22, 10:48 PM
|
Intestinal colonization by Klebsiella pneumoniae is recognized as a pivotal prerequisite for its systemic dissemination and subsequent invasive infection; however, the metabolic basis and regulatory mechanisms underlying this process remain poorly understood. In this study, we employed transposon insertion sequencing (Tn-seq) to systematically identify metabolic genes associated with intestinal colonization in K. pneumoniae. By integrating high-throughput screening with in vivo phenotypic validation, we identified the global transcriptional regulator DksA as a key factor that markedly enhances K. pneumoniae intestinal colonization. Mechanistically, DksA may contribute to intestinal colonization through modulation of central carbon metabolism. Integrated transcriptomic and metabolomic analyses, together with homology modeling and molecular dynamics simulations, suggested that DksA may contribute to allosteric remodeling of key RNA polymerase structural domains, thereby influencing the expression of metabolic genes, carbon source utilization, and intestinal colonization by K. pneumoniae. Collectively, our findings highlight the important role of metabolic plasticity associated with transcriptional regulation in K. pneumoniae intestinal colonization and provide a theoretical framework and potential targets for developing anti-colonization strategies that exploit pathogen metabolic vulnerabilities.
|
|
Scooped by
mhryu@live.com
October 22, 4:28 PM
|
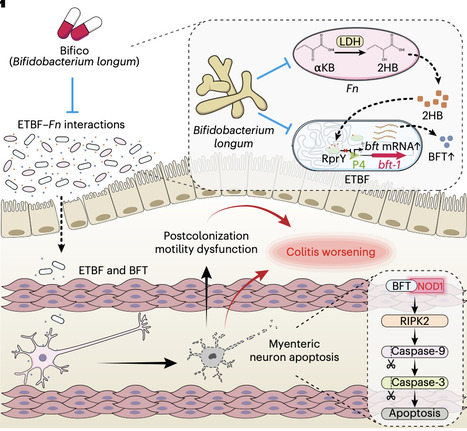
Gut dysmotility is a common long-term effect of enterotoxigenic Bacteroides fragilis (ETBF) colonization but the underlying mechanisms are not clear. Here, using mouse colonization models, we found that ETBF-secreted B. fragilis toxin (BFT) causes loss of colonic myenteric neurons, which are crucial regulators of intestinal motility. BFT induced apoptosis in these neurons through the NOD1–RIPK2–CASP9–CASP3 pathway, leading to gut dysmotility that persisted even after the clearance of ETBF. Analyses of human inflammatory bowel disease and colorectal cancer patient cohorts showed a positive correlation between ETBF and Fusobacterium nucleatum abundance. In mice, F. nucleatum-derived 2-hydroxybutyric acid enhanced BFT production in ETBF by alleviating RprY-mediated suppression and aggravated neurotoxicity and dysmotility. This enteric neuron loss and dysmotility enhanced susceptibility to dextran sulfate sodium-induced colitis in mice. Therapeutically, a Bifidobacterium-based probiotic attenuated neuron toxicity and improved intestinal motility. Our findings reveal bacterial interactions regulating intestinal function and a potential clinical therapeutic approach to preventing outcomes of ETBF colonization. Fusobacterium nucleatum promotes toxin expression in enterotoxigenic Bacteroides fragilis that induces apoptosis in enteric neurons. This eventually leads to gut dysmotility that persists even after bacterial clearance but can be alleviated by using a probiotic cocktail.
|
|
Scooped by
mhryu@live.com
October 22, 12:00 PM
|
Eukaryotic-like proteins (ELP) are often found in intracellular symbionts of insects, plants and evolutionary early-branching sponges, however their functions remain largely uncharacterised. Here we show that the recombinant expression of single genes of various ELP classes can confer on free-living cyanobacteria the capacity to escape phagocytosis by sponges and thus enable intracellular survival. The findings establish ELPs as molecular factors that can control sponge-microbe interactions, which has implications for the evolution of microbial symbiosis.
|
|
Scooped by
mhryu@live.com
October 22, 11:54 AM
|
Over the last decade, there has been a shift in biotechnology from genome reading (sequencing) to genome writing (engineering). While significant progress has been made in prokaryotes and yeast, genome writing remains in its infancy in plants. Unlike their complex nuclear genomes, the compact and biotechnologically attractive plastid genome represents an ideal target to launch plant genome writing at scale.
|
|
Scooped by
mhryu@live.com
October 22, 11:48 AM
|
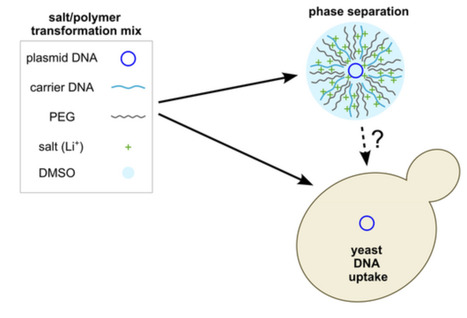
Yeasts are model organisms and protein production hosts of choice, in part due to their ease of genetic manipulation. Yeast transformation with DNA on plasmids or integrated into the genome has been developed since the 1970s and is a routine method in many laboratories. However, transformation frequencies are surprisingly low, with common methods yielding around 1% of transformed cells in budding yeast Saccharomyces cerevisiae. In alternative species, there is lower transformation frequency and unreasonably large quantities of input DNA may be required. Surprisingly, the mechanisms by which yeast cells take up DNA are not known. Here we argue that three approaches could deliver a step-change in yeast transformation frequency and efficiency. First, investigating the mechanisms of DNA uptake. The requirement for salts and extraneous polymers—carrier DNA and PEG—in standard protocols suggests that phase separation of DNA may enhance DNA uptake into cells or subsequent nuclear delivery. Second, accounting for the physiological or growth state of the yeast including cell cycle phase and nutrient or stress conditions. Third, systematically determining the factors that affect transformation efficiency using statistical Design of Experiments. These approaches could allow superefficient transformation of S. cerevisiae, and accelerate genetic manipulation of non-model yeasts.
|
|
Scooped by
mhryu@live.com
October 22, 11:26 AM
|
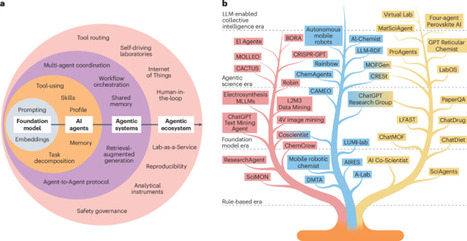
Artificial intelligence (AI) agents are emerging in scientific research as systems that can transform the accumulated decisions of scientists into traceable workflows. In fact, modern scientific practice depends on continual coordination of knowledge dispersed across literature, data, theory and experiment, which, however, is often made through manual, case-by-case judgement. This Primer introduces agentic AI systems as a general method to structure this process through natural language interfaces, serving as a computational brain in the laboratory. Central to this framework are practical workflows for in-depth literature search and knowledge mining, orchestration of physics-based simulations, and integration with automated experimental platforms. We begin by describing the foundations of AI agents, including task decomposition, memory, tool use and communication between agents, and by distinguishing single-agent and multi-agent designs, as well as fully autonomous and human-in-the-loop configurations. Equally important are the means by which these systems are evaluated, documented and reported reproducibly. Finally, we examine major bottlenecks, including hallucination, tool misuse, limited interpretability and safety risks, and outline a future in which coordinated networks of agents, instruments and researchers transform scientific discovery from isolated tasks into continuous adaptive experimentation. Agentic artificial intelligence (AI) systems enhance scientific workflows by structuring knowledge coordination through natural language interfaces. In this Primer, Feng et al. examine the challenges and opportunities involved in integrating AI agents in the laboratory.
|
|
Scooped by
mhryu@live.com
October 22, 11:05 AM
|
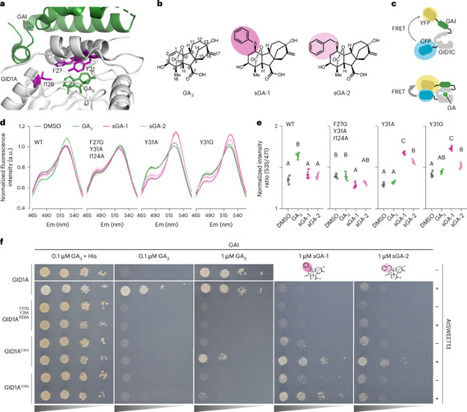
Gibberellins (GAs) influence cell division and elongation, profoundly shaping plant architecture and yield. GA perception occurs when bioactive GAs bind the receptor GID1, promoting DELLA degradation and activating transcriptional programs. While GA signalling in the root endodermis is essential for promoting root elongation, the contributions of other layers have remained unclear. Here we developed a synthetic GA (sGA) that exhibits little or no binding to endogenous GID1, together with a modified GID1 (mGID1) engineered to selectively recognize sGA, enabling cell-specific activation of GA signalling in vivo. Using this system in Arabidopsis, we demonstrate that coordinated action of GA signalling in the endodermis and epidermis is required for full root elongation. Moreover, the distal spatial distribution of GA signalling and GA biosynthetic enzymes supports a model in which bioactive GAs are transported from GA3ox-expressing tissues to the endodermis and epidermis to coordinate root elongation. The sGA–mGID1 system provides a versatile platform for spatially precise reprogramming of hormone signalling, enabling synthetic control of developmental processes such as root–shoot growth balance, thereby advancing applications in plant synthetic biology and sustainable crop improvement. A synthetic gibberellin (GA)-receptor system enables spatial and temporal control of GA signalling, revealing that coordinated GA signalling in the endodermis and epidermis is required for full root elongation and providing a tool for resolving hormone action.
|
|
|
Scooped by
mhryu@live.com
Today, 2:52 PM
|
In this perspective, we discuss the evolving landscape of next-generation probiotics (NGPs), challenging the ambiguity of the term and proposing a framework for unifying the concepts associated with the field. We highlight the potential of novel candidates, including lactic acid bacteria and other commensal microorganisms from untapped taxa and genetically engineered strains, while examining their applications for human health. Rather than defining NGPs as a distinct category, we propose using the term to describe microbial-based health-promoting products in development, particularly where novelty relates to the organism, mechanism, engineering strategy, formulation, or targeted function. We address their safety concerns and regulatory challenges, while clarifying the divergence between therapeutic and dietary supplement pathways. We argue that as these products become more fully characterized and their safety, efficacy, intended use, and regulatory pathway are established, more specific terminology may take precedence, including the long-accepted probiotic category once efficacy and safety are established, or the live biotherapeutic products category if they are specifically developed for therapeutic applications. We suggest that the future of the ‘beneficial microbes for health’ field depends on striking a balance between the benefits of innovation and the necessity of rigorous safety and efficacy considerations.
|
|
Scooped by
mhryu@live.com
Today, 2:02 PM
|
Replication of low-copy-number iteron plasmids is tightly controlled by initiator proteins that coordinate replication initiation with transcriptional autoregulation. In the F-plasmid system, RepE fulfills both functions through a balance between monomeric and dimeric forms. Here, random mutagenesis of the repE gene combined with high-stringency selection was used to identify variants that modulate plasmid copy number. Two point mutations, A78S and Q45L, were identified and shown by qPCR and ddPCR to increase the copy number of plasmid pFO1 relative to the parental repE(R118P) allele. The mutations exhibited growth-phase-dependent effects, with A78S having a stronger impact during exponential growth and Q45L during stationary phase. A double mutant showed a cumulative effect, particularly in exponential phase. RepE transcription increased in the presence of pFO1 and correlated with plasmid copy number, supporting a titration-based regulatory model. These findings demonstrate that engineering RepE variants enables tunable control of plasmid copy number, offering a useful strategy for optimizing gene dosage in synthetic biology and biotechnological applications.
|
|
Scooped by
mhryu@live.com
Today, 1:47 PM
|
Efficient extracellular secretion remains a major bottleneck in microbial cell factories, motivating the search for chassis organisms with native high-throughput transport capacity. Klebsiella quasipneumoniae (Kqp), a recently delineated member of the Klebsiella pneumoniae species complex, encodes diverse secretion-related systems, including RND-type efflux pumps, type II and type VI secretion systems (T2SS/T6SS), outer membrane vesicle (OMV) pathways, and capsular polysaccharide (CPS) export machinery. In clinical settings, these systems contribute to opportunistic pathogenicity, antimicrobial resistance, and host adaptation; in biotechnology, they may provide transport modules that can be detoxified and redirected for extracellular production. This review examines the feasibility of reprogramming Kqp from a pathogen-related bacterium into a safe industrial secretion chassis. We summarize the molecular architecture, substrate-recognition logic, and regulatory networks of major secretion pathways, then discuss CRISPR-Cas-driven removal of virulence and resistance determinants, biocontainment, and synthetic-biology strategies for reallocating secretory capacity. We further survey potential applications in bulk chemicals, human milk oligosaccharides, recombinant proteins, and industrial enzymes. Finally, we identify key barriers to industrial translation, including growth-secretion energetic trade-offs, periplasmic protein misfolding, endotoxin-associated downstream processing, OMV heterogeneity, scale-up robustness, and quantitative safety validation. Establishing Kqp as an industrial chassis will require evidence-tiered strain selection, quantitative secretion-capacity measurement, regulatory decoupling of resistance-associated pumps, and One Health-aligned safety assessment. Direct evidence in K. quasipneumoniae remains limited; therefore, this review distinguishes direct Kqp observations from studies in other KpSC members or K. oxytoca, evidence from non-Klebsiella hosts, and forward-looking proposals. OMV yield is considered separately from lipid A/endotoxin-related properties.
|
|
Scooped by
mhryu@live.com
Today, 1:29 PM
|
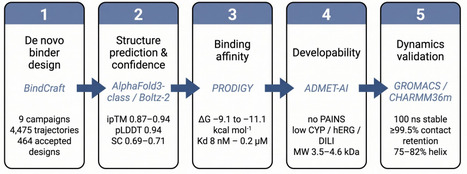
UnibindMD is an integrated computational platform that turns a protein target into validated de novo peptide-binder leads by chaining five orthogonal stages: deep-learning binder design (BindCraft), independent structure prediction and confidence assessment (AlphaFold3-class / Boltz-2), binding-affinity prediction (PRODIGY), developability profiling (ADMET-AI), and all-atom molecular-dynamics validation (GROMACS/CHARMM36m). The platform targets a persistent weakness of single-method design, namely that confidence metrics such as ipTM and pLDDT are self-consistent but not independent, so a design predicted to be confident may still be physically unstable. We demonstrate UnibindMD on three Andes virus (ANDV) intervention points, a setting of urgent need given a case fatality rate approaching 40%, a 2026 MV Hondius cruise-ship outbreak (13 cases, 3 deaths), and no approved therapeutic: the Gn-Gc glycoprotein complex (viral entry), the L-protein endonuclease domain (cap-snatching; PDB 6Q99), and the host VEGFR2 kinase (vascular permeability; PDB 2XIR). Across nine BindCraft campaigns (4,475 trajectories) the platform produced 464 accepted binders (acceptance 2.3-67.6%), with top candidates reaching ipTM = 0.94 and pLDDT = 0.94. The three selected leads were predicted by PRODIGY to bind with G = -9.1 to -11.1 kcal mol-1 (Kd 8 nM-0.2 μM), and all three remained stably bound over 100 ns of unrestrained molecular dynamics (receptor CαRMSD ≤ 2.6 Å, retained peptide fold, ≥ 99.5% of frames preserving more than half of the initial interface contacts, and 75-82% helical content). ADMET-AI flagged the expected poor oral bioavailability of large polar peptides and no PAINS alerts. By converting single-prediction design into a cross-validated, dynamics-tested workflow, UnibindMD provides a rapid, scalable, and generalizable path from any protein target to validated peptide-binder candidates.
|
|
Scooped by
mhryu@live.com
October 22, 11:12 PM
|
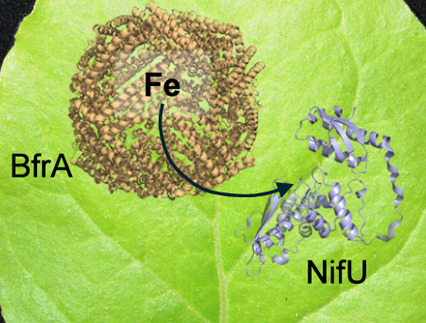
Generating crops that utilize atmospheric nitrogen rather than soil nitrogen requires the transfer of nitrogenase structural genes together with genes required for the biosynthesis of nitrogenase iron−sulfur cofactors. Scaffold proteins, such as NifU, are the starting point of nitrogenase cofactor assembly. While NifU has been successfully produced in plants, its optimal production was achieved with iron supplementation in the nutrient solution. This suggests a faulty connection with the endogenous iron trafficking network, indicating the need for specific elements targeting iron to NifU. Here, we identify bacterioferritin A (BfrA) as an iron donor to NifU. Co-expression with BfrA increases NifU production in Nicotiana benthamiana, leading to doubling the production of NifH, a nitrogenase structural protein that is also required for the synthesis of more complex nitrogenase cofactors. This work underscores the importance of transferring metal delivery systems alongside metalloproteins when engineering heterologous systems.
|
|
Scooped by
mhryu@live.com
October 22, 10:58 PM
|
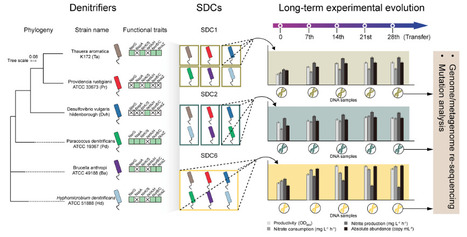
Understanding how interspecific interactions shape ecosystem functioning is a fundamental and longstanding question in ecology, especially along an evolutionary trajectory. Here, we used experimental evolution to evaluate how microbial interactions and adaptive mutations shaped community-level functions in synthetic denitrifying communities (SDCs) along a species richness gradient (one, two, and six species). Our results showed that competitive microbial interactions were observed and significantly (P < 0.05) enhanced denitrification rates in the six-species SDCs during experimental evolution. Genome and metagenome re-sequencing analysis revealed the highest fixed non-synonymous and frameshift mutations within the six-species SDCs, among genes involved in nutrient transport, DNA repair, and secretion pathways. AlphaFold2 protein structure prediction, molecular docking, and Evo 2 host fitness prediction indicated that the fixed mutations could improve host strain fitness, and partial least squares path modeling (PLS-PM) further revealed that interspecific competition accelerated adaptive mutations, thereby reinforcing community-level functions, indicating potential Red Queen dynamics. This study offers novel insights into our understanding of eco-evolutionary mechanisms linking species richness, microbial interactions, adaptive mutations, and community functions, with important implications for active nitrogen removal and greenhouse gas mitigation in diverse ecosystems.
|
|
Scooped by
mhryu@live.com
October 22, 4:34 PM
|
Prokaryotic genomes encode a vast repertoire of noncoding elements such as ribozymes, riboswitches, and CRISPR RNAs. The sequence-level diversity of these intergenic elements precludes alignment-based clustering that would enable annotation and discovery of novel noncoding RNA families. Here, we show that a genomic language model fine-tuned on prokaryotic intergenic sequences learns representations of noncoding elements from DNA sequence. We used these embeddings to organize 376 million intergenic regions from 1.3 terabase pairs of prokaryotic genomes and metagenomes into 5.14 million clusters. This approach nominated 20,149 clusters of putative structured RNAs with a candidate protein partner. Periodicity in token entropies revealed internal architecture, recovering CRISPR, TIGR and VIPR arrays together with their cognate effector proteins from sequence alone. Targeted clustering of bridge recombinase-adjacent sequences identified bridge RNA variants with longer guide recognition sequences than any characterized ortholog. Finally, we directed teams of AI agents to mine the database under researcher-defined criteria, uncovering divergent riboswitches, ribozymes, and other structured RNAs with biological hypotheses for their function. We release a publicly available database that transforms the dark matter of prokaryotic genomes into an explorable resource for human-agent molecular discovery.
|
|
Scooped by
mhryu@live.com
October 22, 12:44 PM
|
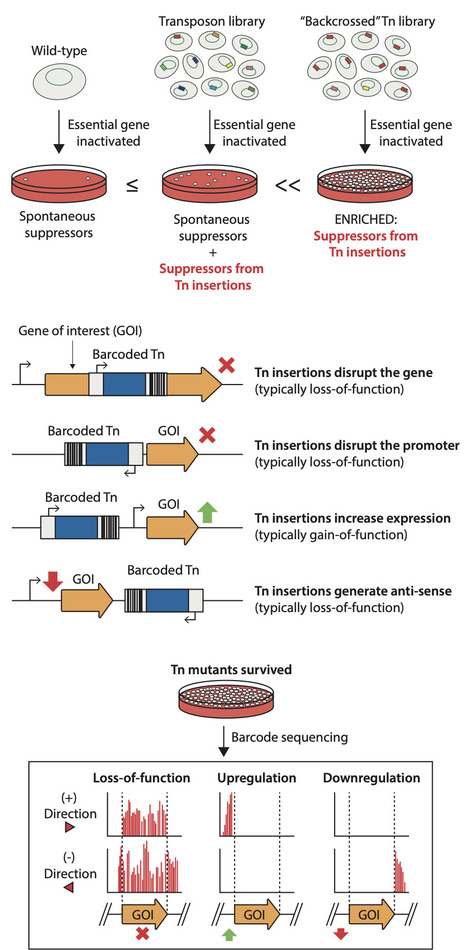
Historically, suppressor analysis has been a powerful tool for uncovering the functions of essential genes. In this study, we report Sup-seq, a transposon-sequencing technique with random DNA barcodes that enables genome-wide detection of genetic suppression. When applied to seven essential genes in Streptococcus pneumoniae, Sup-seq revealed 53 suppressors across various cellular pathways. These suppressors function through diverse mechanisms, such as gene inactivation, overexpression, or downregulation, depending on the locations and orientations of the transposon insertions. Our proof-of-concept experiments identified a previously uncharacterized transcriptional regulator that controls chaperone and ribosomal protein expression, as well as a mechanism that bypasses the requirement for acetate kinase for growth. Furthermore, we found that phosphatidylglycerol is conditionally essential in pneumococcus, revealing an unexpected link between phospholipid synthesis and potassium transport. These findings demonstrate that Sup-seq is a facile approach for providing mechanistic insights into the functions of essential genes. Additionally, Sup-seq can be adapted to other bacteria, aiding in the identification of new antimicrobial targets. Here the authors present ‘Sup-seq’, a transposon-based sequencing technique that enables genome-wide detection of genetic suppression in bacteria. They use it to identify 53 genetic suppressors of seven essential genes in pneumococcus, revealing new links among metabolism, ribosome assembly, and membrane lipid synthesis.
|
|
Scooped by
mhryu@live.com
October 22, 11:58 AM
|
Bacillus subtilis spore surface display has been widely used to immobilize single recombinant enzymes, yet many industrially relevant reactions are multi-enzyme cascades. This study examined whether two distinct, catalytically active enzymes can be co-displayed on a single spore, using the two-step hydrolysis of carboxymethyl cellulose (CMC) to glucose as a model cascade. An endoglucanase (EglS) and a beta-glucosidase (Unbgl1A) were expressed in E. coli, characterised, and surface-displayed on Bacillus subtilis WB800N by fusion to the coat anchors CotB, CotC and CotG. The best constructs were combined either by mixing two single-enzyme spore strains or by co-displaying both enzymes on one strain, and compared with the free-enzyme mixture on CMC. CotG was the most effective anchor for EglS and the only functional anchor for Unbgl1A, with improved thermal stability relative to the free enzymes and activity retained over repeated reuse: CotG-EglS retained approximately 46% activity after 12 h (versus approximately 27% for the free enzyme), and CotG-Unbgl1A retained approximately 30% activity after 8 h, whereas free Unbgl1A lost all activity within 6 h. The mixed single-enzyme spores gave the highest glucose yield (2.15 mM), exceeding both the single dual-enzyme strain (1.57 mM) and the free-enzyme mix (1.35 mM). Two functional enzymes were co-displayed on one spore, but mixing separate single-enzyme spores gave higher yield under the tested conditions and allowed the enzyme ratio to be tuned.
|
|
Scooped by
mhryu@live.com
October 22, 11:51 AM
|
Environmental monitoring of contaminated freshwater systems still lacks curated genomic resources tailored to bioremediation-oriented screening. In this study, we developed a curated catalogue of 546 genes from 80 microbial species associated with heavy-metal resistance, hydrocarbon degradation, detoxification and related functions and applied it to river metagenomes from four municipalities in Pará, Brazilian Amazon: Altamira, Bragança, Marabá and Paragominas. Functional annotation was performed using eggNOG-mapper and metagenomic reads were aligned against the curated catalogue with Bowtie2. Mercury- and copper-related genes were the most frequently represented functional categories across all river systems, a pattern that may reflect both sequence representation in the metagenomes and the unequal composition of the curated reference catalogue. Bragança showed the largest detected gene repertoire, whereas diversity, Jaccard, UpSet and Dunn-based analyses revealed marked spatial heterogeneity and sites of localized functional enrichment. Recurrently associated bacterial taxa included Acinetobacter, Enterobacter, Pseudomonas, Stenotrophomonas, Cupriavidus and Sphingopyxis. These findings indicate that the catalogue provides a targeted genomic reference for environmental screening and bioprospecting, although metagenomic detection represents functional potential rather than experimentally confirmed remediation activity.
|
|
Scooped by
mhryu@live.com
October 22, 11:46 AM
|
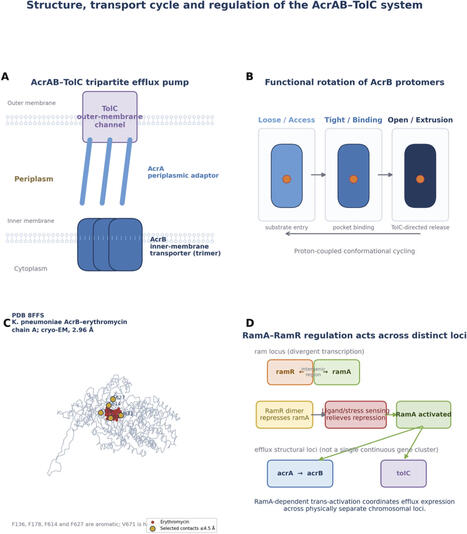
Lipopolysaccharide (LPS) is well known for its immunostimulatory activity and its contribution to the permeability barrier of the Gram-negative outer membrane. Yet LPS is essential for growth in nearly all Gram-negative bacteria, even under laboratory conditions where its barrier function should be less critical. LPS contributes to envelope mechanics, membrane biogenesis, and cell division, but whether any of these functions accounts for its essentiality remains unclear. Here, we systematically tested these possibilities by depleting LPS in E. coli and Klebsiella pneumoniae. Osmoprotection largely preserved cell-envelope integrity but not viability, demonstrating that the mechanical contribution of LPS alone cannot explain its essentiality. Genetic perturbations that increase peptidoglycan rigidity or alter phospholipid homeostasis partially improved growth, supporting independent roles for LPS in envelope mechanics and membrane biogenesis. An unbiased suppressor screen further identified activation of the MarA regulon as a potent mechanism for surviving LPS depletion. Protection required transport by the AcrAB-TolC multidrug efflux pump, revealing an unexpected connection between efflux and cell-envelope homeostasis. Thus, LPS essentiality emerges from multiple non-redundant contributions to Gram-negative cell-envelope physiology.
|
|
Scooped by
mhryu@live.com
October 22, 11:15 AM
|
The coral microbiome is believed to play a crucial role in sustaining corals and enabling adaptation to perturbations. A ubiquitous group of coral-associated bacteria, known as Endozoicomonas, are hypothesized to provide essential metabolites to corals, although detailed dynamics of the interactions between host and microbe are poorly understood. Here multi-omics and laboratory validation experiments reveal that Endozoicomonas encodes genes for the metabolism of coral-derived testosterone and converts ubiquitous cholesterol into testosterone and progesterone, both important steroids that accumulate within the coral holobiont during thermal stress. Field sampling along a natural temperature gradient showed general suppression of coral steroid hormones under thermal stress and during white syndrome manifestation. Provision of testosterone or progesterone to corals during laboratory thermal stress induced transcriptional reprogramming of the coral innate immune response, suggesting microbial involvement in host immune modulation. These findings highlight a previously unrecognized potential mechanism of microbial contribution to coral resilience. Multi-omics and laboratory experiments reveal that a coral-associated bacterium can produce testosterone and progesterone; these two hormones alter the coral host’s innate immune responses during thermal stress.
|







suel gm, During biofilm development, cells inside the biofilm differentiate into two mutually exclusive cell types: motile cells and matrix cells. Motile cells are known to produce γ-PGA. Matrix cells are known to produce various ECM components that hold the biofilm together.