

Single-stranded DNA with hundreds to thousands of bases is a critical reagent in biotechnology. While traditional chemical synthesis is not suitable for efficient production of such long oligonucleotides, M13 bacteriophage offers a scalable pathway towards custom, long, single-stranded DNA production. However, conventional helper plasmid-phagemid systems suffer from batch-to-batch variability and off-target packaging. To address these limitations, we previously engineered eScaf, an E. coli strain with the M13 genome stably integrated into its chromosome. eScaf produces circular, single-stranded DNA with high purity, but yields of only 100 and 200 μg of DNA per liter of culture, too low for cost-effective scale-up. Here, we systematically investigate the genomic and environmental determinants of cssDNA yield in eScaf, revealing that cssDNA yield depends on both the number and chromosomal loci of integrated M13 genomes. Combining these insights with optimized media, we achieve ~4 mg/L of cssDNA in shake flasks, a 40-fold improvement. We also evaluate fed-batch bioreactor feeding strategies for cssDNA production using eScaf. Our new strain, eScaf-IS3, enables cost-effective production of cssDNA for high-throughput therapeutic screening, operon-length knock-in donor production, and large-scale DNA origami assembly.

RMH
100.8K views |
+74 today

 Your new post is loading...
 Scoop.it! 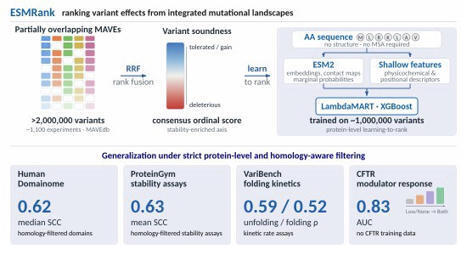
Predicting the functional effects of missense variants is limited both by predictor performance and by the lack of principled strategies to integrate heterogeneous multiplexed assays of variant effect (MAVEs), including deep mutational scanning, into transferable sequence-function models. Here, we show that redundancy among partially overlapping deep mutational scanning experiments, often treated as noise, yields a consensus landscape of mutational constraint. We introduce “variant soundness”, an overlap-aware rank-fusion score that uses cross-experiment consistency to derive an assay-agnostic ordering of mutational tolerance within proteins. Applied to ~1100 MAVEs spanning over two million missense variants, this approach reveals a mutational landscape enriched for structural stability constraints. Guided by this integrated signal, we develop ESMRank, a sequence-based learning-to-rank predictor combining ESM2 representations with physicochemical descriptors. Under strict protein-level partitioning, ESMRank outperforms widely used predictors across abundance-, folding-, and stability-related assays in ProteinGym and Domainome. Predicted deleterious variants are enriched for ClinVar pathogenicity and, in CFTR, correlate with folding efficiency, channel activity, and pharmacological responsiveness. Together, these results turn assay redundancy into a scalable resource for interpretable missense variant prediction.
mhryu@live.com's insight:
ESMRank takes a protein sequence and ranks all possible single amino acid substitutions by predicted tolerance, with predicted loss of function variants at the bottom. Training was dominated by human proteins 
Scoop.it!
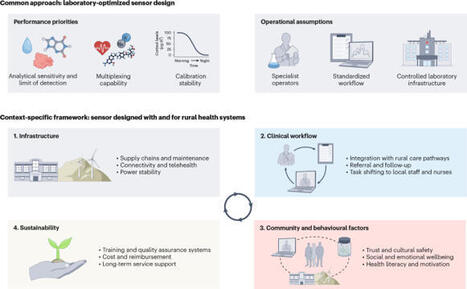
Point-of-care sensors have the potential to improve health equity in rural areas, where access to healthcare infrastructure and services is often limited. However, their readiness for scale-up and real-world deployment is judged mainly by device performance under controlled conditions. We argue that translational readiness should instead reflect how well a sensor fits the people, workflows, infrastructure and geography of its intended settings. A device that performs well in the laboratory may fail in rural settings when its design does not account for local operating conditions, workforce capacity and follow-up pathways. Here we introduce a geography-dependent readiness framework for translating point-of-care sensors into rural healthcare. The framework considers community priorities, usability, care-pathway integration and logistical constraints as upfront design criteria rather than downstream obstacles. Grounded in fieldwork across rural Australia, the framework could also apply to other regions where infrastructure, access and continuity of care are unevenly distributed. This Perspective presents a geography-dependent framework for assessing the readiness of point-of-care sensors for translation into rural healthcare settings, emphasizing the role of local context in guiding their design and deployment.
Scoop.it!
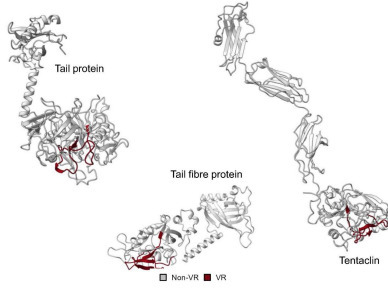
Diversity-generating retroelements (DGRs) are genetic elements that use error-prone reverse transcription to hypermutate specific target genes, most often the receptor-binding proteins of phages. They are enriched in human gut viruses, where they are hypothesised to enable phages expand their host range. Yet DGR-mediated host-range expansion across bacterial species has so far only been predicted in silico and has not been examined directly within complex microbial communities. To test this hypothesis in situ, we tracked phage-host interactions in the human gut, combining bulk metagenomic, virus-like particle (VLP), and Hi-C sequencing of longitudinal faecal samples collected over a five-week period. This culture-independent approach identified 380 interactions between 97 phages and 76 bacterial hosts. Nearly a third of these phages (31 of 97) encoded a DGR, and DGR-encoding phages were more likely to interact with multiple bacterial species. Moreover, nucleotide diversity in DGR-targeted genes increased with the number of hosts a phage interacted with and was concentrated in the surface-exposed, DGR-targeted variable region of its receptor-binding proteins. Together, these results provide in situ evidence that DGRs enable phages to expand their host range in the human gut.
Scoop.it!
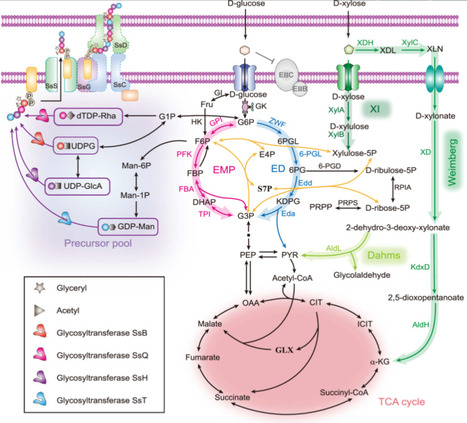
Currently, the production of high-value chemicals from natural lignocellulose is of great significance for a circular economy. In this study, a robust platform was developed based on Sphingomonas sanxanigenens NXdP, which co-utilizes glucose and xylose to synthesize polysaccharide sanxan, for efficient bioconversion of lignocellulose-derived sugars. Sanxan is a promising high-value product due to its high viscosity at low concentrations, excellent acid-gelling capabilities, and unique rheological characteristics. Through a cell growth-based screening strategy and gene manipulation, a key transporter (0233) required for the uptake of both glucose and xylose was identified, clarifying the co-utilization mechanism. Additionally, the metabolic pathways of glucose and xylose were individually blocked to assess their contributions and activities. Furthermore, the metabolic network of strain NXdP was reprogrammed via a “flux-redirection” strategy to develop a potential microbial host with improved sanxan production on mixed sugars. The final engineered strain S09 achieved a sanxan titer of 20.25 ± 0.78 g/kg under mixed-sugar fermentation, representing a 13.51% ± 4.38% increase compared with the original strain NXdP. The mechanism underlying this enhanced production was revealed by central carbon metabolism (CCM) analysis. Overall, this study provides insights into the efficient conversion of lignocellulose-derived sugars and offers genetic components for the green production of other high-value-added products.
mhryu@live.com's insight:
meng
Scoop.it!
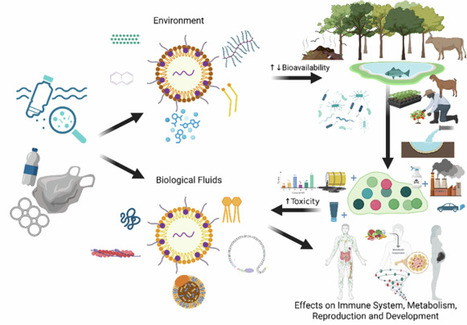
Recent data indicate plastics and associated chemicals pose risks to all forms of life - from plants, microbes, and animals to humans and their developing embryos. Plastics decompose over time into Micro and Nanoplastics (MNPs), which are present as particles or fibrils. MNPs are surrounded by other substances through hydrophobic and hydrophilic interactions, known as eco-coronas in the environment, and within living organisms as bio-coronas. We highlight key biological phenomena that may help us better understand the unique and complex impact of a Trojan horse-like exposure to the bio-corona’s cargo of MNPs. These could modify, enhance, or inhibit hormones and other active substances essential for homeostasis. The investigation of MNPs with bio-corona is of high public health relevance.
mhryu@live.com's insight:
a corona is the layer of substances that coats a micro or nanoplastic particle (MNP) after the particle meets a surrounding medium. It is a surface coating
Scoop.it!
From
pubs

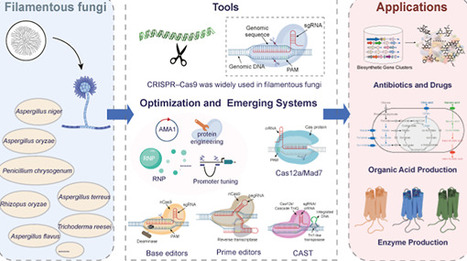
Filamentous fungi are important microbial cell factories for producing antibiotics, organic acids, enzymes, and other valuable products, with broad applications in food, agriculture, and medicine. However, inefficient and imprecise genome editing remains a major bottleneck limiting rational strain engineering. This review summarizes major genome-editing tools in filamentous fungi, including Cre-loxP, TALENs, and CRISPR–Cas systems, with emphasis on their mechanisms, advantages, and limitations. We highlight CRISPR–Cas9 as the predominant platform and discuss optimization strategies involving sgRNA design, engineered Cas9 variants, promoter engineering, AMA1-based replicative plasmids, and ribonucleoprotein delivery. Emerging technologies, including base editing, prime editing, Cas12a, and Mad7, are also discussed for their potential to broaden editing capabilities. Finally, we outline future perspectives focusing on extending the application of existing genome-editing systems in filamentous fungi and discovering novel genome-editing tools from large-scale omics data sets. This review will accelerate the development of filamentous fungi as versatile platforms for biomanufacturing.
Scoop.it!

This book explores recent, fully updated techniques involving computational approaches to the study of synthetic biology.
Scoop.it!
From
academic
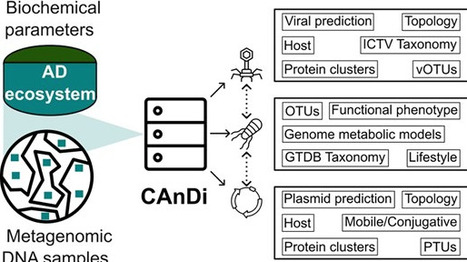
Methanogenic environments harbor complex prokaryotic consortia that drive the anaerobic conversion of organic matter into methane and carbon dioxide. Beyond being key to the global carbon cycle, such processes have been steered into powerful biotechnologies for energy production in bioreactors. Decoding the functional and taxonomic landscape of these communities requires comprehensive surveys, including bacteriophages and plasmids, which, however, remain vastly underrepresented in current databases. Here, we present the microbial Catalogue of Anaerobic Digestion (CAnDi), a database comprising 36 480 prokaryotic genomes spanning 11 149 species recovered from 1256 metagenomic samples collected across bioreactors worldwide. The reconstructed metagenomes also encode ∼1.9 million viral and over 130 000 plasmid sequences. Identity clustering revealed ∼536 000 candidate viral species, of which 85% are novel relative to existing databases, as well as over 15 000 unique plasmids. Clusters of viral and plasmid proteins were generated de novo, totalling 1 721 182 and 123 813 distinct protein families, respectively. Together, these resources substantially advance previous systematic approaches to characterize anaerobic digestion microbiomes, providing an expanded foundation for investigating the ecological and evolutionary roles of the virome and plasmidome in methane-producing ecosystems. CAnDi is accessible via https://www.candi-db.org/.
Scoop.it!
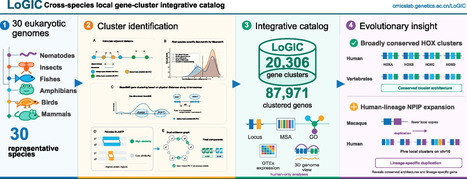
Homologous genes, especially those sharing close genomic locations and forming gene clusters, often have compensatory roles, therefore pose remarkable challenges for deciphering the functions and relationships of these genes. A comprehensive database of local homologous gene clusters with multimodal information would greatly facilitate such research. Here, we present LoGIC (https://omicslab.genetics.ac.cn/LoGIC/), a Local Gene-cluster Integrative Catalog that includes 20 306 local gene clusters identified from 30 representative eukaryotic species. In addition to the basic genomic features of each gene cluster, LoGIC also allows users to examine the sequence similarity of gene members within a cluster, the functional preference of clustered genes within a species, and the cross-species conservation of gene clusters. Information of tissue-wide expression profile and relationship with chromatin Hi-C structure is also available for human gene clusters. Together, LoGIC can facilitate studies of tandem gene duplication, paralog functional divergence, lineage-specific gene-family expansion, and evolutionary conservation or remodelling of homologous gene clusters across-species.
Scoop.it!
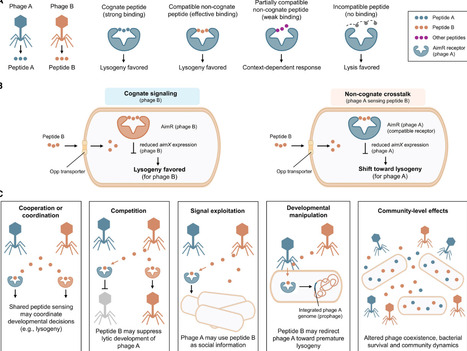
The arbitrium system has transformed our understanding of phage biology by demonstrating that temperate phages communicate through extracellular peptides to coordinate lysis-lysogeny decisions. Initially considered a highly specific signaling pathway restricted to individual phage lineages, arbitrium communication has now been shown to extend beyond lineage boundaries in some phage systems. Recent studies have shown that non-cognate arbitrium peptides can interact with heterologous receptors, providing evidence of inter-phage crosstalk. Here, we synthesize current knowledge of the molecular basis of arbitrium signaling, the structural determinants of peptide-receptor specificity, and the potential ecological and evolutionary consequences of inter-phage communication. We suggest that arbitrium crosstalk may shape phage ecology by mediating competitive and potentially cooperative interactions within shared signaling environments. We also discuss its potential uses in engineered phage systems, synthetic biology, and microbiome engineering. Collectively, the available evidence broadens the concept of viral communication beyond lineage-specific signaling, while the ecological significance and practical applications of arbitrium crosstalk remain to be established.
Scoop.it!
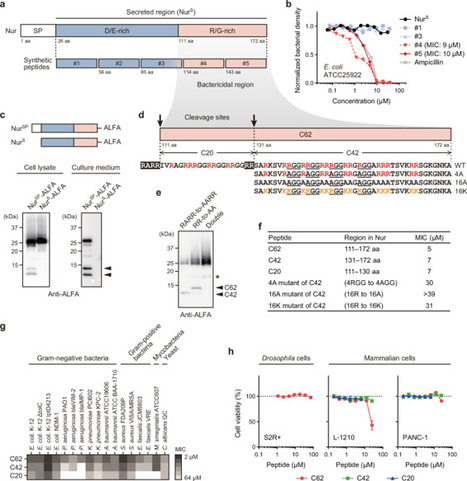
Antibiotic resistance is a global health threat requiring antimicrobials with alternative modes of action. Growing evidence indicates that bacteria organize key intracellular processes through biomolecular condensates and other forms of spatial organization, yet whether such spatial organization can be exploited as an antimicrobial target remains unclear. Here, we identify intrinsically disordered antimicrobial peptides, generated from the Drosophila sleep-inducing protein Nemuri, that inhibit bacterial growth by targeting ribosomal organization. Mechanistically, these peptides enter bacterial cells without overt membrane disruption and accumulate in ribosome-rich regions, where they bind broadly to RNA and ribosomal surfaces through multivalent interactions. This engagement remodels ribosome-rich regions into condensates with reduced ribosomal mobility, thereby suppressing global protein synthesis and distinguishing these peptides from canonical site-specific ribosome-targeting antibiotics. Furthermore, Nemuri-derived peptides exhibit broad-spectrum antimicrobial activity, including activity against drug-resistant pathogens, with low cytotoxicity toward Drosophila and mammalian cells. Together, our findings identify the organization and material dynamics of bacterial ribosomes as a biophysical vulnerability that can be exploited by antimicrobial peptides, revealing how a naturally evolved metazoan host-defense system turns intracellular organization itself into an antibacterial target. Nishinami et al. show that peptides from the sleep-inducing protein Nemuri, condense bacterial ribosomes and inhibit protein synthesis, revealing a biophysical antimicrobial strategy targeting intracellular organization.
Scoop.it!

Sequence-only predictors can score RNAs without experimental structures, but continuously updated training pools can reabsorb benchmark chains. Test overlap and within-training redundancy are often handled together, although they affect evaluation differently. DUET combines a relational graph tower with two convolutional sequence towers built on frozen RNA language models. We removed 98 of 388 training and validation records reaching 80% identity to canonical test chains, retained 110 sequence-identical records, and isolated 115 homology components across folds. Across 10 paired seeds, DUET reached 0.240 +/- 0.013 MCC versus 0.192 +/- 0.031 for CoBRA-M3 on the same data (Delta = +0.047; Holm p = 0.0073), with parallel gains in AUC and AUPRC. DUET and GRASSP were statistically indistinguishable on external sets (0.246 versus 0.243 MCC). Real three-dimensional topology improved MCC by +0.037 over shuffled edges and by +0.042 over predicted secondary structure. Increasing the training pool from 80 to 290 records raised DUET by +0.135 MCC. Benchmark evaluation therefore requires separate controls for test overlap and within-training redundancy.
mhryu@live.com's insight:
no binding partner prediction. Input: an RNA nucleotide sequence only (one chain) Output: a binding probability for each nucleotide, meaning the chance that it belongs to the binding site of a small molecule.
Scoop.it!

Microbial ecosystems can now be measured across genomic, transcriptomic, proteomic, metabolomic, and environmental dimensions, yet their emergent functions remain difficult to predict and control. Artificial intelligence (AI) can integrate heterogeneous data, but predictive accuracy alone does not establish causality, transferability, or controllability. This review developed a five-layer framework: observe, infer, predict, design, and learn that connects microbial measurement to prospective intervention and experimental feedback. We critically synthesise evidence from sequence and protein learning, multi-omics integration, metabolic modelling, synthetic microbial communities, environmental biotechnology, and temporal forecasting. Recent primary studies illustrate progress in forecasting community trajectories, predicting microbial competition and designing consortia for pollutant degradation; they also reveal important boundaries between model performance and demonstrated ecosystem control. We examine risks arising from compositional data, information leakage, dependent observations, batch effects, uncertain calibration, sparse perturbation data, ecological instability, and evolutionary change. We propose operational criteria for ecosystem programmability and a staged validation pathway from controlled systems to field-relevant environments. Progress will depend on standardised context-rich data, hybrid AI–mechanistic models, prospective benchmarks, independent replication, and safety-constrained design.
|
Scoop.it!
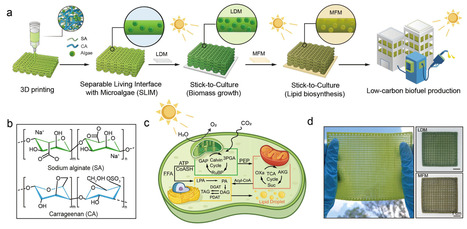
The accelerating global energy demand, coupled with diminishing fossil fuel reserves and growing carbon emissions, underscores the urgent need for sustainable energy solutions. Autotrophic microalgae are promising cell factories for converting CO2 into renewable biofuels. However, conventional microalgal biofuels typically depend on energy-intensive or nutrient-deficient regimes that suppress microalgal proliferation, resulting in a critical trade-off between biomass growth and lipid biosynthesis. Here, we report a separable living interface with microalgae (SLIM) platform that employs extracellular-matrix-mimicking hydrogel scaffolds to spatially organize microalgae and dynamically reconfigure their microenvironment, enabling spatiotemporal switching between biomass accumulation or lipid induction. The SLIM platform supports high-density growth of microalgae while facilitating precision substrate-driven metabolic regulation, enabling low-carbon operation and improved lipid production (8.89 g L−1). This work establishes an engineered living material that overcomes essential bottlenecks in microalgal biofuel production and offers a generalizable blueprint for biohybrid systems in advanced biomanufacturing. Balancing biomass growth and lipid accumulation remains a challenge in microalgal biofuel production. Here the authors report extracellular matrix-mimicking hydrogel scaffolds that dynamically reconfigure the microenvironment of spatially organized microalgae, allowing controlled switching between the two states.
mhryu@live.com's insight:
1str, prints living microalgae (Nannochloropsis oceanica) mixed into a hydrogel bioink (alginate+carrageenan), using an extrusion 3D bioprinter. The printed sheet is moved from one medium-containing substrate to another. in step one, the sheet sits on nitrogen-rich, low-salinity LDM substrate (optimized at 2 g/L nitrate and 10 g/L NaCl), where cells proliferate and the sheet turns dark green, building biomass. In step two, the same sheet is moved to nitrogen-limited, higher-salinity MFM substrate (0.5 g/L nitrate, 21.22 g/L NaCl) for 14 days, where nutrient and salt stress redirects carbon into fatty acid and triacylglycerol synthesis, lipid droplets form, and the sheet turns dark brown.
Scoop.it!

The Arg/N-degron pathway regulates important agronomic traits. In plants, the PROTEOLYSIS 1 (PRT1) protein binds to aromatic amino-terminal residues and recruits ubiquitin (Ub) conjugating enzymes to ubiquitinate substrate proteins. Here, we demonstrate that PRT1 recruits the UBC35-UEV1A complex via RING1 of PRT1. This stimulates UBC35 to produce K63-linked Ub chains. To afford chemical control over Ub signalling, we construct a synthetic substrate of PRT1 with an N-terminal calmodulin-binding peptide (CBP). The addition of the CBP sequence to the substrate creates an off switch for PRT1-dependent ubiquitination that is regulated by calcium and calmodulin. Finally, we produce a set of nanobodies that are recognised by PRT1 and induce the ubiquitination of a reporter protein. Altogether, we introduce and biochemically validate several tools to both redirect and chemically control the Arg/N-degron pathway. The Arg/N-degron pathway plays a central role in plant development, germination, and submersion tolerance. Here, the authors show that PRT1 recruits the UBC35-UEV1A complex via RING1 of PRT1, demonstrate an off-switch for calmodulin-controlled ubiquitination, and engineer nanobodies that can redirect PRT1.
mhryu@live.com's insight:
1str, To downregulate components of the N-degron pathway, we designed artificial microRNAs (amiRNAs) cloned as pre-amiRNAs that were embedded, as introns, within the CBP-RFP reporters. In this system, the presence of RFP indicates a transformed cell or protoplast with correctly spliced out pre-amiRNAs. Built on the Arabidopsis miR319a scaffold, with the targeting sequence designed using the WMD3 Web MicroRNA Designer. Cloned as a pre-amiRNA inside an intron of the reporter gene (CBP-RFP). The cell processes the intron into the mature amiRNA, which silences the target mRNA.
Scoop.it!
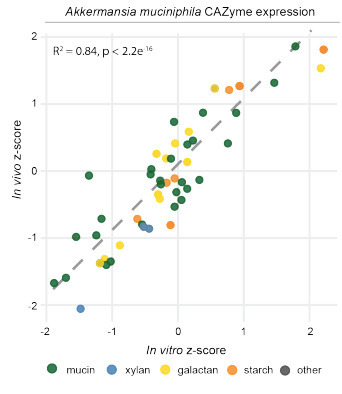
Dietary fiber deficiency shifts the gut microbiota to degrade host mucus, compromising intestinal homeostasis. Mechanistically linking in vitro metabolic dynamics of fiber degradation to in vivo outcomes remains challenging due to a lack of representative model systems. Introducing spatial mucin structures to a defined Oligo-Mouse-Microbiota (OMM12) in vitro consortium allowed us to recapitulate the spatio-temporal dynamics of xylooligosaccharide (XOS) processing in gnotobiotic mice. XOS supplementation rapidly established a stable community state in vitro and in vivo across compositional, metabolic and transcriptional levels. XOS supplementation established distinct functional guilds: Bifidobacterium animalis and Blautia coccoides partitioning xylose and arabinose catabolism, whilst profiting from XOS degradation by Bacteroides caecimuris. Along the colonic axis, a spatial XOS gradient exposes Bl. coccoides's substrate preference for xylose and drives longitudinal functional adaptations. Altogether, our predictive platform proves that regional microenvironments govern prebiotic efficacy, highlighting spatial cascades as essential parameters for targeted microbiome interventions.
mhryu@live.com's insight:
sauer u, syncom
Scoop.it!
From
academic
The fundamental biochemical and stereochemical difference in how cell membrane lipids are built between Archaea on one side, and Bacteria and Eukarya on the other is known as the lipid divide. It is often assumed to translate in the impossibility of combining membranes of both types. It was established early when little sampling of the biodiversity was available. Since then, exceptions have been revealed and accumulated. Here, we review those exceptions and complement them with a broad survey, revealing diverse bacteria with mixed membrane-lipid features. Together with liposome assays and synthetic-biology experiments, studies on these organisms suggest that the lipid divide can be bridged. We conclude by discussing the evolutionary implications for LUCA, the origin of the prokaryotic domains and eukaryogenesis.
Scoop.it!
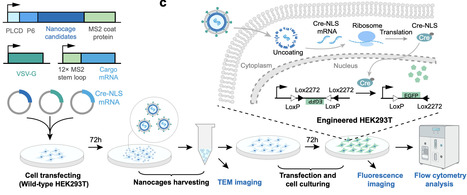
Safe and effective gene delivery remains a central challenge for therapeutic applications. While non-viral and viral vectors have enabled substantial progress, their reliance on non-human components often triggers immune responses, limiting their use in chronic treatments. Here, we develop DeepDelivery, an artificial intelligence-driven platform to identify human proteins with scaffold potential for mRNA delivery. Proteome-scale screening nominated 512 candidates, and experimental validation show that 12 of 15 selected human proteins functioned as scaffold components that facilitated particle formation and mRNA delivery within a modular engineering framework. Notably, several tripartite motif (TRIM) family proteins, typically linked to antiviral responses, exhibit strong assembly and delivery activity. Model interpretability and perturbation analyses identify TRIM domain features that influenced prediction scores and guide TRIM25-specific domain engineering to improve delivery. Together, these results establish a framework for mining the human proteome for scaffold components of programmable mRNA delivery systems and for further exploring the design space of human-derived delivery architectures. Protein-based carriers could provide biocompatible options for mRNA delivery. Here, authors develop an AI-guided platform to identify and engineer human-derived protein scaffolds that assemble into particles and support mRNA delivery in cultured cells.
mhryu@live.com's insight:
Build the training dataset (V1, V2) from InterPro viral capsid families and unrelated negatives. Train and select the model architecture (the one dimensional convolutional network), then train Model 1 on V1 and Model 2 on V2. Run the human proteome (UniProt entries, 100 to 1024 residues) through both trained models. A protein scoring above 0.5 in both becomes a candidate.
Scoop.it!
From
pubs
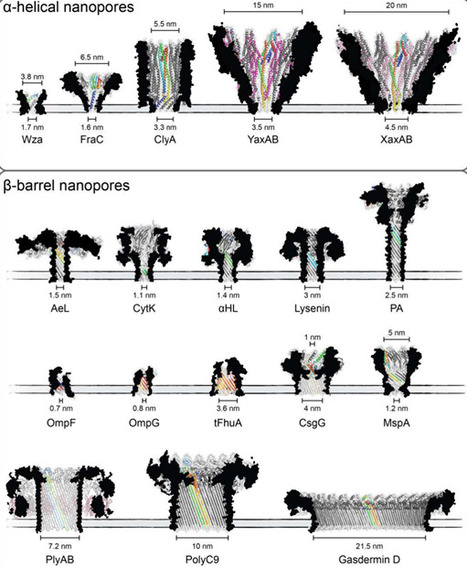
Protein-based biological nanopores have emerged as versatile tools for sequencing biopolymers, identifying molecules, and probing reactions at the single-molecule level. Many biological nanopores have been characterized, revealing the significance of nanopore size, shape, and chemical composition in molecular detection and discrimination. Advancements in understanding the nanofluidic properties of nanopores have complemented engineering approaches expanding the possible applications of nanopore technology. Through a combination of structural insights, rational design, systematic screening, and molecular dynamics simulations, researchers have developed nanopores capable of distinguishing between ions, nucleic acids, peptides, and folded proteins with remarkable specificity and sensitivity. Building upon three decades of progress, this Review highlights recent engineering innovations, discusses ongoing challenges, and explores the prospects in the field.
Scoop.it!
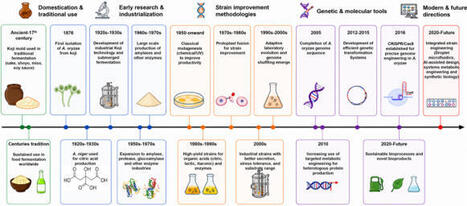
Filamentous fungi of the genus Aspergillus, because of their exceptional secretory capacity and metabolic versatility have been extensively applied in traditional food fermentation and industrial biotechnology for production of diverse bioproducts. Their history of domestication and industrial application has enabled major progress in enzyme production; however, challenges associated with the filamentous growth and strain engineering for future biomanufacturing development still remain. This review summarizes the advances across the spectrum of strain-improvement methodologies, from classical mutagenesis, protoplast fusion, genome shuffling, and adaptive laboratory evolution to the modern era of CRISPR enabled genome editing, synthetic biology, and systems-level metabolic engineering. Previous reviews have primarily focused on individual Aspergillus species, specific industrial products or particular strain improvement approaches. In this review, we provide a comprehensive overview of classical and modern strain engineering technologies and industrial scale up across a wide range of Aspergillus species for industrial enzyme production. In particular, emerging non-GMO evolution tools such as atmospheric and room temperature plasma (ARTP) mutagenesis, and modern molecular biotechnology coupled with microdroplet-based ultrahigh-throughput screening, now allow the researchers to generate and screen a large number of Aspergillus species mutants with single-cell precision. The discovery of high-performance strains with enhanced secretion, stress tolerance, substrate utilization, and favorable enzymatic profiles for food and bioprocessing applications has been accelerated by the strain innovations. Collectively, these advances establish A. oryzae and the related species as increasingly reliable, scalable, and programmable hosts that can meet the biomanufacturing requirements of future circular and sustainable bioeconomies.
mhryu@live.com's insight:
m-1str, protoplast fusion
Scoop.it!
Long non-coding RNAs (lncRNAs) have diverse biological functions, but experimentally validated evidence remains scattered across the literature and is often context dependent. EVLncRNAs 4.0 (https://www.sdklab-biophysics-dzu.net/EVLncRNAs4/) updates our manually curated database of functional lncRNAs supported by low-throughput experiments. For this release, 16 581 publications indexed between 15 May 2023 and 15 May 2026 were retrieved and manually screened. Compared with EVLncRNAs 3.0, EVLncRNAs 4.0 contains 7808 functional lncRNAs (+26%) from 199 species (+23%), 2313 diseases (+34%), 28 757 curated functional records (+21%), and 18 380 molecular interactions (+25%). EVLncRNAs 4.0 also introduces two AI-assisted functions built on the curated evidence base. AI Search uses an enhanced retrieval-augmented generation framework with adaptive selection across Quick, Expanded, and Exhaustive retrieval modes and generates answers from retrieved EVLncRNAs records with PMID-linked citations. Benchmark evaluation showed complete request execution, high retrieval coverage and full citation validity within the supplied evidence context. Functional Overview retrieves and aggregates lncRNA- and species-specific curated records before generating concise evidence-based summaries. In both functions, language models synthesize retrieved evidence rather than serve as independent sources of biological knowledge. EVLncRNAs 4.0 therefore provides an expanded, experimentally grounded resource for accessing and interpreting functional lncRNA evidence across species and biological contexts.
Scoop.it!
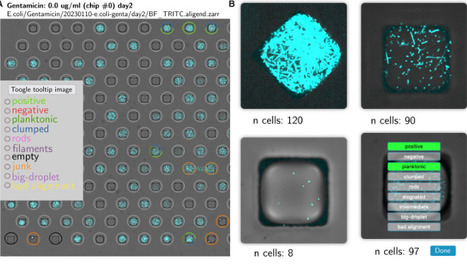
Bacterial morphology is modified by antibiotic stress depending on the antibiotic mechanism of action, while changes in morphology can contribute to cell survival. Here we present a workflow that generates morphological signatures for the progeny of individual bacterial cells across 168 antibiotic conditions. The workflow uses stationary microfluidic droplets to encapsulate and grow bacteria and confocal microscopy to image the droplet content. A custom image analysis pipeline labels morphologies in a subset of images to train a neural network. The network yields multidimensional morphological signatures from 82,000 droplets, showing the co-existence of different morphologies even for the progeny of individual cells. The signatures differ with antibiotic type and concentration, providing a way to distinguish antibiotics by their mode of action. Combined with digital detection of bacterial survival within droplets, this workflow provides a quantitative platform to investigate antibiotic resistance and identify the antimicrobial activity of unknown compounds.
mhryu@live.com's insight:
tool
Scoop.it!

Biomolecular condensates have so far been studied in terms of their structural, compositional, and functional properties. However, condensate enzymatic activity - a key aspect of cellular metabolism - remains unexplored due to the complexity of the system. In this study, using a combination of experimental, computational and theoretical techniques, we have discovered that the non-equilibrium activity which originates from catalytic reactions couples with the environment through various feedback mechanisms across five orders of magnitude of length scales. We observe that condensed enzymes catalyze more rapidly in the presence of crowding proteins and show that the increased enzymatic activity within these droplets stems from the emergence of lower-energy protein conformations induced by the highly crowded environment. Despite the crowding in the environment of the droplet, which might suggest an effective increase in its overall viscosity, we find that it becomes more agile, as evidenced by the observation of enhanced diffusion and macroscopic flow, due to the enzymatic activity. These findings shed light on the dynamic interplay between enzymatic activity, composition and crowding in condensates, and their roles on the mobility and accessibility of various functional units in these environments, expanding our understanding of liquid-liquid phase separation in metabolically active conditions. Enzymes operate in highly crowded cellular environments, yet the impact of crowding on their function remains poorly understood. Here, the authors demonstrate that enzyme activity increases molecular mobility in crowded protein droplets, suggesting a feedback mechanism between catalysis and the physical properties of the surrounding environment.
mhryu@live.com's insight:
llps, Crowded protein droplets allow the partitioning of enzymes to reach high enzymatic activity compared to dilute conditions. Computational dissection of enzyme conformations reveals the atomistic changes in the presence of protein crowding. Fluorescently tagged crowder proteins (scale bars 5 µm) and nanoparticle tracking experiments indicate that there is faster movement in the crowded droplet environment in the presence of enzymatic activity (scale bars 5 µm). Large-scale flows in the droplets emerge due to chemical gradients produced by enzymes.
Scoop.it!
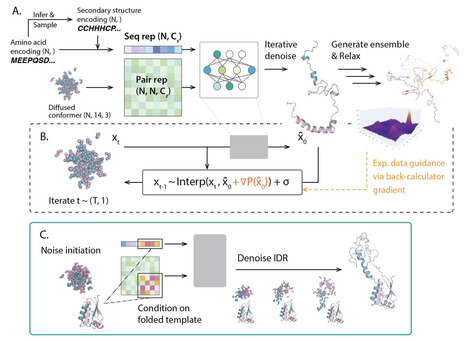
Although machine learning has transformed protein structure prediction of folded protein ground states with remarkable accuracy, intrinsically disordered proteins and regions (IDPs/IDRs) are defined by diverse and dynamical structural ensembles that are predicted with low confidence by algorithms such as AlphaFold and RoseTTAFold. We present a machine learning method, IDPForge (Intrinsically Disordered Protein, FOlded and disordered Region GEnerator), that exploits a transformer protein language diffusion model to create all-atom IDP ensembles and IDR ensembles that maintains the folded domains. IDPForge does not require sequence-specific training, back transformations from coarse-grained representations, nor ensemble reweighting. The created IDP/IDR conformational ensembles show good agreement with both NMR and SAXS solution experimental data, and options for biasing with experimental restraints are available if desired. IDPForge with these diverse capabilities facilitates integrative and structural studies for proteins that contain intrinsic disorder, and is available as an open source resource for general use. Intrinsically disordered proteins and regions (IDPs/IDRs) are defined by diverse structural ensembles. We present IDPForge that exploits a transformer protein language diffusion model to create all-atom IDP and IDR ensembles that maintain folded domains with options to conform to experimental data.
mhryu@live.com's insight:
IDPForge generates structures for disordered regions one already defined.
Scoop.it!
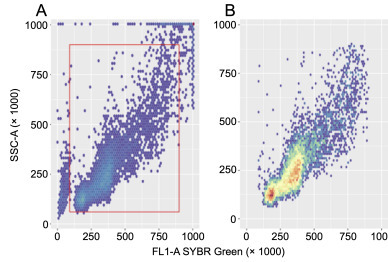
Karyotypic variation in fungal populations from ecological, clinical, and industrial environments and is also a hallmark of many cancers. The evolutionary dynamics of karyotypic vriants (i.e., changes in the number of chromosomes, either as complete sets or single chromosomes) are the same as any de novo mutation. To quantify karyotypic subpopulations within evolving populations, an unbiased computational method is required to identify subpopulations and quantify their abundance. Flow cytometry is the standard method to measure the genome size from large populations of individual cells. Cell populations typically include cells in all stages of the cell cycle (G0/G1 before DNA replication, S phase during replication, and G2/M after DNA replication but before cell division). Existing mathematical models can be fit to genome-size distributions to determine the base ploidy of a population, but existing algorithms assume a single base ploidy. Mixed-karyotype populations therefore require manual gating to subdivide the flow cytometry data before analysis, which introduces bias, as it is not always clear where or when gates should be drawn. To overcome this, we developed a new method that identifies and quantifies karyotypic subpopulations from flow cytometry data. The open-source R package ploidyPeaks builds on existing Bioconductor tools to import and manipulate standard .fcs flow cytometry files. Using reference populations of known ploidy, we established thresholds to distinguish single-genome size populations and identify candidate mixed-karyotype samples. We then developed a peak-detection algorithm to identify G0/G1 and G2/M peaks, resolve karyotypic subpopulations within mixed populations, and apply nonlinear least-squares fitting of the Dean-Jett-Fox cell-cycle model to each subpopulation, providing a confidence measure for model fit. Compared with existing methods, our approach quantifies genome size in mixed-karyotype populations without manual intervention while providing an objective measure of model fit. We demonstrate the utility of this method on 240 replicate populations initiated from 20 Candida albicans genetic backgrounds evolving for 100 generations to the antifungal drug fluconazole.
mhryu@live.com's insight:
inoculated from frozen stocks into YPD and grown overnight, then fixed in ethanol, treated with RNase A, and stained overnight at room temperature with SYBR Green.
|
The DNA that ends up as single strands is not the integrated genome. It comes from a phagemid flanked by the M13 packaging sequence. The M13 proteins made from the chromosome act on the phagemid the way a helper phage or helper plasmid would. The phagemid is copied into a circular single strand, packaged into M13 particles, and secreted into the culture medium.