Your new post is loading...

|
Scooped by
mhryu@live.com
Today, 1:51 AM
|
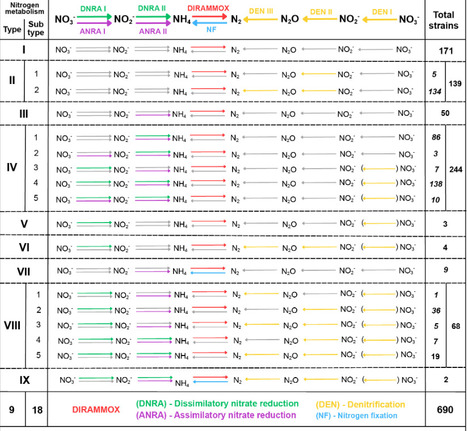
Our knowledge of the nitrogen cycle has been advanced by the discovery of anammox bacteria, ammonia-oxidizing archaea, and comammox bacteria, along with their high diversity. Direct ammonia oxidation (dirammox), a recently discovered heterotrophic nitrification pathway, converts ammonium and organic nitrogen directly to N2 through the DnfABC multienzyme system, further expanding our understanding of the nitrogen cycle. However, the taxonomic distribution, prevalence, and nitrogen metabolic potential of microorganisms encoding dirammox genes remain poorly understood. Here, large-scale genome mining identified complete dnfABC gene clusters in 690 genomes representing 3 phyla and 26 genera. The dnfABC cluster showed a striking bimodal prevalence: it was nearly ubiquitous in Alcaligenes, Delftia, and Microvirgula but occurred sporadically in most other genera. These microorganisms were recovered from diverse environments, particularly organic nitrogen-rich habitats. Comparative analyses of DnfABC phylogeny, gene cluster organization, GC content, and flanking mobile genetic elements indicate that the dnfABC cluster is conserved within several core genera, whereas its sporadic distribution in other lineages suggests that horizontal gene transfer may have contributed to its dissemination. The dnfABC cluster co-occurs with genes associated with assimilatory nitrate/nitrite reduction to ammonium, dissimilatory nitrate/nitrite reduction to ammonium, denitrification, and/or nitrogen fixation, but not with marker genes for nitrification or anammox, indicating diverse nitrogen metabolic potential among potential dirammox microorganisms. Functional assays further showed that diverse dnfABC clusters consistently conferred ammonium- and glutamine-oxidizing activities in Escherichia coli, despite exhibiting low activities in their native hosts. These results suggest that dirammox is a broadly distributed, metabolically flexible, and tightly regulated pathway and may represent an overlooked route of nitrogen transformation in organic-rich habitats.
|
|
Scooped by
mhryu@live.com
Today, 1:33 AM
|
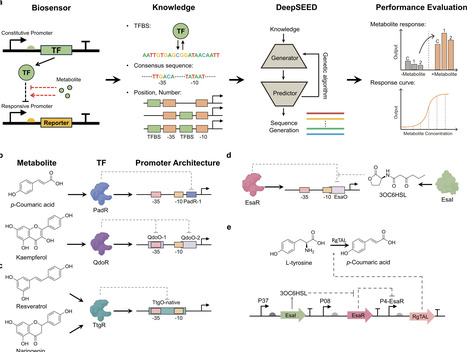
Transcription factor-based biosensors are essential for the scalable bioproduction of plant natural products. However, a major bottleneck in developing high-performance biosensors is the engineering of responsive promoters due to the vast sequence space and poorly understood sequence–function relationships. Here, we extend our previously published DeepSEED framework into an artificial intelligence-driven pipeline for efficient and extensible biosensor-responsive promoter optimization. This strategy integrates deep learning with domain-specific biological expertise. By fixing core promoter elements, varying the number and position of transcription factor binding sites (TFBSs) to define promoter backbones, and subsequently optimizing the flanking sequences, we enable the de novo design of promoters. We systematically apply this strategy to optimize a panel of biosensors across different transcription factors and regulatory modes, including four metabolite-responsive biosensors and a quorum-sensing-based environmental sensor. The resulting biosensors exhibit diverse dynamic behaviors with markedly improved sensitivity, expanded dynamic range, and increased maximum output. For instance, the optimized naringenin biosensor demonstrates a 32-fold broader dynamic range and a 27-fold higher maximum output compared to its parent. To illustrate its utility in metabolic engineering, we employed a DeepSEED-designed promoter to dynamically regulate a p-coumaric acid biosynthesis pathway, boosting production by 117.0%. Our work extends the previously published DeepSEED framework into a powerful pipeline for biosensor optimization and delivers a portfolio of high-performance regulatory elements, addressing a critical need and facilitating the development of more intelligent and productive metabolic engineering systems.
|
|
Scooped by
mhryu@live.com
Today, 1:22 AM
|
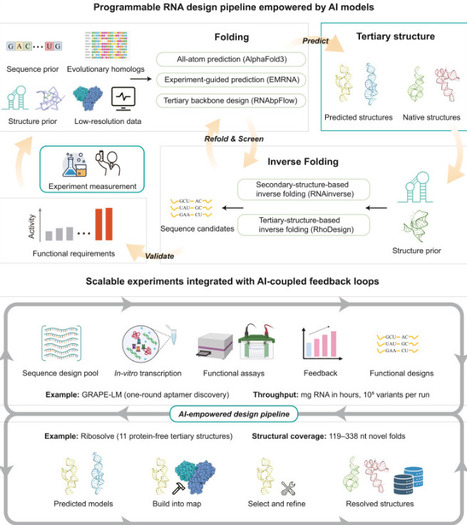
Functional RNA design is central to synthetic biology and RNA therapeutics. Recent AI advances leverage tertiary structure to interpret function and guide sequence generation, replacing traditional trial-and-error workflows. Experimental methods assisted by AI models continuously expand the structural data foundation and ultimately enable scalable and programmable RNA design.
|
|
Scooped by
mhryu@live.com
Today, 1:15 AM
|
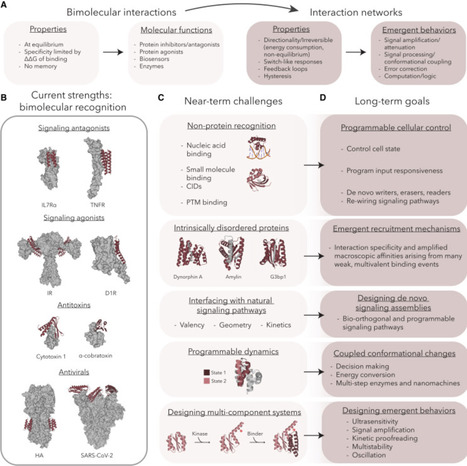
Driven by advances in artificial intelligence, de novo protein design can now generate diverse molecular functions from the ground up, achieving remarkable success in engineering binders against many macromolecular targets. Here, we provide a perspective on how de novo design can progress beyond bimolecular interactions toward the bottom-up construction of protein reaction networks. We highlight how programmable, modular, and controllable de novo components can drive complex, emergent biological behaviors. Overcoming current challenges—including designable protein dynamics, multi-step functionalities, and non-equilibrium systems—is essential for this transition. With continued foundational method improvements, the field is poised to pioneer precise protein perturbations for systems-level insight and to engineer emergent cellular behaviors like signal amplification, decision-making, and error correction de novo to fundamentally advance synthetic biology, biotechnology, and medicine.
|
|
Scooped by
mhryu@live.com
Today, 1:10 AM
|
A weakness in glycobiology research is the insufficient integration of approaches used to connect glycan structure to function. From glycogene knockouts to understanding glycan structures, dynamics, and interactions, to determining whether a single glycan structure underlies a specific cellular function, no single approach provides sufficient information to establish these relationships. We argue that resolving glycan structure–function relationships requires the integration of complementary approaches spanning genetics, analytical glycomics, glycoengineering, structural biology, and computational modelling. The field stands at a transition point: from cataloguing glycan structures to understanding their diverse roles in regulating the biology of health and disease. Such an integrated approach will be critical for translating structural glycan information into mechanistic insights and, ultimately, informed glycoengineering and therapeutic design.
|
|
Scooped by
mhryu@live.com
Today, 12:49 AM
|
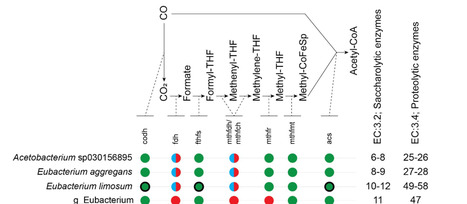
Syngas conversion to carboxylic acids can be achieved through the combined actions of acetogens and chain elongators. Mixed cultures enriched with these functional groups can serve as robust and resilient catalysts for bio-based production, but developing efficient and stable communities remains challenging. In this study, we aimed to enrich a mixed microbial culture for efficient syngas-driven chain elongation using carbon monoxide (CO) and acetate as substrates. Three strategies were employed: (i) conventional enrichment using anaerobic sludge only, (ii) sludge augmentation with known syngas fermenters (Clostridium autoethanogenum, Clostridium carboxidivorans, Eubacterium limosum, and Acetobacterium wieringae) to enhance carboxydotrophic activity, and (iii) sludge augmentation with chain elongator Clostridium kluyveri to stimulate chain elongation. Enrichments were performed in batch cultures with stepwise increases in CO partial pressure across successive transfers until a 100% CO (170 kPa) headspace was reached. Ultimately, all enrichments grew under a 100% CO headspace, simultaneously utilizing CO and acetate, with butyrate as main product. 16S rRNA gene and metagenomic analyses revealed that Eubacterium became predominant. Several genera (Aminobacterium, Anaerotignum, Oscillibacter, and Proteiniphilum) not known to ferment CO, were also enriched. Based on physiological and metagenomic analyses, we hypothesize that Eubacterium consumes CO and acetate, producing butyrate and extracellular polymeric substance (EPS)-like material. Metagenomic data further suggests that components derived from this EPS-like material may serve as substrate for the other enriched genera. This study demonstrates that augmentation can steer mixed microbial communities toward targeted product formation and highlights the critical role of initial culture conditions in shaping community composition and functionality.
|
|
Scooped by
mhryu@live.com
Today, 12:44 AM
|
Klebsiella pneumoniae is an increasingly important opportunistic pathogen worldwide, particularly due to the emergence and spread of hypervirulent K. pneumoniae (hvKP), a clinically significant pathotype first identified in Taiwan. This study aimed to molecularly identify hvKP isolates screened for coexisting virulence determinants and evaluate the association between specific virulence genes and hypervirulence. Clinical specimens were collected from hospitalized patients, and K. pneumoniae isolates were identified phenotypically using microscopic culture characteristics and biochemical assays and genotypically confirmed by PCR targeting the khe gene. Hypermucoviscosity regulators prmpA and prmpA2, siderophore-encoding genes iucA and iroN, and other virulence-associated genes including wzi, magA, wzyK2, wabG, and peg-344 were investigated. Eighty-two capsulated non-motile K. pneumoniae isolates were confirmed with prmpA and prmpA2 detected in 42 and 29 isolates, respectively, whereas iucA was identified in 31 isolates and iroN in 6 isolates only. Based on the molecular criteria, 34 isolates were classified as hvKP and 48 as classical K. pneumoniae (cKP). A strong association was observed between the peg-344 gene and hypervirulence (p = 0.0001), whereas hypermucoviscosity showed a weaker correlation. However, low-speed sedimentation assay (LSSA) analysis demonstrated a significant relationship between the hypermucoviscous phenotype and genotypic hypervirulence (p = 0.041). These findings highlight the molecular determinants of hvKP and identify peg-344 as the gene most strongly associated with hypervirulence; however, its moderate sensitivity (58.8%) and absence in over 40% of hvKP isolates indicate that it is better suited as a supportive confirmatory marker than a standalone diagnostic biomarker for hvKP
|
|
Scooped by
mhryu@live.com
Today, 12:30 AM
|
CRISPR-based RNA cleavage holds promise for gene knockdown but its applications in metabolic engineering remain largely unexplored. Here we reconstitute three mammalian CRISPR-based RNA cleavage systems to arrest the intermediate metabolic reactions in the yeast Candida viswanathii to produce hydroxydecanoic acid (HDA), an important nylon precursor. We identify hfCas13d as the optimal effector and next develop a circular guide RNA (CgRNA) precursor architecture to enhance and extend the knockdown of genes involved in HDA production. This hfCas13d/CgRNA precursor platform also represses genes in other yeasts and E. coli, demonstrating broad microbial applicability. Single and dual CgRNA precursor arrays enable multiplexed pathway repression, with dual arrays reducing byproduct formation and increasing HDA titer to 17.8 g/L in shake flasks. Bioreactor fermentation further boosts HDA production to 37.2 g/L. Together, this work establishes hfCas13d/CgRNA as a programmable RNA regulatory platform for engineering microbial cell factories. CRISPR-based RNA cleavage holds promise for gene knockdown but its applications in metabolic engineering remain largely unexplored. Here the authors develop a circular guide RNA architecture to enhance the knockdown of genes involved in HDA production.
|
|
Scooped by
mhryu@live.com
Today, 12:18 AM
|
Cell-free DNA (cfDNA) carries extensive biological and clinical information, reflecting cell death, immune processes and tissue-specific chromatin organization. However, its short-fragment size and low abundance limit comprehensive molecular characterization by conventional approaches. Nanopore sequencing enables direct, single-molecule analysis of cfDNA, preserving sequence, methylation and fragmentation features without amplification bias. Recent advances in pore chemistry, basecalling and library preparation have made nanopore-based cfDNA analysis increasingly feasible across applications including oncology, prenatal diagnostics, transplantation monitoring and infectious disease. This Primer provides a methodological framework for cfDNA nanopore sequencing, covering pre-analytical variables, library preparation for short DNA fragments, sequencing strategies and downstream data analysis. Emphasis is placed on basecalling, methylation detection, fragmentomic profiling and integration of multi-omic signals. We discuss key technical challenges, including sequencing accuracy, fragment-length bias and limitations in detecting low-frequency variants, and highlight practical considerations for reproducibility, quality control and reporting. Finally, we outline emerging directions, including single-molecule methylome analysis, improved computational models and integration of multilayer cfDNA signals for diagnostic applications. Cell-free DNA is generally fragmented and present in low abundance, making its detection and characterization challenging. In this Primer, Korabečná et al. discuss the use of nanopore sequencing for cell-free analysis from blood samples, as well as associated technical challenges.
|
|
Scooped by
mhryu@live.com
October 15, 11:59 PM
|
Gram-negative bacteria build their cell envelope by transporting various sugar-based precursors across the inner membrane on the shared lipid carrier undecaprenyl (also known as C55). Because the cellular pool of undecaprenyl is finite and used by multiple competing pathways, it must be efficiently regenerated to sustain envelope assembly. Yet the enzymes that release polysaccharide cargo from the lipid carrier have remained elusive, leaving a major gap in our understanding of bacterial envelope biogenesis. Here we identify three periplasmic enzymes in E. coli - UprA, UprB, and UprC - that hydrolyze the bond linking polysaccharide precursors to undecaprenyl. UprA releases elongating peptidoglycan glycan strands from the inner membrane, catalyzing the long-sought terminal step of peptidoglycan synthesis. UprB cleaves the undecaprenyl-linked initiating unit common to the enterobacterial common antigen and O-antigen pathways, while UprC is the likely release factor for the colanic acid pathway. These enzymes define a widespread Upr-like protein family (formerly DUF218) as polysaccharide release factors, revealing a unifying mechanism by which bacteria coordinate envelope assembly. Because all three Upr-like proteins share a conserved catalytic core, this family represents a potential target for antibiotics that simultaneously disrupt multiple envelope pathways.
|
|
Scooped by
mhryu@live.com
October 15, 11:17 PM
|
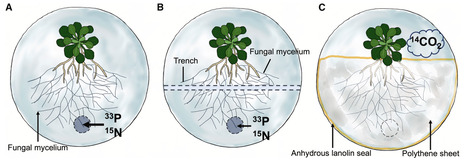
Arabidopsis thaliana, a widely used genetic model species, is considered incapable of forming mycorrhizal associations, which support nutrient acquisition in most other land plants. Mucoromycotina “fine root endophytes” (MFREs) are a recently characterized group of mycorrhizal fungi capable of forming nutritional mutualisms with a wide range of plants, supplying nitrogen and phosphorus to host plants and gaining access to photosynthetically derived carbon. Here, we demonstrate that a wild-isolated MFRE strain, Lyc-1, successfully colonizes Arabidopsis roots under monoaxenic conditions without affecting plant growth. Using a combination of conventional light and epifluorescence microscopy, we demonstrate extensive extra- and intraradical colonization of the root cortex. Stable- and radio-isotope tracing further revealed bidirectional nutrient exchange between the host plant and the fungus. To our knowledge, this is the first demonstration of a functional, mycorrhiza-like nutritional mutualism in Arabidopsis. As Arabidopsis lacks a complete set of common symbiosis signaling pathway genes, our findings suggest that Arabidopsis-Lyc-1 symbiosis may be controlled by an alternative, so far uncharacterized, symbiotic pathway. Thus, our study not only supports the existence of multiple evolutionary routes to plant-fungal nutritional mutualisms but also establishes Arabidopsis as a genetically tractable model to study plant-fungal symbiosis, thereby opening new opportunities to dissect the molecular basis and evolution of mycorrhizal symbioses.
|
|
Scooped by
mhryu@live.com
October 15, 10:18 PM
|
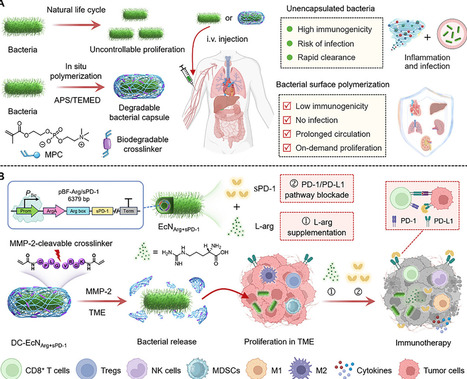
Engineered bacteria have emerged as a promising therapeutic modality but face safety risks and delivery challenges in clinical practice. Herein, we develop a programmable encapsulation technology that formulates individual bacteria with a thin formulation layer of cross-linked polymers, which confers live bacteria with reduced immunogenicity and restricted proliferation in healthy organs. To leverage these benefits, an engineered E> coli Nissle strain capable of converting tumor-accumulated ammonia into L-arginine and secreting soluble programmed cell death protein 1 (sPD-1) was encapsulated to synthesize a degradable bacterial capsule, optimizing both biosafety and delivery processes while preserving therapeutic function. Upon reaching the tumors, matrix metalloproteinase-2 triggers bacterial release and local proliferation, achieving L-arginine-driven tumor immune microenvironment modulation and sustained PD-L1 blockade, ultimately initiating robust antitumor immune responses. Overall, this programmable encapsulation platform tackles both safety risks and delivery challenges of bacterial medicines, broadening the clinical application prospects of live bacterial therapeutics.
|
|
Scooped by
mhryu@live.com
October 15, 5:20 PM
|
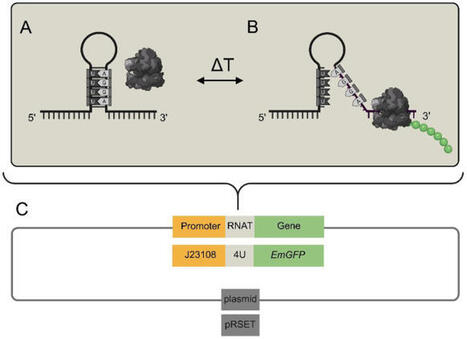
Precision fermentation is a key technology for the future sustainable production of food proteins, bulk ingredients, and high-value recombinant products. However, process efficiency is often limited by the coupling of biomass formation and recombinant protein synthesis, leading to substantial carbon losses into non-recyclable biomass. Particularly for cost-sensitive bulk products such as food proteins, economically viable production requires highly efficient and dynamically controllable bioprocesses. RNA thermometers (RNATs) offer a promising inducer-free strategy for temperature-dependent control of recombinant protein production and dynamic bioprocess regulation, with the potential to improve carbon efficiency and product yields. Results In this study, a fluorescent reporter protein (EmGFP) under the control of a fourU (4U) RNA thermometer was evaluated in E. coli BL21 across a temperature range of 25–40 °C. Temperature-dependent regulation enabled maximal recombinant protein production at 35 °C, resulting in a 2.6-fold higher specific productivity compared to the derepressed control. The strongest RNAT-mediated regulatory effect was observed at 35.5 °C, highlighting the relevance of precise temperature control for recombinant protein production. Evaluation of altered carbon partitioning revealed a substantially increased carbon allocation toward product formation, while carbon partitioning into biomass and CO2 remained largely unchanged. Furthermore, the product-per-acetate yield increased more than threefold at induction temperature, indicating controllable carbon partitioning and reveals potential for decoupling approaches of biomass formation from recombinant protein production. This study demonstrates the successful implementation of an RNAT-controlled recombinant expression system enabling temperature-dependent regulation of protein production in E. coli. Quantitative analysis revealed controllable carbon partitioning toward recombinant protein synthesis and defined target temperatures for future process induction and switching. In contrast to conventional inducer-based systems, RNAT-mediated regulation enables dynamic control without chemical additives, supporting more resource-efficient bioprocess operation. However, fully exploiting RNAT-based process control will require advanced model-based strategies integrating temperature and feeding regimes, particularly for cost-sensitive precision fermentation processes such as food protein production. Overall, RNATs represent promising molecular control elements for future high-efficiency precision fermentation processes.
|
|
|
Scooped by
mhryu@live.com
Today, 1:48 AM
|
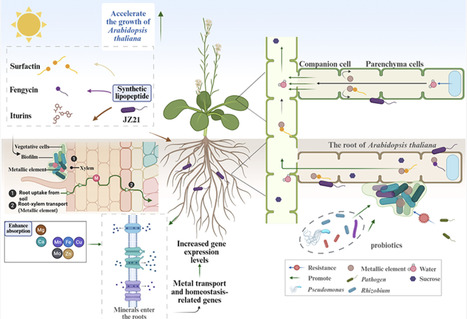
The plant-beneficial bacterium Bacillus velezensis and related Bacillus species are increasingly recognized as important contributors to plant growth and health, yet the ecological functions of their secondary metabolites in situ remain poorly understood. Here, we investigated how lipopeptides produced by Bacillus species influence plant performance and root-associated microbial communities. Using Arabidopsis thaliana as a model host, we show that bacterial lipopeptides produced during root colonization promote plant growth by enhancing the uptake of essential mineral elements, including magnesium, calcium, iron, and zinc. Transcriptomic analyses revealed that lipopeptides upregulated host genes involved in metal transport and homeostasis in roots, while downregulating negative regulators of metal uptake. The lipopeptide production reshaped the composition of root-associated bacterial communities by increasing the relative abundance of putatively beneficial taxa and altering overall rhizosphere community structure. Functional genes indicated that lipopeptide enrichment was associated with microbial pathways related to metal transport, nutrient acquisition, and stress adaptation, suggesting that lipopeptides modulate both plant and microbial traits relevant to nutrient cycling. These results provide new insight into the ecological roles of B. velezensis-derived metabolites in plant-associated microbiomes and highlight lipopeptides as promising targets for harnessing beneficial plant–microbe interactions in sustainable agriculture.
|
|
Scooped by
mhryu@live.com
Today, 1:30 AM
|
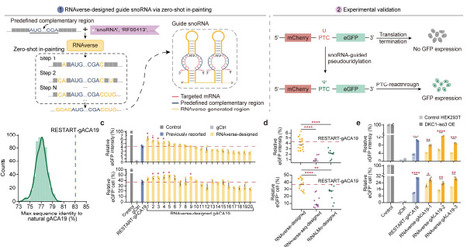
RNA engineering is bottlenecked by the tight coupling among sequence, structure, and function, leaving the field reliant on heuristics or screening. We introduce RNAverse, a multimodal generative model that learns a conditionable design grammar from RNA sequences, experimental structural probing profiles, and functional annotations. Beyond setting a new state of the art across 16 predictive modeling tasks, RNAverse inverts into generative design across four distinct RNA classes through three regimes, requiring experimental testing of no more than 20 candidates per-task. Zero-shot in-painting generates snoRNAs that promote premature termination codon (PTC) readthrough, and chimeric aptazymes for ligand-controlled gene regulation, exceeding rational design or large-scale screens. In both settings, RNAverse outperforms family-adapted sequence-only baselines. Fine-tuned in-painting extends to an unseen functional class, yielding IGFBP3-binding aptamers with picomolar affinity that outperform SELEX winners. Iterative optimization engineers 5′ UTRs that outperform clinical benchmarks, enhancing protein production in cell-based assays for both the Zolgensma gene therapy and the BNT162b2 COVID-19 vaccine, and boosting antibody responses to BNT162b2 in mice. Together, these results establish multimodal generative learning as a route from predictive RNA modeling to programmable specification of functional RNAs.
|
|
Scooped by
mhryu@live.com
Today, 1:18 AM
|
Rare earth elements (REE) are increasingly mobilized from mine residues and other industrial sources, creating a need for rapid screening of chemically complex waters where conventional ICP-based analysis is impractical on site. Here, we evaluate a lanthanide-gated enzymatic switch biosensor, for matrix-aware monitoring of mine-influenced waters. We quantified REE-dependent activation and systematic interference from co-occurring metals and anions typical of mine waters, and defined operating windows that preserve lanthanide responsiveness. REE activation was readily detectable at ~100 nM, whereas inhibition emerged in the low-micromolar range and became pronounced near ~10 µM for key interferents, corresponding to a practical tolerance of roughly 1:100 (REE:inhibiting element). Inhibition was strongest for AlIII, FeIII, GaIII, ZnII, and SnII. Combined machine learning, molecular dynamics, and density functional theory analyses indicate distinct inhibition modes, including coordination of FeIII, ZnII, and SnII at the LanM EF4 hand loop and favorable binding of [Al(OH)4]-/[Ga(OH)4]- near the TEM-1 substrate-recognition region. Consistent with these predictions, citrate reduced inhibition from hydrolysing metals, especially AlIII and SnII. Together, these findings define matrix-dependent operating envelopes, mechanistic design rules, and practical mitigation strategies for deploying REE biosensors in chemically complex waters.
|
|
Scooped by
mhryu@live.com
Today, 1:12 AM
|
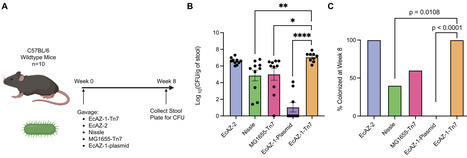
Native bacteria are strains isolated from the host and often require greater development and optimization of genetic manipulation tools than lab-adapted strains. We have previously shown that the native murine E, coli strain EcAZ-1, which colonizes the conventional mouse gut long term, is genetically tractable, and the engineered strain can elicit physiological effects while residing in the mouse gut. Given the interest in using EcAZ-1 and other native E. coli as a chassis for heterologous gene expression in the mouse gut, researchers need more efficient tools to rapidly and reliably introduce heterologous gene expression into the strain. Here, we demonstrate that a gene expression cassette can be integrated into the EcAZ-1 genome with high efficiency using the site-specific Tn7 transposon. Furthermore, we demonstrate that EcAZ-1 modified to express sfGFP using this method stably engrafts in the gut of conventional mouse. Taken together, Tn7-mediated integration is an effective method for stably introducing genetic cassettes into the native murine strain EcAZ-1 for expression in the mouse gut.
|
|
Scooped by
mhryu@live.com
Today, 1:07 AM
|
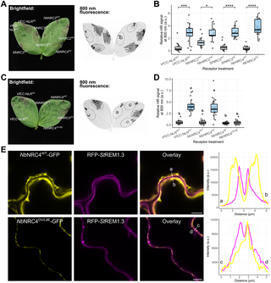
Agrobacterium-mediated transient expression has revolutionised plant research, enabling numerous landmark discoveries across diverse areas of plant biology. Yet this powerful approach remains largely confined to solanaceous species, leaving most economically important crop families without a comparable rapid assay platform. Here, we show that an engineered Rhizobium rhizogenes strain, AS109, mediates efficient transient expression across diverse dicot species spanning multiple taxonomic families, consistently outperforming commonly used laboratory agrobacterial strains. Leveraging the broad host range of AS109, we establish a suite of functional assays in faba bean (Vicia faba), including protein localization, RNA interference-mediated gene silencing, cell-surface elicitor recognition screens, nucleotide-binding leucine-rich repeat receptor (NLR) activation, and infection cell biology at the host-pathogen interface. We further demonstrate that both singleton NLRs and sensor-helper NLR pairs from Solanaceae retain effector recognition and cell death activity when transferred into faba bean, establishing a rapid platform for evaluating cross-family transferability of disease-resistance genes. AS109 thus provides an accessible and versatile chassis for functional genomics in non-model crops, bridging the widening gap between hypothesis generation and experimental validation across diverse plant species.
|
|
Scooped by
mhryu@live.com
Today, 12:45 AM
|
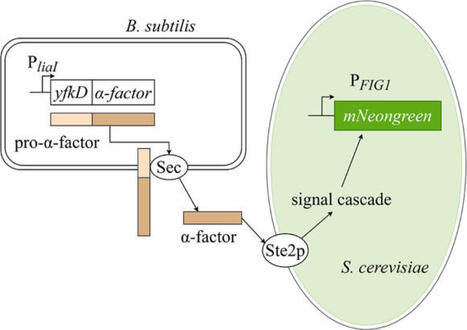
Microbial communities found in nature are complex and thrive in harmony, consisting of organisms derived from several kingdoms. Hence, they need to be able to communicate by quorum-sensing molecules to co-exist in close proximity. Being able to engineer artificial inter-kingdom communication for biotechnological approaches, such as degradation of complex biomaterials in one fermenter using controlled signaling between microorganisms, would be beneficial. Therefore, we wanted to establish a proof of principle by enabling the Gram-positive bacterium Bacillus subtilis to communicate in a controlled manner with the eukaryotic yeast Saccharomyces cerevisiae by exploiting the yeast’s pheromone system. We genetically modified B. subtilis to heterologously produce the α-pheromone after bacitracin induction, which in turn would activate the production of the green fluorescent protein mNeongreen upon pheromone recognition by S. cerevisiae. Co-cultivation in modified growth medium and induction of pheromone expression was successful and repeatable. Yeast activation and production of mNeongreen was visible not only by analysis but also by epifluorescence microscopy of the co-culture. This study successfully demonstrated controlled interkingdom signaling between microorganisms by leveraging their native communication systems.
|
|
Scooped by
mhryu@live.com
Today, 12:36 AM
|
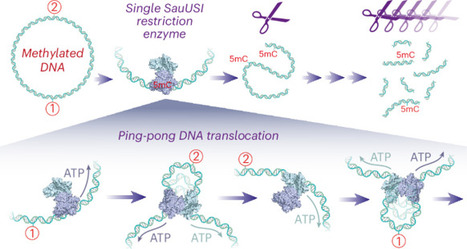
SauUSI is a dimeric, ATP-dependent type IV restriction enzyme that protects Staphylococcus aureus by cleaving nonself DNA containing 5-methylcytosine or 5-hydroxymethylcytosine. Using biophysical, single-molecule and nanopore sequencing methods, we show that 5-methylcytosine recognition first induces ATP-driven unidirectional translocation by one helicase-like subunit, displacing its target recognition domain (TRD). In half of events, the partner TRD can then bind the liberated modified site, stabilizing a growing DNA loop. On symmetrically methylated DNA, looping is more frequent and both motors can engage in bidirectional loop translocation. Cleavage occurs stochastically throughout translocation but events that increase dwell time such as rebinding methylated sites elevate the chance of cleavage. Interactions with other SauUSI dimers or protein roadblocks influence cleavage distributions but are not required for nuclease activation. While the first cleavages principally generate 1-nt 3ʹ overhangs or blunt ends, multiple binding–translocation–cleavage cycles by individual enzymes ultimately fragment the modified nonself DNA, neutralizing its threat. Using single-molecule physics and biochemical assays, Shaw, Hughes-Games and Torres Montaguth et al. reveal how the bacterial defense enzyme SauUSI repeatedly scans and makes multiple DNA breaks, efficiently fragmenting invading genetic material.
|
|
Scooped by
mhryu@live.com
Today, 12:20 AM
|
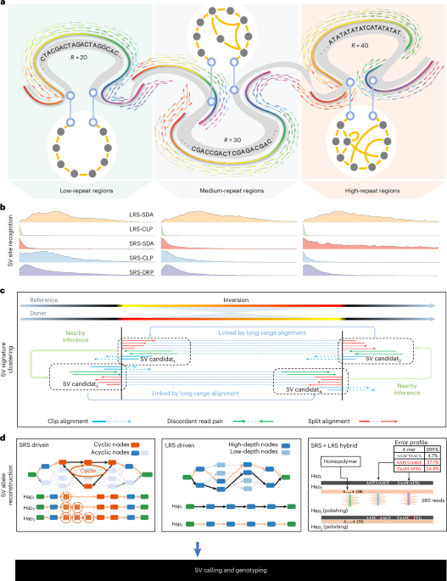
Structural variants (SVs) are a major source of genomic diversity, yet their discovery remains challenging due to repetitive genomic contexts, alignment ambiguity and the trade-off between sequencing cost and read length. Here we introduce HitSV, which implements repetitiveness- and signature-density-aware breakpoint recognition coupled with precise haplotype-resolved local assembly, thereby enabling base-resolution SV reconstruction and genotyping in various sequencing technologies. On the HG002 benchmark, HitSV achieved 15–72% (long read), 10–36% (short read) and 12% (hybrid sequencing), respectively, higher genotype F1 scores than state-of-the-art SV callers. Applied to the 1KGP phase 4 cohort, HitSV expanded the SV callset by 31.5%, substantially reshaping allele-frequency landscapes. In a large Chinese long-read cohort, HitSV uncovered tandem-repeat-mobile-element composite arrays as a prevalent, multi-allelic class of complex SVs, highlighting composite repeat architectures as an important and previously underrecognized contributor to human structural variation. HitSV models the interaction between genomic repetitiveness and signature intensity to improve structural variant detection.
|
|
Scooped by
mhryu@live.com
Today, 12:17 AM
|
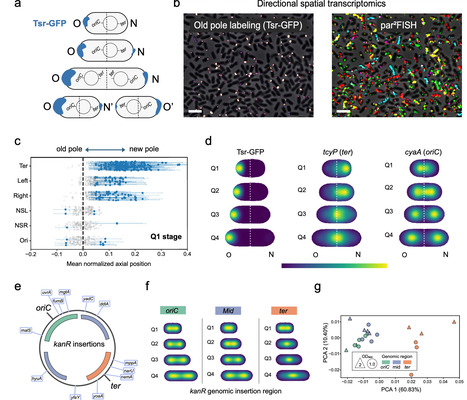
Elucidating the intracellular organization of bacterial transcriptomes may reveal new modes of gene regulation. Recent work reported that chromosomal gene position determines mRNA localization within E. coli cells, implying a widespread genome-guided organizing mechanism. Yet this interpretation contrasts with biophysical expectations based on translation and diffusion and may be confounded by nascent transcription. Here, we examine this phenomenon by analyzing intracellular spatial transcriptomics datasets spanning different methods, bacterial species, growth conditions, and thousands of genes. We confirm the presence of genome-associated localization signatures that track chromosome macrodomain dynamics across the cell cycle and establish causality by repositioning a gene at distinct chromosomal loci. Quantitative analysis, however, shows that these signatures generally involve only a minority of transcripts and are of magnitudes consistent with reported rates of nascent transcription. Moreover, genome-associated patterns preferentially occur in weakly translated and unstable mRNAs. Together, our results indicate that genome-guided localization is unlikely to be a dominant driver of transcript positioning in bacteria. Genome organization has been previously proposed to dictate mRNA localization in bacteria. Here, the authors use spatial transcriptomic analysis to show that the genomic effect involves only a minority of transcripts, mainly weakly translated and unstable mRNAs.
|
|
Scooped by
mhryu@live.com
October 15, 11:55 PM
|
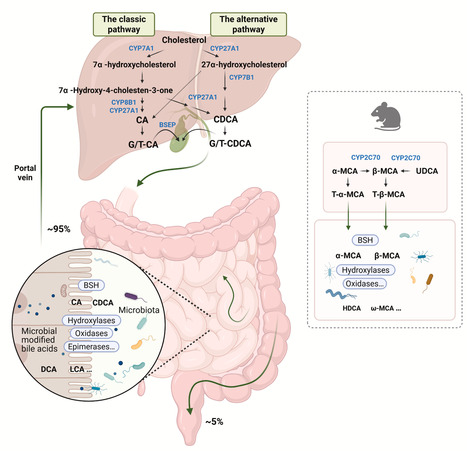
Perturbation of microbial-derived bile acid (BA) metabolism is strongly linked to the pathogenesis of intestinal diseases, including inflammatory bowel disease, enteric infections, and colorectal cancer. Recent advances in artificial intelligence-assisted pipelines have enabled the identification of BA modification reactions, which have uncovered novel targets and the mechanisms contributing to intestinal diseases. In this review, we summarize recent studies on microbial enzymatic pathways and modifications of BAs, and their roles in promoting or preventing intestinal diseases. The mechanisms altering gut microbiota populations and microbial BA metabolic landscapes that influence intestinal diseases are further examined along with how microbial-derived BAs modulate intestinal immune responses. Based on these advances, we suggest different dietary patterns that reshape the gut microbiota community and BAs to prevent intestinal diseases. We further discuss current challenges in basic and translational research on microbial BA modifications in intestinal homeostasis and propose future directions to drive progress in this field.
|
|
Scooped by
mhryu@live.com
October 15, 11:03 PM
|
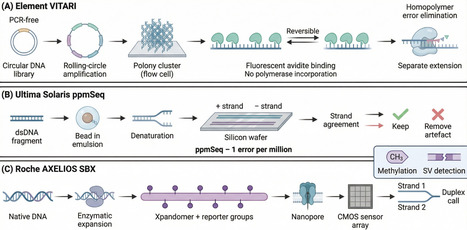
In 2026, three high-throughput sequencing platforms approached the widely publicised sub-$100 30× human genome. Behind these figures, which reflect consumable prices rather than total cost of ownership, the more consequential shift is the end of single-vendor dominance: these systems make comprehensive genomic profiling and multi-omic liquid biopsy feasible in small batches.
|
|
Scooped by
mhryu@live.com
October 15, 6:14 PM
|
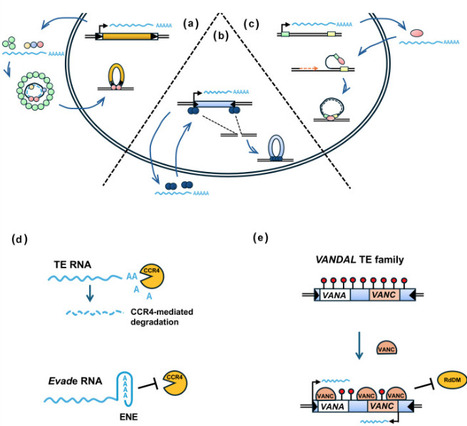
Transposable elements (TEs) constitute a major fraction of eukaryotic genomes and represent one of the most abundant classes of repetitive sequences in plants. Recent advances in sequencing technologies, together with the increasing availability of telomere-to-telomere and pan-genome assemblies, have enabled comprehensive characterization of TE sequences, revealing their complex architectures, structural variation, and evolutionary dynamics across diverse plant species. Because TE mobilization can compromise gene function and genome integrity, plants have evolved sophisticated epigenetic mechanisms to restrict TE activity. In turn, TEs have co-evolved diverse strategies to evade host silencing, reflecting a continuous evolutionary arms race between host genomes and their mobile elements. Beyond their traditional view as genomic parasites, accumulating evidence now demonstrates that TEs are major drivers of genome innovation by providing novel protein-coding modules, cis-regulatory elements, and structural variation that reshape chromatin organization and gene regulatory networks. In this review, we highlight recent advances in our understanding of how TEs promote regulatory innovation, environmental adaptation, and genome evolution in plants.
|