Your new post is loading...

|
Scooped by
mhryu@live.com
Today, 1:47 PM
|
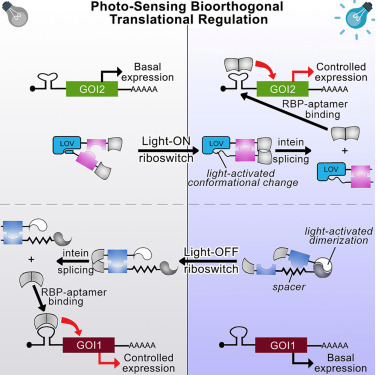
Photoriboswitches are invaluable tools for synthetic biology because they provide direct, precise, and on-demand control of translation. While most existing photoriboswitches are predominantly one-directional, this study presents a bidirectional photoriboswitch platform capable of mimicking the orthogonal RNA-binding protein-mediated up-/downregulation of multiple mRNAs. Inspired by intein-based split protein reconstitution and light-induced protein dimerization, we first engineered a Light-OFF system that uses photodimerization to block intein reconstitution. We then validated that this Light-OFF system can function orthogonally with a Light-ON system that reconstitutes a different split intein pair under light. Together, a bidirectional photoriboswitch is generated, which senses light to orthogonally reconstitute RNA-binding proteins to tune the translation of their respective target mRNAs. We have demonstrated that the photoriboswitch produces bidirectional translation regulation. Importantly, the functional modules of the photoriboswitch can be easily altered to adapt for different purposes, allowing the bidirectional photoriboswitch to serve as a versatile tool for translation regulation.
|
|
Scooped by
mhryu@live.com
Today, 1:40 PM
|
The antibiotic pipeline is in crisis, even though microbial natural products, account for most approved antibacterial drugs and retain vast, untapped biosynthetic potential. Genuinely new scaffolds are rare because discovery remains constrained to a narrow set of cultivable taxa and expressed biosynthetic gene clusters, while most microbial chemistry persists as “dark matter” in uncultured organisms and silent clusters. Each bottleneck is both a discovery and a biotechnology challenge: a predicted cluster must be expressed, its product isolated, and its biosynthesis engineered to workable titres before any hit can realistically progress. This review links the major obstacles in the contemporary pipeline including limited ecological sampling, silent and cryptic clusters, access to uncultured bacteria, inefficient metabolome mining, weak genome–metabolome integration, and incomplete target identification, to the technologies best suited to address them, drawing on recent case studies of integrated workflows rather than single-platform approaches. By evaluating these strategies for both discovery power and bioprocess tractability, we outline a practical framework for turning microbial dark matter into producible, mechanistically understood antibiotic leads.
|
|
Scooped by
mhryu@live.com
Today, 1:24 PM
|
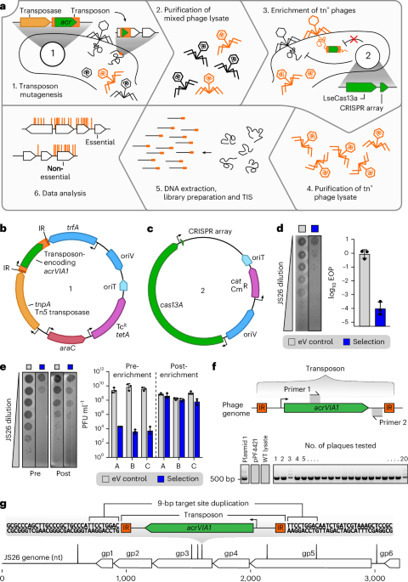
Phages are important drivers of bacterial evolution and have potential as antimicrobials. However, incomplete understanding of phage biology and our inability to rapidly engineer them with new genetic cargo hinder progress towards phage-based therapies. Here we develop an unbiased, genome-wide mutational tool for phages. This approach, phage Tn-seq, uses Tn5 transposon mutagenesis using anti-CRISPR-based selection and deep-sequencing. Phage Tn-seq was successful across diverse phages, including a nucleus-forming jumbo phage and enabled gene essentiality assignment consistent with phage structural proteomics and core gene conservation. In addition, insertion biases allowed prediction of transcriptional direction and early injected and highly expressed regions. We exploit transposons to deliver new cargo to phages within a few days and created an orthogonal artificial intelligence-designed Acr transposon system to generate phage double mutants. Transposon insertion was also achieved in phages with hypermodified DNA. Phage Tn-seq is a versatile tool to advance our understanding and application of phages. Phage Tn-seq uses Tn5 transposon mutagenesis with anti-CRISPR-based selection and deep sequencing as a method to generate unbiased, genome-wide mutations across diverse phages
|
|
Scooped by
mhryu@live.com
Today, 12:34 PM
|
Diverse chemicals biosynthesis demand distinct precursor acetyl-CoA and energy inputs, yet strategies balancing carbon atom economy and energy generation for varied products remain scarce. Here, we design an autonomous hybrid energy-carbon balancing apparatus (ATHENA) to establish a paradigm for redox-driven, dynamically adaptive regulation of carbon conservation and energy synthesis in E. coli. ATHENA features modularization of acetyl-CoA and NADH synthesis pathways for carbon conservation and energy generation, paired with NADH-responsive positive and negative genetic circuits constructed using the BsRex sensor and antisense RNA. This system enables intracellular redox state-dependent rational metabolic flux allocation to balance carbon atom economy and energy generation. Ultimately, ATHENA significantly improves yields and titers of five products with differential acetyl-CoA and energy demands. In the 5-L bioreactor, acetate yield markedly exceeds its native theoretical yield, while that of mevalonate nearly approaches this theoretical upper limit. This work provides a universal strategy for constructing high carbon-yield microbial chassis cells. Metabolic engineering strategies that balance carbon atom economy and energy generation remain scarce. Here the authors engineer E. coli with an NADH-responsive genetic circuit to regulate carbon-conserving and energy-generating modules, improving yields and titres of diverse products.
|
|
Scooped by
mhryu@live.com
Today, 12:24 PM
|
Nitrification is a key process in the global nitrogen cycle, yet the physiological characterization of nitrifiers remains limited due to their low culturability. Although culture-independent approaches have provided important ecological insights, they do not fully resolve the physiological properties of nitrifying microorganisms. Here, we show that common gelling agents (agar, agarose, and gellan gum) contain toxic substances that inhibit the growth of diverse nitrifying microorganisms as well as some heterotrophic bacteria. These inhibitory substances appear to be generated during autoclaving and act via mechanisms distinct from previously reported hydrogen peroxide. Removal of these inhibitory factors using a simple washing strategy enables colony formation in recalcitrant nitrifiers, including Nitrososphaera (ammonia-oxidizing archaea) and Nitrospira (nitrite-oxidizing bacteria). Application of this approach to environmental samples enabled the isolation of multiple difficult-to-culture nitrifying strains, including ammonia-oxidizing and nitrite-oxidizing microorganisms. Together, these findings demonstrate that the limited culturability of nitrifiers on solid medium is largely driven by gelling agent–derived inhibitory substances and can be overcome by a simple and broadly applicable strategy.
|
|
Scooped by
mhryu@live.com
Today, 11:36 AM
|
Selective DNA delivery to specific members of assembled bacterial communities remains challenging. Bacterial conjugation enables efficient DNA delivery, but transfer to non-target recipients limits its specificity within mixed communities. Here, we repurpose plasmid entry exclusion (Eex) as a recipient-side gate to control conjugative DNA delivery. We demonstrate selective plasmid delivery to Eex-negative recipients within populations containing both Eex-expressing and Eex-negative cells. This recipient selectivity was maintained at increased cell densities and during prolonged mating. By combining RP4-type and F-type conjugation systems with their corresponding exclusion modules, we further directed DNA delivery from distinct donors to defined recipient populations. RP4-derived Eex also functioned in environmental bacteria, including Pseudomonas putida and Sphingobium japonicum, enabling recipient-specific exclusion within a multispecies mixture. Furthermore, repeated cycles of Eex-guided conjugation and selection altered community composition after assembly. These results establish entry exclusion as a recipient-side strategy for selective conjugative DNA delivery and, when combined with selection, for controlling the composition of assembled bacterial communities.
|
|
Scooped by
mhryu@live.com
Today, 11:24 AM
|
Fungal biotechnology has emerged as a promising platform for the development of sustainable biocomposites, leveraging the intrinsic ability of fungi to transform complex polymeric substrates into structurally integrated materials. This review critically evaluates mycelium-based biocomposites (MBCs), with particular emphasis on the relationships between fungal biosynthesis, substrate transformation, processing strategies, and resulting material properties. Key biochemical components, including chitin, chitosan, and β-glucans, are examined in terms of their molecular structures, biosynthetic pathways, and contributions to composite performance. The chemical modification of polymeric substrates during fungal colonization, including enzymatic degradation, substrate remodelling, and interfacial bonding mechanisms, is critically discussed. The review further examines chemical functionalization, hybrid reinforcement, and densification strategies for tailoring mechanical performance, thermal insulation, fire resistance, and durability, highlighting recent advances in processing-driven material optimization. Applications in packaging, construction, insulation, environmental remediation, and functional materials are critically reviewed alongside sustainability considerations, including biodegradability, circularity, and life-cycle impacts. Finally, current challenges and future research directions are discussed, emphasising the integration of synthetic biology, advanced materials chemistry, and digital bio-fabrication to enable scalable, high-performance, and multifunctional MBCs.
|
|
Scooped by
mhryu@live.com
Today, 10:57 AM
|
Yeasts serve as premier eukaryotic hosts for microbial cell factories, enabling the production of recombinant proteins, biofuels, and high-value natural products. Precise transcriptional control is paramount for balancing complex metabolic pathways and maximizing target product yields. Promoters, as the key regulatory elements governing transcription initiation, have thus become a focal point of metabolic and synthetic biology engineering. This review surveys promoter architecture, function, and engineering in yeast. It outlines promoter structure, from core elements to upstream regulatory sequences, and reviews current and emerging strategies for discovery, optimization, and de novo design, including random mutagenesis, combinatorial assembly, rational design based on transcription factor binding sites, intron-mediated enhancement, cross-species approaches. Importantly, this review highlights the rapidly growing role of machine learning (ML) and artificial intelligence (AI) in promoter engineering, from predictive modeling of promoter strength to in silico generation of synthetic variants. Special emphasis is placed on synthetic promoter engineering to yield tunable and inducible toolkits. The review also highlights applications in yeast cell factories, such as metabolic pathway optimization, dynamic regulation, high-throughput strain screening, and recombinant protein production. By summarizing these recent advances, it serves as a practical resource for researchers aiming to achieve precise metabolic control and improve bioproduction outcomes in yeast.
|
|
Scooped by
mhryu@live.com
September 25, 4:35 PM
|
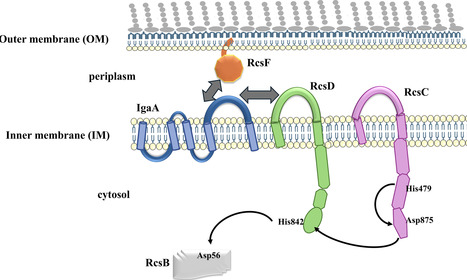
Two-component systems (TCS) formed of sensor histidine kinases (HK) and response regulators (RR) are the primary prokaryotic signal transduction pathways. In Enterobacteriaceae, some TCS evolved architectural complexity, by integrating auxiliary proteins into the HK– RR dyad. The Regulator of Capsule Synthesis (Rcs), a conserved phosphorelay system in Enterobacteriaceae, recruits several auxiliary proteins, including an outer membrane lipoprotein, RcsF, which senses cell envelope threats and transduces the signal to downstream components. A second auxiliary protein located in the inner membrane, RcsD, transfers the phosphoryl group from the hybrid histidine kinase RcsC to the response regulator RcsB. RcsD is formed of a large periplasmic domain (Asn43-Asn308) sandwiched between two transmembrane helices and a cytosolic domain. For RcsD, the role and evolutionary benefit of maintaining a large periplasmic domain while the main phosphostransfer reaction occurs in the cytosol is not clear. Here, we show that RcsF and RcsD exhibit strong co-evolutionary coupling independent of host speciation (partial Mantel r = 0.950, p < 0.0001). Structure-guided docking and 500-ns molecular dynamics simulations predicted a stable interaction between RcsF and RcsD (MM-GBSA ΔG ≈ –95 kcal/mol), suggesting their functional coupling. Functional assays in Escherichia coli K12 MG1655 revealed that RcsD periplasmic domain represses basal Rcs activity in unstressed cells (~5-fold derepression) yet is essential for stress-induced activation (1.9–2.5-fold). Overexpression of rcsF beyond the chromosomal level induces Rcs signal activation only when the RcsD periplasmic domain is present. These findings demonstrate how co-evolution of an auxiliary protein generates novel regulatory function in the Rcs phosphorelay and provide insights into the role of RcsD periplasmic domain in stress signal transduction.
|
|
Scooped by
mhryu@live.com
September 25, 4:28 PM
|
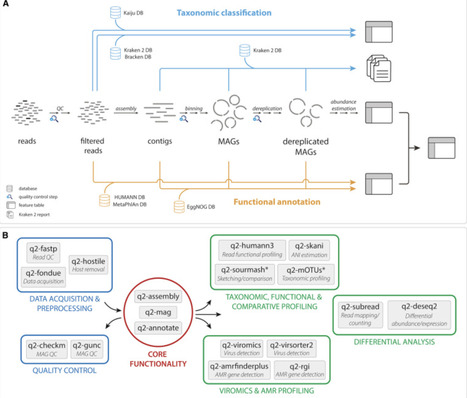
Metagenome sequencing has revolutionized functional microbiome analysis across diverse ecosystems but is fraught with technical hurdles. We introduce MOSHPIT (MOdular SHotgun metagenome Pipelines with Integrated provenance Tracking; https://moshpit.qiime2.org)—software built on the QIIME 2/rachis framework (Q2F) that integrates best-in-class CAMI II- and LEMMI-validated metagenome tools with robust provenance tracking and multiple user interfaces—enabling streamlined, reproducible metagenome analysis for all expertise levels. By building on Q2F, MOSHPIT enhances scalability, interoperability, and reproducibility in complex workflows, democratizing and accelerating discovery at the frontiers of metagenomics.
|
|
Scooped by
mhryu@live.com
September 25, 4:14 PM
|
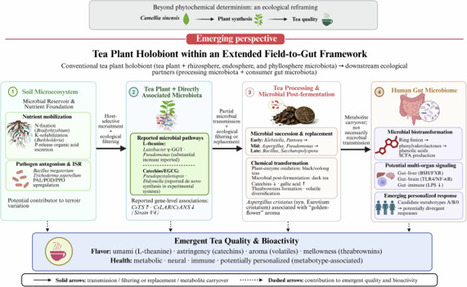
Across cultivation, processing, and consumption, tea quality and bioactivity are shaped by plant-microbiota interactions. This review develops a field-to-gut framework centered on the tea plant holobiont and downstream microbial partners. Within that frame, rhizosphere and endophytic microbiota are examined as regulators of flavor precursors, host metabolism, and defense. Processing microbiota are considered for remodeling tea chemistry and gut microbiota for bioactivating tea compounds, with microbial transmission, ecological replacement, and metabolite carryover distinguished and translational directions identified in microbiome-guided cultivation, controlled fermentation, and metabotype-informed nutrition.
|
|
Scooped by
mhryu@live.com
September 25, 3:19 PM
|
FAIDR-Desi is a non-technical user interface for design of intrinsically disordered regions (IDRs) based on greedy optimization of sequence features (simple sequence statistics correlated with IDR function, including amino acid composition, residue patterning and short linear interaction motifs), using more features (127) than other IDR design approaches. FAIDR-Desi can be used to design synthetic IDR mimics to establish feature sufficiency for function, or to knock out features to show necessity for function, exemplified with experiments on the yeast Cox15 mitochondrial targeting signal.
|
|
Scooped by
mhryu@live.com
September 25, 2:55 PM
|
Protein synthesis forms the very basis of life. Cells rely on a sophisticated machinery of factors for the translation of an RNA template into a correct amino acid sequence. We owe a lot of our understanding of this process to specific small-molecule modulators. Since the discovery of antibiotics inhibiting bacterial protein synthesis, small molecules have served as probes to dissect the translation apparatus and as medications to fight disease. Advances in sequencing-based techniques and structural methods now provide for a highly detailed mechanistic understanding. Recent evidence shows that many translation modulators act in an RNA- or a peptide-sequence-selective manner. These molecules present exciting opportunities to better understand the translation machinery and provide new venues to modulate the expression of certain protein subpopulations. In this Review, we summarize recent findings on sequence-selective translation modulators, with emphasis on eukaryotes, the techniques that enabled their discovery, and offer an outlook into their potential applications. Small-molecule modulators of protein synthesis are invaluable tools for dissecting the translation apparatus. This Review summarizes the mechanisms of molecules that disrupt translation in an amino acid or mRNA sequence-dependent manner, with focus on eukaryotic systems. It discusses relevant techniques used in their characterization and offers perspectives into their potential applications.
|
|
|
Scooped by
mhryu@live.com
Today, 1:44 PM
|
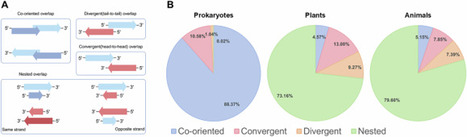
Overlapping genes are widely distributed across the genomes of nearly all living organisms. Although thousands of overlapping genes have been identified and studied in several species, a comprehensive annotation of these genes is lacking, and the forces shaping their evolution remain poorly understood. Here, we present a comparative analysis of overlapping genes in 253 prokaryotic genomes, 115 plant genomes, and 111 animal genomes. We find that co-oriented overlapping genes predominate in prokaryotes, whereas nested overlapping genes are the most common type in eukaryotes. In prokaryotes, overlapping genes are significantly shorter than non-overlapping genes, and 69% of co-oriented overlaps occur in translational phase 2. The number of overlapping genes is strongly correlated with genome size in prokaryotes but not in eukaryotes. We estimated the rates at which overlapping genes are gained and lost over evolutionary time and found that loss of overlapping orthologs is the highest rate in eukaryotes, whereas the highest rate in prokaryotes is the gain of overlapping genes from adjacent genes. Expression analyses revealed that, compared with neighboring genes, nested genes exhibit lower expression levels, altered expression correlations, and higher tissue-specific expression patterns in selected eukaryotic species. Based on these patterns, we propose distinct models to explain the evolutionary pressures acting on overlapping genes in prokaryotes and eukaryotes. Together, our findings provide new insights into the evolution of overlapping genes.
|
|
Scooped by
mhryu@live.com
Today, 1:28 PM
|
Adaptive laboratory evolution (ALE) is a powerful strategy for exploring functional solution spaces, yet its pace is fundamentally constrained by low spontaneous mutation rates. Here, we present a broadly applicable genetic device that accelerates genome diversification in Gram-negative bacteria through the transient action of a dual base editor (ACBE3). This construct combines cytidine and adenosine deaminase activities with a single-stranded DNA-binding module, enabling efficient genome-wide mutagenesis without reliance on host-specific replication or repair systems. The expression of ACBE3 increased mutation rates by up to four orders of magnitude in E. coli and Pseudomonas putida, outperforming classical random mutagenesis approaches such as UV irradiation and chemical treatment. Whole-genome sequencing revealed extensive and uniformly distributed mutations, predominantly consistent with deamination signatures, as well as a broader spectrum of nucleotide changes and small indels. The system displayed strong orthogonality, functioning across diverse bacterial taxa. As a proof of concept, coupling ACBE3-mediated diversification with selective growth conditions enabled the rapid evolution of Acinetobacter calcoaceticus P320, yielding mutant strains with enhanced growth on n-dodecane within only a few induction cycles. This approach decouples diversification from selection, enabling the controlled exploration of adaptive landscapes at unprecedented speeds. While high mutational loads impose cellular burdens and require tight regulatory control, the platform offers a versatile route to accelerate microbial evolution at both single-strain and community levels. Our results thus document dual base editing as a powerful framework for fostering evolution in microbial biotechnology.
|
|
Scooped by
mhryu@live.com
Today, 12:49 PM
|
Microbial CO₂ assimilation provides a strategy for sustainable biomanufacturing. However, CO₂ reduction requires substantial energy input, while native energy systems are tightly regulated and extensively consumed by endogenous metabolism, limiting assimilation efficiency. Here, we design and construct an orthogonal energy system in E. coli, achieving an intracellular NUDH concentration exceeding 7.0 mM and an NUDH/NUD⁺ ratio above 55. Subsequently, we design a CO₂-formate-acetyl-CoA-pyruvate-malate pathway and reprogram key enzymes for NUDH-dependent operation. Integration of the OES-driven CAM* pathway with a rationally rewired native electron transport chain supports growth of the engineered E. coli on CO₂ and methanol, with a doubling time of 8.6 ± 0.3 h and a maximum OD₆₀₀ of 37.5 ± 5.8. Moreover, the platform extends CO₂ assimilation to biosynthesis of multiple chemicals from distinct CAM* pathway nodes, including mevalonate, lactate, 2,3-butanediol, and succinate. Together, these results establish a framework for reprogramming microbial energy metabolism and expanding one-carbon biotransformation. Microbial CO₂ reduction requires substantial energy input which limits assimilation efficiency. Here the authors create a dedicated reducing-power supply for CO₂ assimilation in E. coli and couple this with methanol oxidation to support the production of four chemicals from distinct metabolic nodes.
|
|
Scooped by
mhryu@live.com
Today, 12:27 PM
|
Carbohydrate-active enzymes (CAZymes) drive the turnover of polysaccharides in the oceans. However, the scale of diversity in polysaccharide utilization and genome organization strategies has yet to be characterized, particularly in marine ecosystems. In this paper, we introduce mpcgcdb.com as an interactive web catalog for marine CAZyme gene clusters, their associated metagenome-assembled genomes, and relevant enzyme families found in marine metagenomic samples. This database contains nearly 290,000 marine CAZyme gene clusters from more than 22,000 genomes, enabling user-initiated exploratory visualizations for genome organization networks and enzyme phylogeny. This tool will assist researchers in developing hypotheses about gene function, connecting species to metabolic niches, and identifying gaps where future work is needed to understand different pathways in marine glycan cycling.
|
|
Scooped by
mhryu@live.com
Today, 12:18 PM
|
Maintaining microbially mediated phosphorus mobilization under intensive nitrogen fertilization is difficult in acidified paddy soils. Using a decade-long field experiment in a rice–oilseed rape rotation, we compared an ordered manure-substitution series, in which manure input increased while mineral nitrogen input decreased, with a mineral nitrogen dose gradient. Increasing mineral nitrogen input was associated with lower diversity and lower absolute abundance of the phoD-harboring bacterial guild and a marked shift in community composition. Across the manure-substitution series, soil pH increased from 5.94 to 6.62, potential alkaline phosphatase activity increased by up to 158%, and Olsen-extractable phosphorus increased by up to 182%. The diversity, composition, and absolute abundance of the phoD guild changed progressively across the same series. Distance-based redundancy analysis identified soil pH as the strongest measured marginal correlate of community composition, whereas variation partitioning indicated that the effects of pH, carbon, and phosphorus were largely shared. An exploratory structural equation model was consistent with conditional associations of both community composition and absolute gene abundance with alkaline phosphatase activity, which was, in turn, associated with Olsen-extractable phosphorus. The highest observed means for gene abundance, grain yield, and grain phosphorus removal occurred in the intermediate manure treatment, whereas treatment-level copper and zinc concentrations increased towards the highest-manure treatment. Together, these findings indicate that manure substitution can counteract mineral nitrogen-associated deterioration of the phoD guild and support microbial phosphorus mobilization, although the agronomic and metal responses require confirmation across sites and seasons.
|
|
Scooped by
mhryu@live.com
Today, 11:30 AM
|
Programmable genome editing has advanced from nucleases through base and prime editors, yielding an increasingly precise and versatile editing toolkit that is nonetheless best suited to small, localized changes. Efficient, site-specific writing of kilobase-scale DNA, including whole genes, regulatory elements, and multi-component cassettes, remains a central unmet goal, with implications for dissecting regulatory logic, building physiologically accurate disease models, and engineering mutation- and gene-agnostic therapies. Here, we review recent advances in programmable large-DNA writing in the human genome across four mechanistic categories: recombinase-, prime editing-, transposase-, and retrotransposon-mediated technologies. Across these platforms, high catalytic activity and precise control over the integration site have tended to trade off against one another, a tension further compounded by declining efficiency from immortalized lines to primary cells to in vivo models. We also discuss the key limitations facing efficient ex vivo and in vivo large-DNA writing applications, and consider how current and future advances in biomolecule evolution and delivery systems can help address these challenges.
|
|
Scooped by
mhryu@live.com
Today, 11:01 AM
|
Decolorization remains a major challenge in the treatment of dye-containing wastewater. Here, we engineered E. coli with a chromosomally integrated minimal Psl polysaccharide synthesis operon, pslA-G, to create a Psl biofilm-based living bioflocculant enabling autonomous dye capture, sedimentation, and controllable recovery. The engineered bioflocculant exhibited rapid self-flocculation, achieving approximately 85% sedimentation within 8 h. The bioflocculant mediated the removal of Reactive Red 24, Acid Orange 7, and Congo red in a dye-dependent manner, achieving its highest removal efficiency of approximately 90% for Congo red. Importantly, dye-loaded aggregates could be disassembled via PslG-triggered Psl polysaccharide hydrolysis, enabling controlled dye release. This work establishes a proof-of-concept living bioflocculation approach that integrates capture, sedimentation, and recovery for potential dye wastewater treatment.
|
|
Scooped by
mhryu@live.com
Today, 10:37 AM
|
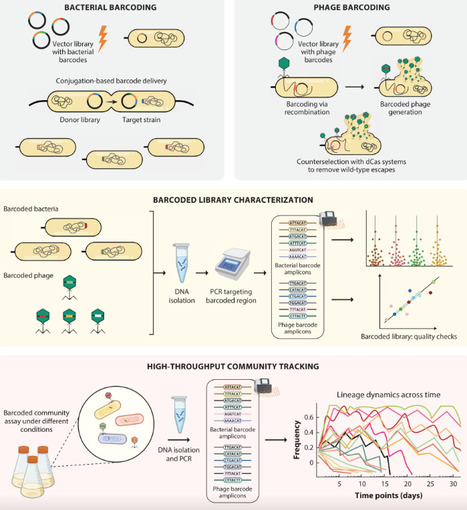
Bacteriophages, viruses that parasitize bacteria, hold tremendous potential as antimicrobial agents, microbiome modulators, and industrial biocontrol tools; yet clinical and environmental applications remain frustratingly inconsistent. Decades of research on isolated phage-bacteria pairs have revealed fundamental mechanisms governing infection specificity, coevolutionary arms races, and resistance trade-offs. These foundational studies, however, do not help predict outcomes when phages encounter multi-species assemblages characteristic of natural ecosystems. In this review, we integrate recent advances examining how interaction complexity shapes phage efficacy across four scales, gradually from simple phage-bacteria pairs to phage-bacterial communities. At every scale, emergent properties arise from complex interactions. Dissecting these dynamics requires technologies that can track multiple lineages simultaneously. DNA barcoding, which inserts unique genetic identifiers into bacterial and phage genomes, offers a promising solution. While barcoding all members in a synthetic community is unrealistic, we propose that even foundational reference sets of barcoded phage-bacteria pairs would enable systematic investigation of resistance evolution, competitive interactions, and functional outcomes in realistic contexts. Bridging laboratory insights and field performance demands integrating genetic engineering, high-throughput tracking, functional profiling, and predictive modeling into a coordinated research framework.
|
|
Scooped by
mhryu@live.com
September 25, 4:33 PM
|
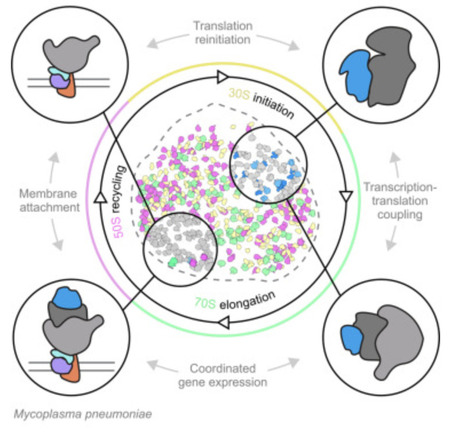
Translation is a central process in gene expression. Its regulation is complex, depends on factors that include cell state and the subcellular environment, and is subject to modulation via crosstalk to processes such as transcription or translocation. Here, we used cryo-electron tomography of native and antibiotic-perturbed Mycoplasma pneumoniae cells to resolve 140 maps that recapitulate bacterial translation during the initiation, elongation, and recycling phases. We visualized multiple transcription-translation complexes, allowing us to propose a threading-based translation reinitiation mechanism and to provide structural evidence for a long-hypothesized supercomplex that coordinates transcription, translation, and membrane attachment. We resolved abundant membrane-associated large ribosomal subunits and suggest that dissociation from membranes depends on the conditional initiation of new translation, consistent with a potentially conserved mechanism in mammalian cells. This work visualizes the multilayered control of bacterial translation and demonstrates the power of in-cell structural biology to investigate regulatory circuits in gene expression.
|
|
Scooped by
mhryu@live.com
September 25, 4:20 PM
|
A major goal in bottom-up synthetic biology is the construction of multicellular synthetic systems capable of coordinated and robust collective behaviours. However, robustness is often limited by noise and variability arising from increased molecular complexity. Whilst communication has been implemented in synthetic multi-cellular systems, the ability for communication to suppress cell free gene expression variability in populations of synthetic cells remain unexplored. To address this, we encapsulated the Lux and Las quorum sensing gene circuits in lipid vesicles under cell-free conditions to test the effect of communication on reducing cell-free gene expression variability across the population. Our results show that communication, limiting expression resources, and membrane surface effects can reduce gene expression variability. Resource limited Gillespie simulations for transcription and translation show that communication-mediated coupling reduces population-level expression noise under constrained and excess resource conditions. Together, our work provides simple strategies to reduce gene expression variability and thereby improve robustness in synthetic multicellular systems, an important criteria for the future applications of synthetic cells.
|
|
Scooped by
mhryu@live.com
September 25, 3:25 PM
|
Bacteria adapt to changing environments to ensure survival. Responding to new environmental conditions and internal states through regulation of gene expression is a fundamental biological process. One group of regulators of gene expression is regulatory RNAs. Among their many functions, there is a group of relatively short RNA molecules (‘small RNAs’, sRNAs) which typically modulate gene expression by base-pairing to a cognate target RNA or by directly binding to a protein, leading to a regulatory output. Although regulatory sRNAs have been studied in great detail in many medically important bacteria, we know much less about them in the World Health Organization priority pathogen Acinetobacter baumannii. In this review, I outline our current understanding and highlight recent progress of sRNA-mediated gene regulation in A. baumannii and provide an outlook for future areas to study.
|
|
Scooped by
mhryu@live.com
September 25, 3:18 PM
|
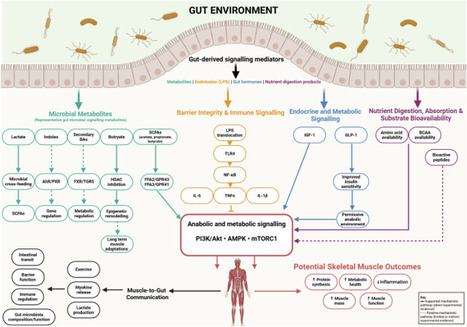
Skeletal muscle has a profound influence on metabolic health, functional capacity and resilience across the lifespan. Beyond dietary protein and physical activity, the gut microbiome may have the capacity to impact skeletal muscle mass and function through the bidirectional network termed the gut-muscle axis (GMA). Fermented protein foods (FPFs) are protein rich matrices transformed by microbial activity that integrate modified protein structures, bioactive peptides, and live microorganisms, or components thereof, which could modulate skeletal muscle physiology. Here, we review mechanistic, preclinical and human evidence concerning the potential of FPFs to influence skeletal muscle health through GMA modulation. Although emerging human studies suggest favourable effects on metabolic regulation, inflammatory pathways and gut microbial ecology, direct evidence that FPFs impact muscle protein synthesis, muscle mass or function remains limited. Furthermore, the literature is characterised by considerable heterogeneity in interventions and outcome measures. Studies integrating comprehensive microbiome characterisation, multi-omics approaches and direct skeletal muscle phenotyping are needed to determine whether FPFs confer advantages over conventional protein foods or more established microbiome targeted strategies such as probiotic supplementation. Such studies will be essential to elucidate their role in targeted nutritional strategies for ageing, metabolic disease, muscle atrophy, and physical performance.
|